Articles in Press

Articles in press have been peer-reviewed and accepted, which are not yet assigned to volumes/issues, but are citable by Digital Object Identifier (DOI).
Display Method:
MAPK signaling in plant metabolic control: integrating phytohormones with primary and secondary metabolism
Mengyun Wu, Shuqun Zhang, Mengmeng Zhang
, Available online  , doi: 10.1016/j.jgg.2026.11.005
Abstract (39) PDF (0)
Abstract:
Mitogen-activated protein kinase (MAPK) cascades are conserved signaling modules that integrate environmental and developmental cues to regulate plant metabolism. This review synthesizes current understanding of how MAPKKK–MAPKK–MAPK cascades coordinate primary metabolism, secondary metabolism, and phytohormone biosynthesis. MAPK cascades regulate photosynthetic capacity, nitrogen fixation and assimilation, and carbon flux through phosphorylation of key metabolic enzymes and transcription factors that balance growth with stress responses. In secondary metabolism, MAPK cascades converge on downstream transcription factors to regulate camalexin, indole glucosinolate, phenylpropanoid, flavonoid, lignin, terpenoid, and alkaloid biosynthesis. We further summarize how MAPKs are involved in the biosynthesis and signaling of ethylene, jasmonic acid, salicylic acid, auxin, and abscisic acid, revealing extensive crosstalk between kinase cascades and hormonal networks. We propose an integrative model in which MAPK cascades function as signaling hubs funneling upstream cues into three coordinated arms: transcription factor phosphorylation, direct modulation of structural enzymes, and phytohormone crosstalk. Future research should address key knowledge gaps, including incomplete MAPK cascades, limited understanding of MAPK substrates and their specificity in metabolic regulation, feedback mechanisms preventing metabolic overactivation, and the conservation of these principles across crop species. Understanding MAPK-mediated metabolic control holds promise for engineering stress-resilient, pathogen-resistant, and high-yield crops.
Lung development: from static snapshots to dynamic lineage trajectories
Qing Li, Yifei Bian, Yongting Fang, Hongbin Ji
, Available online  , doi: 10.1016/j.jgg.2026.11.004
Abstract (27) PDF (0)
Abstract:
The cellular diversity of the mammalian lung arises from tightly regulated spatiotemporal events. While traditional genetics and single-cell transcriptomics elucidate key mechanisms of lung organogenesis, current models of branching morphogenesis and lineage commitment rely heavily on endpoint mapping or inferred trajectories derived from discrete snapshots. These approaches lack the temporal and clonal resolution necessary to empirically trace lineage histories, leaving the precise timing of fate commitment and cellular origins largely obscured. In light of this, this review synthesizes the molecular and biomechanical drivers of lung development, highlighting ongoing debates regarding progenitor plasticity. We then introduce the Lung Spatiotemporal Cell Atlas of Phylogenetic Encoding (LungSCAPE), a conceptual roadmap that integrates emerging lineage-recording technologies with multi-omics to reconstruct clonal phylogenies. We outline its principles and discuss how this framework can generate a comprehensive atlas of lung developmental cell fates and address long-standing questions in the field.
Molecular regulation of anthocyanin biosynthesis in soybean
Minghui Xing, Xuejie Zhao, Dongmei Wang, Shuhua Liu, Huanbin Zheng, Liangliang Li, Siqi Yang, Zhaoming Qi, Qing Lu, Fengning Xiang, Xianzhong Feng, Weiqiang Li, Xiang Gao, Xiaojian Yin
, Available online  , doi: 10.1016/j.jgg.2026.11.001
Abstract (50) PDF (0)
Abstract:
Anthocyanins are specialized flavonoid pigments that contribute to pigmentation in soybean (Glycine max) seed coats, hypocotyls, stems, and flowers, as well as to photoprotection, stress adaptation, and nutritional quality. Although the core anthocyanin biosynthetic pathway is broadly conserved in plants, recent advances highlight anthocyanin accumulation in soybean as a systems-level output of flavonoid metabolic partitioning, tissue-specific transcriptional regulation, and environmental responsiveness. Here, we summarize recent progress in soybean anthocyanin biosynthesis and regulation, with emphasis on metabolic partitioning, tissue-specific regulatory modules, light-responsive pathways, and the functional diversification of anthocyanin-related MYB regulators. We further discuss unresolved questions related to pigment stabilization, stress-associated functions, multilayer regulatory networks, and precision manipulation of anthocyanin-related traits. These perspectives provide a framework for molecular breeding, regulatory engineering, and metabolic pathway optimization aimed at improving soybean quality, nutritional value, and stress resilience.
Developing a high-precision adenine base editor with AC motif preference through reverse engineering
Yifan Wang, Qiaoxia Xu, Lin Yang, Xiaoxi Huang, Yifeng Gui, Hui Yang, Jinhui Li, Yingnan Li
, Available online  , doi: 10.1016/j.jgg.2026.11.003
Abstract (40) PDF (0)
Abstract:
Exome-wide characterization of genetic diversity and population-specific functional variants in indigenous tribal populations of India
Sudeshna Datta, Deepak Jena, Arup Ghosh, Viplov Kumar Biswas, Sudarshana Jena, Mamuni Swain, Vinay More, Soumendu Mahapatra, Rasmita Mishra, Bhawna Gupta, Analabha Basu, Punit Prasad, Sunil Kumar Raghav
, Available online  , doi: 10.1016/j.jgg.2026.11.002
Abstract (28) PDF (0)
Abstract:
India harbors over 4600 endogamous ethnolinguistic groups shaped by multiple waves of migration, long-term isolation, and social stratification. Odisha, an eastern state of India, is home to 62 indigenous tribal groups, many of which remain genetically understudied. We perform high-coverage whole-exome sequencing of 13 tribal groups (n = 515) from Odisha, collectively termed OdiTribe. We identify 625,596 single-nucleotide variants (SNVs), of which 37.02% are rare, and 5.54% (n = 34,667) are unreported non-singletons (AC > 1). Population genetic analyses reveal that the OdiTribe populations broadly overlap with Austroasiatic- and Dravidian-speaking tribal populations, while also exhibiting heterogeneity across subgroups. Outgroup f3 statistics reveal high shared drift between Bonda and the Nicobarese. Approximately 43% of the individuals show elevated inbreeding coefficients (≥0.0156), and 14.17% carry ROH segments >8 Mb, indicative of recent consanguinity. Functional annotation identifies pathogenic variants in HBB, HBD, BTD, and BRCA2 genes, as well as multiple homozygous high-confidence predicted loss-of-function (HC-pLoF) variants across the dataset. Our study provides exome-wide insights into genetic diversity, endogamy, and population-specific variants of potential clinical relevance among endogamous indigenous tribal populations, underscoring the importance of including isolated and underrepresented groups in population and medical genetics research.
Activating death receptor 5 exposes a cancer cell vulnerability to natural killer and T cell attack
Zhen Chen, Gabriele Casagrande Raffi, Ziji Zhao, Chaopin Yang, Jian Chen, Xuezhao Feng, Yanxing Chen, Marit Steur, Pietro Morandi, Cor Lieftink, Siyi Lu, Huamei Xie, Haoran Zhong, Jing Wang, Lisa Willemsen, Hugo Horlings, Roderick Beijersbergen, Krijn K. Dijkstra, Song Gao, Ruidong Xue, Rene Bernards, Liqin Wang
, Available online  , doi: 10.1016/j.jgg.2026.10.015
Abstract (54) PDF (0)
Abstract:
Immunotherapy has revolutionized cancer care, yet resistance frequently undermines its efficacy. Although tumor-intrinsic resistance pathways have been mapped, how immune pressure regulates death-receptor vulnerability and anti-apoptotic buffering remains less defined. Using genome-wide CRISPR-Cas9 screening in cancer cells co-cultured with natural killer (NK) cells, we identify CFLAR, encoding cFLIP, as a dominant suppressor of death-receptor signaling that impairs NK- and T-cell cytotoxicity. CFLAR depletion sensitizes cancer cells to NK- and T-cell attack in vitro and in vivo. Single-cell and spatial transcriptomic analyses link high CFLAR expression to reduced lymphocyte infiltration and poorer responses to immune checkpoint blockade. Mechanistically, immune effector cell-induced inflammation increases CFLAR and death receptor 5 (DR5) expression in association with p38–MAPK signaling, balancing survival and death under immune pressure. The DR5 agonist conatumumab enhances lymphocyte-mediated killing and improves responses to cellular therapies, checkpoint blockade, and chemo-immunotherapy. Combining DR5 agonism with NKG2A blockade co-activates NK- and T-cell responses and overcomes anti-PD-1 resistance in MHC-I-deficient tumors. These findings identify the DR5–cFLIP axis as an immune-pressure-associated, pharmacologically targetable vulnerability and support rational combinations of DR5 agonists with multiple immunotherapies for refractory cancers.
Hybridization reshapes maize phenotypic plasticity through coordinated changes in environmental responses and genetic architecture
Yilan Cao, Xiangjian Gou, Zhenwei Zhang, Haitao Li, He Pei, Shuang Zhang, Lingju Zeng, Haoyan Yang, Xianlong Song, Xiaoyan Qian, Nannan Liu, Yingjie Xiao, Jianbing Yan, Tingting Guo
, Available online  , doi: 10.1016/j.jgg.2026.10.016
Abstract (84) PDF (0)
Abstract:
Phenotypic plasticity is central to crop adaptation under climate variability, yet how hybridization reshapes it remains unclear. Here, we integrate multi-environment phenotypes, environmental variables, and genomic data from 456 maize inbred lines and 912 corresponding hybrids evaluated across 11 environments to dissect the environmental and genetic bases of plasticity for flowering time, plant height, and ear weight. Reaction-norm analyses reveal two- to three-fold greater plasticity variation in inbreds than hybrids, indicating more uniform plastic responses in hybrids. Environmental association analyses demonstrate that inbreds and hybrids respond to the same environmental factors with different magnitudes or even opposite directions (e.g., correlations between diurnal temperature range and flowering time ranged from −0.73 to −0.14 in inbreds but 0.22 to 0.46 in hybrids). Genome-wide association analyses identify distinct parental loci for plasticity, with more loci and larger effects in hybrids. Functional validation shows that ZmBAF60_13, a chromatin-remodeling gene found only in the inbred panel, negatively regulates flowering-time plasticity. Integrating multiple environmental indices improves phenotype prediction accuracy by up to 16% over single-index models. Together, this study reveals that hybrids exhibit reduced plasticity variation and differ from inbreds in environmental responses and genetic associations, providing a framework for predicting crop performance under climate variability.
Lactylation at the crossroads: a metabolic-epigenetic-immune integrator in intestinal homeostasis and disease
Jingli Ao, Yingwei Li, Xinbei Lin, Bufu Tang, Lianxiang Luo
, Available online  , doi: 10.1016/j.jgg.2026.10.010
Abstract (39) PDF (0)
Abstract:
Lysine lactylation (Kla) is a metabolite-sensitive post-translational modification implicated in DNA repair, tumorigenesis, chemoresistance, and innate immunity. Recent studies also implicate Kla in intestinal disorders. This review synthesizes current evidence on protein lactylation in intestinal inflammation, inflammatory bowel disease, colorectal cancer, irritable bowel syndrome, intestinal fibrosis, radiation-induced injury, and enteric infection. We distinguish lactate-mediated receptor signaling and metabolic effects from covalent Kla, compare evidence across cell types and disease stages, and assess preclinical strategies targeting the lactate-lactylation axis. We also discuss microbiota-host metabolic crosstalk and mucus-penetrating nanodelivery, including the cross-border emulsification effect, an oil-in-water double-emulsion concept that describes traversal of the mucus barrier rather than a biochemical form of lactylation. Collectively, the available evidence supports a metabolism-epigenetics-immunity framework for gut pathophysiology, while highlighting unresolved questions concerning stereochemistry, site specificity, causality, safety, and prospective clinical validation.
A yak pangenome reveals whole-genome structural variations underlying domestication and introgression from cattle
Yi Liu, Yuan Xu, Jiawei Lu, Xian Guo, Xingdong Wang, Niyazbekova Zhannur, Gaoyue Liu, Zhengrong Yuan, Yu Jiang, Yi Ma
, Available online  , doi: 10.1016/j.jgg.2026.10.012
Abstract (135) PDF (0)
Abstract:
From loci to mechanisms: wheat autoimmunity as a window into plant immune activation
Wenling Li, Miao He, Fei Xiong, Qiuhong Wu, Haiyan Jia, Zhiyong Liu
, Available online  , doi: 10.1016/j.jgg.2026.10.013
Abstract (59) PDF (3)
Abstract:
Wheat (Triticum aestivum L.) is a major staple crop, and developing broad-spectrum resistance to multiple fungal pathogens is essential for sustainable production. Wheat autoimmunity (WAI) mutants represent genetic resources that spontaneously activate immune responses under non-stress conditions, exhibiting lesion-mimic or leaf-tip necrosis phenotypes and enhanced resistance to multiple foliar diseases, including powdery mildew and rusts. Increasing evidence indicates that WAI arises from dysregulation of immune homeostasis across multiple regulatory levels, involving receptor activation, cellular homeostasis, and membrane transport processes, ultimately converging on evolutionarily conserved defense outputs such as reactive oxygen species (ROS) accumulation, activation of defense-related genes, ion fluxes, and programmed cell death (PCD). Recent advances have identified key WAI-associated loci, including nucleotide-binding and leucine-rich-repeat receptors (NLRs), membrane proteins, regulatory components, and transporters, revealing mechanistically distinct pathways underlying constitutive immunity and its trade-off with growth. This review summarizes recent progress in the genetics and molecular mechanisms of WAI, critically assesses its roles in broad-spectrum, multi-disease resistance, and discusses strategies to minimize the growth and fitness costs. Furthermore, we propose that structure-guided allele optimization combined with spatiotemporal regulation of gene expression holds promise for the rational engineering of WAI alleles that confer durable disease resistance while minimizing associated fitness costs.
Protein palmitoylation in disease: decoding its crosstalk with ubiquitination and phosphorylation for pathogenic mechanisms and targeted therapeutics
Sainan Li, Huanjin Suo, Renqiang Hao, Ruohan Wu, Yinming Liang, Eryan Kong, Liu Cao, Binhui Zhou
, Available online  , doi: 10.1016/j.jgg.2026.10.014
Abstract (76) PDF (0)
Abstract:
Protein S-palmitoylation is a reversible lipid modification that governs protein stability, trafficking, and signaling. Rather than an isolated tag, it acts as a spatiotemporal routing signal within an integrated post-translational modification (PTM) network, engaging in three defined crosstalk modes with ubiquitination and phosphorylation: synergistic, antagonistic, and cascading. Although crosstalk with acetylation and myristoylation exists, this review focuses on the ubiquitination- and phosphorylation-centered networks most directly linked to palmitoylation in disease. The same palmitoylation event can stabilize or destabilize substrates, or activate or silence signaling, depending on the specific Zinc finger DHHC domain-containing protein (ZDHHC) isoform, subcellular location, local enzyme availability, and metabolic state. We further introduce the “PTM context signature”, a four-variable predictive framework for disease-specific crosstalk outcomes. By dissecting palmitoylation-centered crosstalk through a disease-oriented lens, we examine how its dysregulation drives oncogenesis, inflammation, metabolic disorders, and neurodegeneration. We then translate these mechanistic principles into a network-centric therapeutic framework, evaluating ZDHHC-directed modulators, peptide-based interface disruptors, and crosstalk-informed combinations that target context-specific nodes and anticipate therapy resistance. Viewing palmitoylation as an integrated component of the cellular PTM network, rather than as an isolated lipid tag, provides a framework for understanding disease pathogenesis and for targeting PTM networks rather than single modifications.
mRNA alternative splicing dysregulation in oocytes causes premature ovarian failure
Wei Yue, Yi-Xiao Li, Di Gao, Rui-Ying Yuan, Hong-Yong Zhang, Shu-Chen Liu, Xu-Feng Li, Zheng-Hui Zhao, Qing-Yuan Sun, Tie-Gang Meng
, Available online  , doi: 10.1016/j.jgg.2026.10.011
Abstract (87) PDF (0)
Abstract:
Primordial follicle survival determines female reproductive lifespan, yet the post-transcriptional mechanisms governing this process remain incompletely understood. Here, we show that the splicing regulator SRSF2 is required for maintaining the ovarian follicle reserve through the regulation of RNA processing in oocytes. Oocyte-specific ablation of Srsf2 using Gdf9-Cre causes complete female infertility, progressive follicular depletion, and increased follicular apoptosis. Linear amplification of complementary DNA ends and sequencing (LACE-seq) identifies 1609 SRSF2-bound transcripts enriched in RNA-regulatory and cellular homeostasis-related processes. Subsequent RNA-seq analysis reveals broad transcriptomic and alternative-splicing dysregulation after Srsf2 loss, including prominent exon 7 skipping of the oocyte-specific transcription factor Nobox and a marked reduction in full-length NOBOX protein. Supplementation with correctly spliced Nobox mRNA partially rescues follicular developmental defects in vitro, supporting Nobox as a functionally important downstream target rather than the sole mediator of the Srsf2-deficient phenotype. Together, these findings identify SRSF2-dependent RNA processing as an important regulator of oocyte and follicle homeostasis and suggest that disruption of SRSF2-regulated splicing may contribute to ovarian insufficiency.
Ancient-modern genomics reveals the geographic origins of the historical Huangqi specimen from the Qing Imperial Pharmacy
Xiaojing Han, Minzhen Yin, Fengyuan Li, Zhengyang Yang, Yuanyang Cai, Yucheng Wang, Huasheng Peng, Luqi Huang
, Available online  , doi: 10.1016/j.jgg.2026.10.009
Abstract (84) PDF (0)
Abstract:
LSD1 exerts crucial functions to drive the progression of meiotic prophase I during spermatogenesis
Tianhua Zhu, Shaogang Qin, Yingying Dong, Hong Li, Qingfeng Yang, Chenmin Zhou, Yibing Bao, Zijian Zhu, Fengchao Wang, Guoliang Xia, Bo Zhou, Chao Wang, Zhengpin Wang
, Available online  , doi: 10.1016/j.jgg.2026.10.008
Abstract (52) PDF (0)
Abstract:
Epigenetic modifications are critical determinants governing the proper initiation and progression of gametogenesis. Accumulating evidence has indicated that epigenetic dysregulation underlies the pathogenesis of infertility. However, detailed information on the individual powerful roles of these epigenetic enzymes in facilitating programmed meiosis remains elusive. This study indicates that lysine-specific demethylase 1 (LSD1) plays critical roles during meiotic prophase I in mouse spermatogenesis. Germ cell-specific deletion of the Lsd1 gene in male mice caused meiotic arrest at the pachytene stage, ultimately leading to infertility. In primary spermatocytes of Lsd1-deficient mice, compromised homologous chromosome synapsis and DNA double-strand break repair activated the DNA damage checkpoint pathway, triggering spermatocyte apoptosis. Mechanistically, LSD1 directly binds to the promoters of meiotic genes, including Prdm9 and Rpa2, to regulate their expression during meiosis. PRDM9 and RPA2 are upregulated upon LSD1 loss and are essential for homologous recombination and DNA double-strand break repair. The promoters of Prdm9 and Rpa2 are also occupied by H3K4me2, whose levels increase upon LSD1 loss. Collectively, this study highlights the crucial role of LSD1 in orchestrating meiotic genes, possibly through its demethylase activity during spermatogenesis, and may help explain the pathology of non-obstructive azoospermia in humans.
DNAJC17 regulates RNA splicing in auditory hair cells and is required for hearing
Luping Zhang, Ziyang Wang, Wenya Fan, Yiliang Huang, Chenxuan Yong, Yuecen Sun, Qingyue Yuan, Guang-Jie Zhu, Ao Li, Tao Yang, Qing Liu, Guoqiang Wan
, Available online  , doi: 10.1016/j.jgg.2026.10.007
Abstract (98) PDF (0)
Abstract:
RNA splicing is essential for gene regulation, but its role in maintaining cochlear hair cell function remains incompletely understood. Here, we identify a heterozygous missense variant (c.401A>C, p.E134A) in DNAJC17, which encodes a spliceosome-associated protein, that segregates with adult-onset progressive hearing loss in a multigenerational family. Hair cell-specific deletion of Dnajc17 in mice results in profound deafness with progressive hair cell degeneration, establishing its essential role in cochlear maintenance. Hair cells lacking DNAJC17 exhibit marked stereocilia disorganization accompanied by widespread alternative splicing defects in the cochlea. Notably, the dual-specificity protein kinase Clk1 that regulates pre-mRNA splicing shows altered exon inclusion, consistent with disruption of splicing control networks. In addition, Dnajc17 p.E134A knock-in mice develop adult-onset progressive hearing loss with stereocilia and mitochondrial abnormalities but preserve hair cell survival, recapitulating the human phenotype. Together, these findings identify DNAJC17 as a causative gene for autosomal dominant hearing loss and identify RNA splicing regulation as a key mechanism required for long-term maintenance of cochlear hair cell integrity.
The photoperiod regulator E1 family negatively modulates salt tolerance by directly repressing GmCIPK25 expression in soybean
Miao Zhang, Aimin Wu, Ziyuan Deng, Xiaohuan Sun, Yuehua Yu, Zhiyong Ni, Xiaoyan Lin, Fanjiang Kong, Qing Sang, Zhihui Sun
, Available online  , doi: 10.1016/j.jgg.2026.10.006
Abstract (111) PDF (0)
Abstract:
Soybean (Glycine max [L.] Merr.) is an important crop whose productivity is severely constrained by salinity. Here, we identify the photoperiod regulator E1 family as negative regulators of soybean salt tolerance. Under salt stress, the e1 e1la e1lb triple mutant (e1-3m) show higher antioxidant enzyme activities and proline accumulation, together with reduced hydrogen peroxide and malondialdehyde levels, whereas E1 overexpression increases salt sensitivity. Integrated RNA-seq and ChIP-seq analyses identify GmCIPK25, a positive regulator of salt tolerance, as a direct target of E1. E1, E1La, and E1Lb bind directly to the GmCIPK25 promoter and repress its transcription. Knockdown of GmCIPK25 attenuates the enhanced salt tolerance of e1-3m, while loss of GmCIPK25 function in the wild-type background increased salt sensitivity. Thus, the E1 family negatively regulates salt tolerance by repressing GmCIPK25, which compromises reactive oxygen species (ROS)-scavenging capacity and osmotic adjustment. These findings link photoperiodic regulation to salt-stress responses and identify GmCIPK25 as a potential target for improving soybean salt tolerance.
GmBBX13 and GmBBX4a/b antagonistically regulate GmCML19 transcription to fine-tune leaf senescence in soybean
Lu Zhai, Rongbo Yang, Zhaoqing Song, Yuntao Xiao, Fengyue Zhao, Wubin Wang, Dongqing Xu
, Available online  , doi: 10.1016/j.jgg.2026.09.010
Abstract (118) PDF (0)
Abstract:
Leaf senescence is a critical developmental process that fundamentally determines crop yield and quality. Here, we report that the B-box CONTAINING PROTEIN 13 (GmBBX13) negatively regulates leaf senescence in soybean. Overexpression of GmBBX13 in soybean significantly delays leaf senescence onset. GmBBX13 physically interacts with two closely related B-box proteins, GmBBX4a and GmBBX4b. Loss of GmBBX4a and GmBBX4b function markedly delays leaf senescence, indicating that GmBBX4a/b serve as positive regulators of senescence. Transcriptomic profiling reveals that GmBBX13 and GmBBX4a/b antagonistically co-regulate approximately 4,700 target genes. Specifically, GmBBX13 represses, whereas GmBBX4a/b activate, the expression of GmCML19, a gene encoding CALMODULIN-LIKE PROTEIN 19, whose product promotes leaf senescence. Collectively, our study reveals a molecular mechanism whereby GmBBX13 and GmBBX4a/b constitute antagonistic regulatory modules that fine-tune the timing of leaf senescence in soybean, offering strategies to improve yield and extend the grain-filling period.
Corrigendum to “Migfilin promotes migration and invasion in glioma by driving EGFR and MMP-2 signalings: A positive feedback loop regulation” [Journal of Genetics and Genomics 44 (2017) 557-565]
Yunwei Ou, Qingnan Wu, Chuanyue Wu, Xuefeng Liu, Yongmei Song, Qimin Zhan
, Available online  , doi: 10.1016/j.jgg.2026.09.009
Abstract (84) PDF (1)
Abstract:
LD protein-associated GAF is functionally linked to age-dependent transcriptional regulation in glia
Qingyuan Wei, Zhi-Xiang Xu, Lihong Sheng
, Available online  , doi: 10.1016/j.jgg.2026.09.007
Abstract (75) PDF (0)
Abstract:
Decoupling skeletal and soft-tissue genetic effects in human craniofacial morphology
Yi Li, Wei Qian, Manfei Zhang, Yingzhi Wu, Xiang Jiao, Lei Wei, Ziyi Xiong, Siyuan Du, Ting Li, Xiongzheng Mu, Yong Zhou, Yongchu Pan, Fan Liu, Sijia Wang
, Available online  , doi: 10.1016/j.jgg.2026.09.006
Abstract (133) PDF (0)
Abstract:
Natural variation in the OsAFB2 promoter enhances rice seedling emergence under submergence
Hui Jiang, Xionglin Zhang, Xuebin Yang, Xinjun Yi, Meiqi Wu, Kehan Di, Hongpu Chen, Kangkang Deng, Hao Wang, Weilan Chen, Ting Li, Bin Tu, Hua Yuan, Zhaohui Zhong, Shigui Li, Bingtian Ma, Liangzhu Kang, Shiwen Tang, Yuping Wang, Jiawei Xiong, Peng Qin
, Available online  , doi: 10.1016/j.jgg.2026.09.008
Abstract (146) PDF (0)
Abstract:
Seedling emergence is a key adaptive trait for water direct-seeded rice, yet its genetic basis remains poorly understood and elite alleles are limited. Here, a genome-wide association study of 307 rice accessions identifies qSLS4, a locus controlling shoot elongation during seedling emergence under submergence. OsAFB2 encodes an auxin receptor and is the causal gene for qSLS4. Its knockout promotes shoot elongation by increasing cell length and cell number, thereby improving seedling emergence rather than promoting coleoptile elongation, and also reduces plant height with only a minor effect on yield per plant. Expression and pharmacological analyses suggest that OsAFB2 affects the final emergence phenotype through auxin-related regulation. In a candidate gene association analysis, a single nucleotide substitution (C to A) at –941 bp in the promoter of OsAFB2 is found to alter its expression level and contribute to variation in seedling emergence. OsDREB6 preferentially activates the OsAFB2HapC promoter, whereas this substitution weakens the activation, resulting in lower expression and enhanced seedling emergence in OsAFB2HapA. Together, these findings establish that OsAFB2 acts as a negative regulator of rice seedling emergence and uncover a promoter variant with potential value for improving water direct-seeded rice.
Transcriptional reprogramming and epigenetic regulation in Paramecium caudatum: a new model for studying extreme germline DNA elimination
Yunyi Gao, Rui Wang, Yueer Yu, Naomi A. Stover, Feng Gao
, Available online  , doi: 10.1016/j.jgg.2026.09.011
Abstract (102) PDF (0)
Abstract:
Metazoans suppress transposons via small RNA-mediated silencing mechanisms, whereas ciliates have evolved a unique repressed strategy by physically eliminating transposons during somatic macronuclear genome development, offering unique models for transposon regulation and programmed DNA rearrangement. The ciliate Paramecium caudatum exhibits the most extreme known DNA elimination among eukaryotes, removing > 98% of its germline genome, yet its molecular basis and underlying mechanisms remain unknown. Here we present a high-quality, near-chromosome-level somatic macronuclear genome assembly, which is exceptionally compact, with ultra-short untranslated regions, and high gene density. Comprehensive transcriptome analysis across 15 life cycle stages reveals extensive transcriptional reprogramming during sexual reproduction. A total of 8260 genes are identified as developmentally regulated, forming 10 co-regulated clusters. While the core DNA elimination machinery appears to be conserved, many regulatory genes exhibit divergent expression patterns compared to related species. We further demonstrate that the histone methyltransferases Ezl1 and Ezl3 deposit the repressive marks H3K9me3/H3K27me3 in a nuclear-specific manner, showing functional specialization distinct from those previously reported in other ciliates. Our findings establish P. caudatum as an exceptional model for studying extreme genome rearrangement and provide new insights into the evolution of epigenetic mechanisms controlling transposon silencing and genome stability in eukaryotes.
PIGSVOME: an integrated resource of porcine genomic structural variants for functional genomics and breeding applications
Qi Bao, Lingsen Zeng, Hui Wen, Junjian Wang, Hongwei Yin, Yongjin Zhang, Lingbin Liu, Francesco Tiezzi, Garrett See, Clint Schwab, Chiara Gini, Christian Maltecca, Kui Li, Lingzhao Fang, Yongju Zhao, Jicai Jiang, Guoqiang Yi
, Available online  , doi: 10.1016/j.jgg.2026.08.009
Abstract (305) PDF (2)
Abstract:
Biallelic ATP8B3 variants cause male infertility and fertilization failure in humans
Daijuan Chen, Guicheng Zhao, Yunchuan Tian, Xiang Wang, Xinyao Tang, Shikun Zhao, Jun Ma, Xiaodong Wang, Yihong Yang, Ying Shen
, Available online  , doi: 10.1016/j.jgg.2026.09.004
Abstract (119) PDF (0)
Abstract:
A wild soybean MADS-box gene GsAGL62 improves seed weight by enhancing cytokinin signaling and cell proliferation
Fei Wu, Wei Li, Dezhou Hu, Zhongyi Yang, Zhen Wang, Zhiyu Wang, Xiaoyue Su, Yifei Yang, Shupeng Dong, Cui Xuan, Yu Zhang, Qinxin Song, Deyue Yu, Fang Huang
, Available online  , doi: 10.1016/j.jgg.2026.09.005
Abstract (121) PDF (0)
Abstract:
Soybean seed weight is a key yield determinant, but the transcriptional mechanisms connecting hormone signaling to seed growth are poorly understood. Here, we identify GsAGL62, a wild soybean MADS-box transcription factor located within a previously mapped hundred-seed weight (HSW) locus and a domestication-associated selective sweep. Functional analyses show that overexpression of GsAGL62 in cultivated soybean significantly increases HSW, whereas ethyl methanesulfonate (EMS)-induced gmagl62 mutants reduce it. Integrated transcriptomic and metabolomic analyses reveal that GsAGL62 enhances cytokinin accumulation and signaling cytokinin-associated responses, accompanied by increased expression of genes involved in cell proliferation. Mechanistically, GsAGL62 directly binds to the promoter of the conserved growth inhibitor GmATPK2 and represses its transcription. Consistently, independent EMS-induced gmatpk2 mutants exhibit increased seed weight, supporting GmATPK2 as a downstream negative regulator of seed growth. Population genetic analyses further reveal strong differentiation of GsAGL62 promoter haplotypes during soybean domestication and improvement. These haplotypes show differential promoter activities and are associated with distinct agronomic performance, suggesting that cis-regulatory variation at GsAGL62 contributes to its selection during soybean improvement. Collectively, our findings establish a regulatory module linking GsAGL62 to cytokinin-associated responses, cell proliferation, and seed growth, and highlight GsAGL62 as a potential target for soybean yield improvement.
A telomere-to-telomere genome assembly of Medicago ruthenica enables QTL mapping and marker development for pod shattering resistance
Qiang Li, Mingliang Wang, Wenying Wang, Jing Gao, Yuan Song, Fantong Meng, Linqing Yu, Jinglong Li, Chien-Hsun Huang, Jun Li
, Available online  , doi: 10.1016/j.jgg.2026.08.001
Abstract (296) PDF (0)
Abstract:
Genetic architecture and inheritance patterns of Leber hereditary optic neuropathy among 419 Chinese pedigrees carrying the ND4 11778G>A mutation
Yanchun Ji, Feilong Meng, Yinglong Gao, Zhipeng Nie, Yunfan He, Juanjuan Zhang, Ya Li, Minglian Zhang, Jianyong Wang, Yongming Zhang, Shihui Wei, Jun Qin Mo, Min-Xin Guan
, Available online  , doi: 10.1016/j.jgg.2026.08.007
Abstract (188) PDF (0)
Abstract:
Leber's hereditary optic neuropathy (LHON) is a mitochondrial disease mainly driven by the m.11778G>A mutation, and its incomplete penetrance and diverse inheritance patterns remain unclear. This study integrates clinical and genetic analyses of 419 Han Chinese pedigrees carrying this mutation, covering 5262 matrilineal relatives. Distinct phenotypic heterogeneity emerges, including sporadic, maternal and complex transmission patterns; 209 pedigrees contain only single affected individuals, which suggests that the m.11778G>A mutation alone fails to cause disease. Full mitochondrial DNA sequencing and haplogroup screening identify multiple mitochondrial genetic modifiers. Haplogroups D4j, M7, M9, and M10 are significantly enriched in maternally inherited families with elevated disease penetrance. Haplotype-specific variants ND4 11696G>A, ND1 3394T>C, and ND6 14502T>C synergistically aggravate mitochondrial dysfunction together with m.11778G>A, and secondary mtDNA mutations disrupting complex I or mitochondrial tRNA metabolism also raise disease susceptibility. Nuclear modifiers PRICKLE3 and YARS2, as well as X-linked sex-specific regulatory factors, are also identified. Overall, LHON results from the interaction of mitochondrial and nuclear genetic factors. This research constructs a comprehensive genetic landscape of LHON, highlights the vital role of modifier genes, and provides theoretical support for precision therapies targeting mitochondrial and nuclear pathways.
GmNIGT2a/2b-mediated transcriptional cascade buffers high-nitrogen-triggered nodule senescence in soybean
Tianli Ge, Xiaolei Yao, Yuan Li, Xin Wang, Suning Li, Fang Xie, Zhichang Chen, Yuefeng Guan
, Available online  , doi: 10.1016/j.jgg.2026.08.006
Abstract (177) PDF (0)
Abstract:
Soil inorganic nitrogen (N) availability is a critical determinant of symbiotic nitrogen fixation efficiency, making it essential for legumes to respond appropriately and effectively to N fluctuations. Here, we identify a pair of high N response factors GmNIGT2a/2b (NITRATE-INDUCIBLE GARP-TYPE TRANSCRIPTIONAL REPRESSOR). GmNIGT2a/2b are activated by GmNLP4a/b (NIN-like proteins) under high N. The nodules of gmnigt2a/2b double mutants exhibit exacerbated reduction in nitrogenase activity and accelerated senescence in response to high N. Integration of RNA-seq and DAP-seq analyses reveals that GmNIGT2a/2b negatively regulate a suite of core N-induced genes, including NAC, WRKY, and bZIP transcription factors as well as trehalose metabolism genes. GmNIGT2b binds to the promoters of SNAP3 and NAC039 and represses their expression, thereby delaying nodule senescence. Our results suggest that GmNIGT2a/2b-mediated transcriptional regulation prevents excessive nodule senescence in response to high N, highlighting the complexity of transcriptional reprogramming for environmental adaptation in nodules.
Decoding immunoglobulin repertoires in zebrafish reveals distinct paradigms for adaptive immune diversification in ancient vertebrates
Nan Zhang, Chongbin Hu, Jianfei Ji, Dongdong Fan, Aifu Lin, Lixin Xiang, Ye Chen, Jianzhong Shao
, Available online  , doi: 10.1016/j.jgg.2026.08.005
Abstract (141) PDF (0)
Abstract:
The evolutionary origins of adaptive immune diversification remain a central question in immunology. Using zebrafish as an early vertebrate model, we systematically dissect the distinct diversification strategies of ancient IgM and IgZ antibody repertoires. While both isotypes share a common V gene pool, they exhibit lineage-specific V/J usage biases during independent V(D)J recombination. Heavy-chain rearrangement employs the recombination-activating gene–non-homologous end joining (RAG–NHEJ) pathway, with IgM demonstrating greater junctional diversity than IgZ. Antigen-driven somatic hypermutation (SHM) primarily targets IgM via activation-induced cytidine deaminase (AID)-primed base excision repair/mismatch repair within primitive germinal center-like clusters. However, unbiased targeting across complementarity determining and framework regions, together with constrained CXCR5–CXCL13 signaling, limits mutation frequency and affinity maturation.Notably, we identify an AID-independent SHM pathway mediated by APOBEC2a that specifically targets TCG/CGT motifs. Furthermore, light-chain V–J recombination is uniquely processed via microhomology-mediated end joining, restricting CDR3 diversity compared with heavy-chain. Conversely, IgZ+ B cells are biased toward plasma cell differentiation, exhibit loosely organized distribution, and undergo minimal antigen-driven SHM, aligning with rapid mucosal defense. Our findings reveal both conserved and distinct diversification mechanisms in teleosts, illuminating layered evolutionary strategies balancing receptor diversity and self-tolerance in early vertebrates.
A ruminant lncRNA atlas uncovers association with epithelial development and barrier integrity
Ge Han, Junru Tian, Fenglei Zhang, Botong Zhou, Yi Wan, Huishan Yue, Miaomiao Chen, Lei Chen, Wen Wang, Zan Huang
, Available online  , doi: 10.1016/j.jgg.2026.08.002
Abstract (183) PDF (0)
Abstract:
TBGR-v2: an integrated pan-genome and evolutionary resource for Brassicaceae genomic research
Lusheng Guo, Zipeng Meng, Xiao Ma, Zhuo Liu, Chenhao Zhang, Jinghua He, Mengyao Shi, Chengcheng Zou, Rong Zhou, Xiaoming Song
, Available online  , doi: 10.1016/j.jgg.2026.08.004
Abstract (320) PDF (0)
Abstract:
Identification and quantification of alternative polyadenylation sites in single cell RNA-seq data using scPAISO
Yongjie Liu, Peiwen Xiong, Songyang Li, Xinjia Liu, Tao Liu, Qinglan Yang, Shuting Wu, Hongyan Peng, Yana Li, Lingling Zhang, Yafei Deng, Yong Zhu, Junping Wang, Youcai Deng
, Available online  , doi: 10.1016/j.jgg.2026.07.013
Abstract (175) PDF (0)
Abstract:
Alternative polyadenylation (APA) generates transcript diversity by producing mRNA isoforms with distinct 3′ untranslated regions (3′ UTRs) or coding sequences. Existing single-cell RNA sequencing (scRNA-seq) methods for APA analysis primarily rely on Read2 data, which lacks precise cleavage site (CS) information and limits accurate polyadenylation site (PAS) identification. Here, we present single-cell PolyAdenylation ISOform quantification (scPAISO), a computational pipeline that leverages the often-discarded Read1 from 3′ tag-based scRNA-seq to enable de novo PAS identification and PAS isoform quantification. Unlike existing approaches, scPAISO directly captures mRNA 3′ end cleavage sites, resulting in stronger AAUAAA motif enrichment, sharper PAS peaks, and improved spatial resolution for resolving closely spaced PASs. Across multiple datasets and biological systems, scPAISO robustly identified PASs and quantified APA dynamics, revealing stage-specific 3′ UTR lengthening during hematopoietic differentiation, widespread 3′ UTR remodeling in systemic sclerosis, and tissue-specific polyadenylation preferences associated with distinct RNA-binding protein programs in mice. scPAISO provides an accurate and scalable framework for single-cell APA analysis, enabling high-resolution characterization of post-transcriptional regulation and transcriptome diversity in development, physiology, and disease.
Upgraded genome with ultra-long Nanopore reads facilitates resistance-gene discovery in sugarcane
Qibin Wu, Chao Zhang, Ge Wang, Wenhui Zou, Dongjiao Wang, Yuanyuan Zhang, Wanying Zhao, Peilin Wang, Zhen Zeng, Haifeng Wang, Youxiong Que
, Available online  , doi: 10.1016/j.jgg.2026.07.012
Abstract (305) PDF (0)
Abstract:
Large-scale low-coverage whole-genome sequencing reveals the genetic architecture of wool and growth traits in fine-wool sheep
Jianxin Shi, Changchun Lin, Peiling Wei, Juncheng Huang, Hanikezi Tulafu, Nana Yan, Sen Wang, Zhangyuan Pan, Weiwei Wu
, Available online  , doi: 10.1016/j.jgg.2026.07.006
Abstract (378) PDF (0)
Abstract:
Breeding sheep with superior growth performance and wool quality is essential for the sustainability of the fine-wool sheep industry. In this study, we perform low-coverage whole-genome sequencing (lcWGS) on 3842 individuals from 5 sheep breeds (4 fine-wool and 1 semi-fine wool) and generate a large genomic dataset. By comparing these breeds with coarse-wool sheep, we characterize the genomic landscape and selection signatures of fine-wool sheep. We identify several known functional genes associated with hair follicle development and skin morphology, including EGFR, KRT74, EDAR, EREG, and GLI2. Furthermore, GWAS of 19 traits identifies 156 candidate genes significantly associated with growth and wool characteristics, including LCORL for body size, EGFR for clean wool yield, and PRDM1 for fiber diameter. Notably, EGFR is detected in both GWAS and selection signature analyses, indicating its important role in phenotype formation and historical selection. Overall, our findings reveal the genetic basis of growth and wool traits in fine-wool and semi-fine wool sheep, highlight EGFR, LCORL, and PRDM1 as candidate genes, and provide valuable genomic resources and candidate markers for future functional validation and molecular breeding.
Incomplete penetrance in neurogenetic disorders: current insights and emerging perspectives
Jiao-Jiao Xu, Dian-Fu Chen, Zhi-Ying Wu
, Available online  , doi: 10.1016/j.jgg.2026.07.005
Abstract (246) PDF (0)
Abstract:
Neurogenetic disorders have been recognized clinically for decades, and advances in clinical and genetic studies have identified more than 1700 monogenic causes of neurological diseases. Various types of mutations, including missense, truncating, and repeat expansions, have been reported in patients with neurogenetic disorders. It is now recognized that incomplete penetrance is common, with some individuals carrying disease-causing mutations remaining clinically unaffected. However, there is currently no comprehensive conceptual framework to categorize or explain these observations. Here, we review and integrate decades of evidence on incomplete penetrance in neurogenetic disorders to clarify its biological and mechanistic bases. Accordingly, four major themes are identified, encompassing genetic modifiers, epigenetic modifications, mosaicism, and environmental factors. These factors may act independently or interactively to influence pathogenic burden and functional network balance, ultimately determining whether a pathogenic mutation manifests clinically. Based on these insights, we highlight emerging perspectives and propose future research to fill gaps in our understanding. A deeper understanding of incomplete penetrance will be essential for generating genetic insights to support more effective genetic counseling, therapeutic interventions, and disease prevention in neurogenetic disorders.
Beyond radiogenomics: advancing imaging integration and multi-omics horizons in cancer precision medicine
Yuming Chen, Huiqin Li, Silu Chen, Hongsheng Ji, Junyi Xin, Mulong Du
, Available online  , doi: 10.1016/j.jgg.2026.07.003
Abstract (278) PDF (0)
Abstract:
Cancer remains the leading cause of death worldwide, presenting substantial challenges to precision medicine due to its complex heterogeneity. Radiogenomics, as a method combining quantitative radiologic data with genomic information, provides a robust analysis framework to assess tumor heterogeneity and cancer progression. Here, we summarize the application of radiogenomics into two key fusion methods: feature-level and decision-level fusion. Feature-level fusion combines multimodal data into a rich feature set to improve the predictive power of models, while decision-level fusion integrates decision results from multiple independent models to improve robustness and reliability. Furthermore, we explore the integration of radiomics with various omics technologies, including transcriptomics, metabolomics, and proteomics. This integration enables a deeper understanding of the dynamic tumor microenvironment, metabolic dysregulation, and cancer progression mechanisms. Finally, we provide a detailed overview of publicly available datasets relevant to radiogenomics research, such as The Cancer Imaging Archive, cBioPortal, UK Biobank and Human Connectome Project; and further describe multiple types of omics data and sample characteristics for each resource for the benefit to readers. In summary, this review charts a path beyond radiogenomics by advancing radiomics and multi-omics horizons to transform precision medicine in cancer.
Evolution of resistance to KRASG12C inhibitor in a non-small cell lung cancer responder
Jiahui Xu, Shijia Wang, Ziming Wang, Jumin Huang, Chun Xie, Tingting Zhao, Yuqing Niu, Yabing Cao, Chen Ming, Elaine Lai-Han Leung
, Available online  , doi: 10.1016/j.jgg.2026.07.002
Abstract (171) PDF (0)
Abstract:
The ZmGF14-6–ZmCDC48 module controls endosperm cell proliferation and kernel weight in maize
Long Zhang, Yongbin Dong, Ju Wu, Feiyan Li, Han Wang, Weigang Yao, Yan Wang, Lingwei Hu, Yiming Yuan, Ran Li, Shengkun Li, Wenyu Li, Yuling Li
, Available online  , doi: 10.1016/j.jgg.2026.07.001
Abstract (253) PDF (0)
Abstract:
Tapt1 deficiency in mice impairs pulmonary lipid homeostasis and normal postnatal respiration by targeting ABCA3 for autophagy-lysosomal degradation
Dandan Tan, Lu Cai, Li Yuan
, Available online  , doi: 10.1016/j.jgg.2026.06.012
Abstract (148) PDF (0)
Abstract:
TAPT1, which encodes a highly conserved multi-pass transmembrane protein termed transmembrane anterior posterior transformation 1 (TAPT1), has been reported as a disease-causing gene, but its physiological role in mice remains to be elucidated. Using Tapt1 knockout and knock-in mice, we indicate that TAPT1 localizes to the endoplasmic reticulum and that Tapt1 deletion causes neonatal lethality due to atelectasis-induced respiratory distress. We further reveal that TAPT1 interacts with the ATP-binding cassette transporter A3 (ABCA3), thereby regulating autophagy-lysosomal degradation of ABCA3. ABCA3 insufficiency reduces surfactant lipids, leading to defective lamellar body formation and surfactant production. This study presents a hitherto unrecognized pathway for ABCA3 protein degradation. Our findings may advance the understanding of genetic determinants of neonatal respiratory distress syndrome, providing valuable insights into the treatment of lung surfactant disorders.
Gapless comparative genomics reveals effector landscape and adaptive selection signatures in Plasmodiophora brassicae
Jianbiao Wang, Shuxin Xuan, Xizhe Sun, Jingning Li, Guanghua Hao, Parham Haddadi, Tayab Soomro, Yanhua Wang, Shuxing Shen, Chunyang Wang, Mengyang Liu, Jianjun Zhao, M. Hossein Borhan, Lisong Ma
, Available online  , doi: 10.1016/j.jgg.2026.06.011
Abstract (289) PDF (0)
Abstract:
Characterizing selection signatures in coding and noncoding regions of 14,886 cancer genomes
Mengyue Zheng, Xiwei Sun, Junren Hou, Minmin Guo, Xinran Liu, Wen Yang, Jian Yang
, Available online  , doi: 10.1016/j.jgg.2026.06.008
Abstract (143) PDF (0)
Abstract:
Clonal selection drives cancer development, but quantifying selection on noncoding somatic mutations remains largely unexplored. Here, we introduce dNdS-Fun, an extension of the dN/dS framework to quantify selection of both coding and noncoding somatic mutations, thereby enhancing the discovery of driver genes. Applying dNdS-Fun to whole-genome sequencing data from 14,886 cancer patients across 31 cancer types, we identify 175 genes under positive selection across multiple cancer types or datasets, as well as 20 previously known driver genes detected through noncoding mutations. Of these, 69 are previously unrecognized as drivers, and 30 are identified solely through noncoding mutations. Furthermore, we observe evidence of negative selection throughout the genome, with significant enrichment in essential and cancer-dependent genes. Sixteen genes exhibit an overall signature of negative selection but show positive selection in noncoding elements, indicating both their conserved functions and adaptive regulatory roles in tumorigenesis. Our study reveals evidence consistent with negative selection of noncoding mutations, providing important insights for future research on their roles in cancer progression.
Tissue-resolved comparative developmental transcriptomics reveals auxin-associated regulatory shifts underlying divergent fruit tissue origins in pomes and drupes
Jiakai Liang, Qiaoming Yu, Yan Wang, Man Xie, Xiangxiang Chen, Peien Feng, Yunxiao Liu, Mingjun Li, Yaqiang Sun, Jia-Long Yao, Fengwang Ma, Yangjun Zou, Tao Zhao
, Available online  , doi: 10.1016/j.jgg.2026.06.006
Abstract (202) PDF (0)
Abstract:
Fruit type divergence in closely related species offers a powerful lens into the evolution of plant developmental programs. Within the Rosaceae family, pomes (e.g., apple) and drupes (e.g., peach) share a common ancestry but exhibit a fundamental difference: pomes recruit the hypanthium to form fleshy tissue, while drupes develop flesh solely from the ovary wall. To uncover the regulatory basis of this divergence, we conduct a comparative tissue-resolved developmental transcriptomic analysis across these fruit types. We identify distinct patterns of auxin-related networks, with pomes showing preferential activation of auxin signaling, transport, and biosynthetic pathways within the hypanthium, accompanied by differences in gibberellin responses. Through integrated network analysis, we identify a set of candidate regulators, including MADS-box, YABBY, and AP2 family genes, that interface with auxin signaling in a tissue-specific manner. Furthermore, we delineate a putative regulatory axis in which upstream transcription factors (FIL, ANT, AP1), potentially modulated by DOF3.2, are differentially associated with the auxin biosynthesis framework (e.g., YUC6) between pomes and drupes. Our findings suggest that fruit type divergence is associated with the spatial redeployment of auxin-related regulatory modules across tissues, providing a molecular framework for understanding fleshy fruit diversification and a basis for future evolutionary-developmental studies.
Yorkie/Scalloped-OVOL-Rac1 axis controls insect wing development by promoting cell proliferation
Jing Zhang, Yanan Zhang, Babar Hussain Chang, Caiyan Huo, Weimin Liu, Jianzhen Zhang, Xiaoming Zhao
, Available online  , doi: 10.1016/j.jgg.2026.06.010
Abstract (121) PDF (1)
Abstract:

The regulation of organ size is a fundamental question in developmental biology, and insect wings provide a powerful model for elucidating the genetic mechanisms underlying morphogenesis. Although the conserved Hippo signaling pathway plays a central role in controlling tissue growth, its precise regulatory network during wing development remains incompletely understood. Here, we identify the zinc finger transcription factor OVOL as a critical mediator of Hippo signaling in insect wing development. We indicate that OVOL is essential for normal wing formation in both Locusta migratoria and Drosophila melanogaster, regulating cell proliferation and trichome patterning. Through transcriptomic analysis and functional validation, we further identify the small GTPase Rac1 as a key downstream effector of OVOL that promotes proliferative growth. Moreover, we find that OVOL expression is directly activated by the Yorkie/Scalloped (Yki/Sd) complex, the core transcriptional effector of the Hippo pathway, without forming a feedback loop. This regulation is mediated through a specific Sd-binding motif (GATAA) within the OVOL promoter. Importantly, Yki/Sd-induced Rac1 expression is dependent on OVOL. Collectively, our findings establish the Yorkie/Sd-OVOL-Rac1 pathway that governs insect wing development by promoting cell proliferation, providing mechanistic insights into organ size regulation in animals.

Construction of a high-quality indexed EMS mutant library and a pipeline for identifying causal mutations in wheat by whole-exome sequencing
Dong Cao, Guangrui Liu, Yunlong Liang, Yaolin Yao, Haiqing Wang, Yun Li, Baolong Liu
, Available online  , doi: 10.1016/j.jgg.2026.06.009
Abstract (366) PDF (5)
Abstract:
CHST5 gene mutations contribute to high myopia by disrupting collagen fiber organization
Meining He, Senwei Tan, Furong Huang, Zhe Cheng, Lan Yao, Haibin Zhao, Kai Wu, Xiaolong Yu, Hongbing Jiang, Qiuping Liu, Xiangtian Zhou, Qi Tian, Zhengmao Hu, Kun Xia
, Available online  , doi: 10.1016/j.jgg.2026.06.007
Abstract (148) PDF (0)
Abstract:
High myopia (HM) is a leading cause of irreversible vision loss in working-age adults. Its pathogenesis is characterized by alterations in the microstructure and composition of collagen fibers, and genetic factors make a substantial contribution. In this study, we identify carbohydrate sulfotransferase 5 (CHST5) as a candidate gene for HM in humans and mice, with its mutations disrupting collagen fiber organization. The c.444C>A (p.S148R) variant in CHST5, a gene critical for sulfating corneal keratan sulfate (KS), completely co-segregates with HM in a Chinese family. Screening of CHST5 variants in 320 HM patients identifies two additional ones. We further find that Chst5 is expressed primarily in the cornea and sclera of mouse ocular tissues, and that the mutant protein CHST5S148R loses its Golgi localization. Homozygous mutant Chst5S126R mice exhibit HM phenotypes, including myopic refractive error (RE), significantly thinner sclera and cornea, notable microstructural changes in scleral and corneal collagen fibers, and shorter corneal KS chains. Our findings suggest that CHST5 NM_024533.5 c.444C>A (p.S148R) causes loss of proper protein localization, likely impairing its sulfotransferase function. This defect disrupts the organization of corneal and scleral collagen fibers and ultimately contributes to the development and progression of HM.
Neuroprotective role of Faecalibacterium prausnitzii-derived butyrate in diabetic gastrointestinal autonomic neuropathy
Lin Guo, Jie Li, Jiaming An, Jiahe Miao, Yang Yi, Keying Zhu, Qing Cai, Sijing Wang, Zhan Su, Xiaolong Ye, Yayun Wang, Minglin Pan, Qingmiao Lu, Bota Cui, Faming Zhang, Jianhua Mao, Xingyin Liu, Yibing Lu, Dafa Ding
, Available online  , doi: 10.1016/j.jgg.2026.06.005
Abstract (175) PDF (0)
Abstract:
Diabetic gastrointestinal autonomic neuropathy (DGAN) is a common yet poorly understood complication of diabetes that is characterized by gastrointestinal dysmotility and enteric neurodegeneration. Here, we investigate whether gut microbiota dysbiosis contributes to DGAN pathogenesis and explore the potential involvement of microbiota-derived metabolites in enteric nervous system (ENS) injury. Gut microbiota profiling reveals disease-associated compositional alterations in patients with DGAN, including depletion of Faecalibacterium-associated signals, with Faecalibacterium prausnitzii identified as a putative species-level annotation and negatively associated with gastrointestinal symptom severity. Fecal microbiota transplantation from patients with DGAN into db/db mice aggravates gut dysmotility and increases apoptosis of ChAT+ and nNOS+ myenteric neurons. Further experiments indicate that butyrate, the predominant metabolite produced by F. prausnitzii, attenuates neuronal apoptosis under high-glucose conditions. This biological process is accompanied by enhanced BCL2 expression, downregulation of cleaved caspase-3, and activation of the PI3K/Akt signaling cascade. Collectively, our findings support the presence of a gut microbiota-ENS axis in DGAN and identify butyrate as an important candidate neuroprotective metabolite associated with F. prausnitzii. These results provide a rationale for microbiota-targeted therapeutic strategies for diabetic enteric neuropathy.
Divergent roles of the Rag GTPases MoGtr1 and MoGtr2 in TOR signaling, autophagy, and pathogenicity of Magnaporthe oryzae
Shengjie Zheng, Tian Chen, Wei Cao, Shuaishuai Wang, Min Guo
, Available online  , doi: 10.1016/j.jgg.2026.06.002
Abstract (192) PDF (0)
Abstract:
In eukaryotes, conserved Rag-family GTPases are crucial for TOR signaling, integrating nutrient sensing with cellular growth. However, their functions in filamentous fungal pathogens are poorly understood. Here, we demonstrate that MoGtr1 and MoGtr2, the Rag GTPase homologs, form a functional heterodimer in Magnaporthe oryzae. Phenotypic analyses reveal that the △Mogtr1 mutant exhibits severe defects in autophagy, asexual reproduction, vacuolar homeostasis, and virulence. In contrast, the △Mogtr2 mutant is primarily impaired in autophagy. The △Mogtr1/2 double mutant shows additive defects in autophagy and mild attenuation in asexual reproduction and pathogenicity. Critically, MoGtr1 acts as an important GTP-dependent molecular switch. The GTP-bound form of MoGtr1 activates TOR signaling, maintains basal autophagy, and confers tolerance to hydrogen peroxide, whereas its GDP-bound form inhibits TOR, promotes autophagy, and increases sensitivity to hydrogen peroxide. Thus, MoGtr1 serves as a master regulator of development and infection, while MoGtr2 fine-tunes autophagy. Our study elucidates the divergent roles of Gtr homologs in a major fungal pathogen and proposes new strategies for disease intervention.
MaWRKY76 enhances symbiotic nodulation and salt tolerance of nodulation by activating NIN in Melilotus albus
Shengsheng Wang, Bao Ao, Qi Yan, Fan Wu, Pan Xu, Zhen Duan, Yunyue Xiao, Yangyang Han, Yuncan Qu, Jiyu Zhang
, Available online  , doi: 10.1016/j.jgg.2026.06.003
Abstract (160) PDF (0)
Abstract:
Legumes establish mutualistic symbiosis with rhizobia to facilitate biological nitrogen fixation. However, the genetic and molecular mechanisms of symbiotic nodulation regulated by salt stress remain largely unexplored. In this study, we identify that MaWRKY76 localizes to the nucleus and possesses transcriptional activation activity in Melilotus albus. MaWRKY76 expression is strongly induced under salt stress and upregulated during nodule formation. Transgenic hairy root composite plants overexpressing MaWRKY76 enhance salt tolerance by increasing antioxidant enzyme activity and alleviating oxidative damage. Furthermore, MaWRKY76 also promotes symbiotic nodulation by directly targeting the promoter region of NODULE INCEPTION (MaNIN), a master regulator of nodule development, and activating its expression. Under salt stress, MaWRKY76 overexpression plants enhance salt tolerance of nodulation by increasing nodule numbers and upregulating MaNIN expression, whereas knockdown of MaWRKY76 produces the opposite effect. Our findings reveal that MaWRKY76 functions as a transcriptional activator of NIN in legume nodulation signaling, and serves as a regulatory node to coordinate salt stress response with symbiotic nodulation.
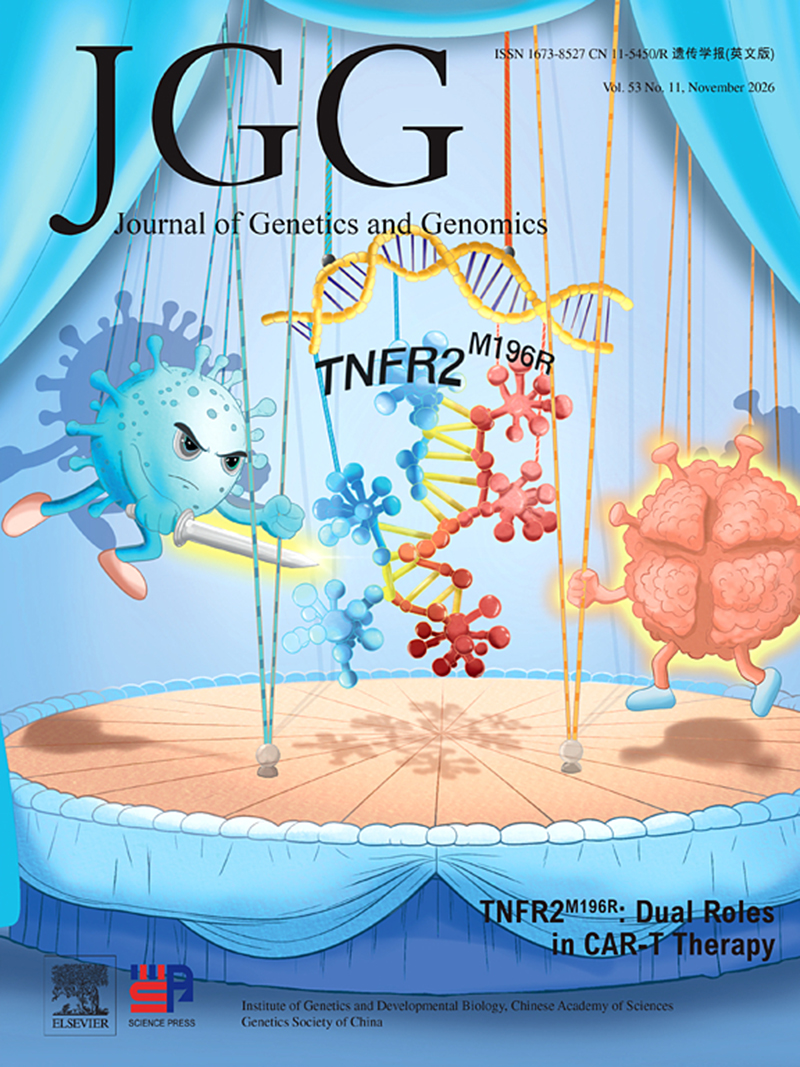
The TNFR2M196R germline variant confers dual impact on CAR-T cell immunotherapy by enhancing T-cell survival and tumor proliferation
Nan Liu, Lu Han, Yuting Lu, Yelei Guo, Jiajia Li, Qi Zhu, Yao Wang, Zhiqiang Wu, Xin Lin, Jianshu Wei, Weidong Han
, Available online  , doi: 10.1016/j.jgg.2026.05.014
Abstract (190) PDF (0)
Abstract:
Despite its remarkable efficacy in hematologic malignancies, chimeric antigen receptor T-cell (CAR-T) therapy is often limited by atypical clinical outcomes, necessitating predictive biomarkers. Here, prompted by a patient with low-tumor-burden non-Hodgkin lymphoma who develops severe and persistent cytokine release syndrome (CRS), we identify the germline TNFR2M196R single-nucleotide polymorphism, rs1061622, as a candidate genetic determinant of atypical responses. The TNFR2M196R variant is found to enhance the antitumor efficacy of CAR-T cells by reducing their apoptosis. Paradoxically, however, patients carrying the TNFR2M196R variant exhibit higher rates of early tumor progression. Further investigation reveals that while the TNFR2M196R mutation increases tumor susceptibility to CAR-T-mediated killing, it also accelerates tumor proliferation kinetics. Together, these results establish the TNFR2M196R SNP as a dual-function genetic determinant that modulates both CAR-T cell efficacy and intrinsic tumor behavior, highlighting the critical impact of germline genetics on cancer immunotherapy.
A high-quality genome assembly delineates the genetic basis of freezing tolerance in the elite wheat cultivar Jimai 22
Guang Qi, Guangwei Li, Rongzhi Zhang, Wei Li, Weilong Guo, Jun Zheng, Haosheng Li, Jianjun Liu, Takao Komatsuda, Cheng Liu
, Available online  , doi: 10.1016/j.jgg.2026.06.001
Abstract (693) PDF (2)
Abstract:
Mov10 mitigates hematopoietic stem cell exhaustion under stress by modulating the Camp-mediated inflammatory pathway
Changlin Zhen, Junbing Pan, Wei Wang, Jiankun Fan, Yonghui Zhang, Dan Liu, Shiheng Zhang, Xiaohuan Huang, Yalan Xiao, Yu Hou, Xueya Zhao, Zhigang Li
, Available online  , doi: 10.1016/j.jgg.2026.05.012
Abstract (124) PDF (0)
Abstract:
The integrity of hematopoietic stem cell (HSC) function is crucial for robust hematopoietic regeneration following stress. Inflammatory responses are pivotal drivers of HSC stress response, yet the precise modulation of inflammatory pathways remains incompletely defined. In this study, we identify the RNA helicase Mov10 as a negative regulator of stress-induced inflammatory pathways in HSC. Our study indicates that Mov10, which is critically required for HSC maintenance, is highly expressed in HSCs, and its loss adversely affects HSC fitness and survival during hematopoietic stress induced by bone marrow transplantation and irradiation (IR). Mechanistically, Mov10 mitigates excessive inflammatory activation to sustain HSC functional integrity during hematopoietic stress, primarily by enhancing the translation of CAMP, which inhibits the interaction between TNF-α and its receptor TNFR1 and suppresses NF-κB activation. Overall, our results imply that Mov10 plays a critical role in averting functional failure of hematopoiesis under stress, presenting viable paths for the therapeutic intervention of relevant diseases.
Multi-omics identification and verification of Dnajb14 as a modulator of retinal ganglion cell survival in glaucoma through ferroptosis
Yu Chen, Jie Wen, Qian Yang, Zixi Chen, Haibo Li, Zhenkai Wu, Quan Cheng, Dan Ji
, Available online  , doi: 10.1016/j.jgg.2026.05.011
Abstract (266) PDF (2)
Abstract:
Glaucoma is a leading cause of irreversible blindness and necessitates the identification of unreported therapeutic targets. Here, we report eight stable plasma protein targets for glaucoma, identified through a large-scale proteome-wide association study integrated with Mendelian randomization and colocalization analyses. Among these candidates, we show that the DnaJ heat shock protein family member B14 (DNAJB14) acts as a critical neuroprotective factor. Using single-cell and bulk transcriptomics, we demonstrate that Dnajb14 is characteristically expressed in retinal ganglion cells (RGCs) and that its abundance is inversely correlated with ferroptosis under ischemic stress. Furthermore, our results reveal that Dnajb14 preserve RGC survival and structural and functional integrity in multiple acute and chronic murine models. Mechanistically, Dnajb14 overexpression robustly alleviate ischemia-reperfusion-induced retinal injury, whereas targeted knockdown exacerbate lipid peroxidation, mitochondrial dysfunction, and ferroptosis. Taken together, our findings highlight that Dnajb14 is a key regulator of RGC survival through inhibition of ferroptosis, thereby making it a promising therapeutic candidate for glaucoma treatment.
Genetic dissection of cadmium accumulation in tea plants under nonphytotoxic field conditions
Shijie Luo, Xinwan Zhang, Weiyi Zhang, Yujia Ren, Dawei Gao, Xiali Ye, Alisdair R. Fernie, Weiwei Wen
, Available online  , doi: 10.1016/j.jgg.2026.05.006
Abstract (179) PDF (0)
Abstract:

Cadmium (Cd) accumulation in tea plants (Camellia sinensis) under chronic, nonphytotoxic field conditions poses a persistent food safety challenge; however, the genetic basis of this process remains uncharacterized. To dissect the genetic architecture of this trait, we quantify leaf Cd content and perform genome-wide association studies (GWAS) across 207 diverse tea accessions which have been cultivated for decades. Quantification analysis of Cd content reveals substantial natural variation in Cd content between young leaves (YL) and mature leaves (ML) under nonphytotoxic environment, with 38.2% of the 173 accessions with paired data intrinsically accumulating more Cd in YL than in ML. GWAS identify 112 quantitative trait loci associated with Cd accumulation. By further integrating the results of GWAS with Cd-responsive transcriptome profiling, we prioritize 14 high-confidence candidate genes. One of these candidate genes, CsHIPP6, which encodes a heavy metal-associated isoprenylated plant protein, is selected for further functional investigation. Heterologous expression in yeast and Arabidopsis demonstrates that CsHIPP6 enhances Cd tolerance and significantly reduces Cd accumulation. Our findings elucidate the genetic architecture underlying long-term Cd accumulation in perennial crops and highlight CsHIPP6 as a valuable target for breeding low-Cd tea cultivars to ensure beverage safety amidst rising environmental contamination.

The development and application of deep learning for stem cell research
Zhenfu Li, Wenjing Zhang, Yuxin Wu, Yaxuan Ye, Yuxin Chen, Jian Wu, Di Chen
, Available online  , doi: 10.1016/j.jgg.2026.05.008
Abstract (254) PDF (0)
Abstract:

Stem cells reside in specific microenvironments where they divide to maintain themselves and differentiate into functional cells that replace old, dead, or damaged cells to maintain tissue homeostasis. Some stem cells could also be cultured in vitro for self-renewal and directed differentiation towards specific lineages for both mechanistic interrogation and clinic investigation. A deeper understanding of the regulatory mechanisms underlying stem cell properties is essential for advancing their translational applications. Deep learning (DL), a branch of artificial intelligence, has been widely and deeply incorporated into different fields including biology. The application of DL in stem cells has revolutionized our research strategies and provided significant technical advantages. Here, we review the latest advances of the application of DL in analyzing bioimaging data and exploring large-scale genomics data in stem cells. We also summarize the limitations of current DL models and challenges of the application of these models in stem cell research.

Molecular basis of blade-pseudo-petiole-sheath axis patterning in Spodiopogon sagittifolius: roles of auxin signaling and lineage-specific regulators
Chao Xu, Yang Wang, Liping Qin, Renyi Ma, Wei Bi, Feng Liu, Zhenwen Liu
, Available online  , doi: 10.1016/j.jgg.2026.05.007
Abstract (189) PDF (0)
Abstract:
Metabolic reprogramming of sucrose flux boosts natural rubber in Taraxacum kok-saghyz
Xiuli Fan, Qingwen Chen, Xiaoguang Song, Liquan Kou, Guodong Wang, Jinfang Chu, Xia Cui, Hong Yu, Xia Xu, Jiayang Li
, Available online  , doi: 10.1016/j.jgg.2026.05.009
Abstract (691) PDF (3)
Abstract:
The transcription factor SlDOF9 directly activates SlELP5 expression to regulate inflorescence length in tomato
Junling Hu, Can Zhu, Xin Li, Lin Yang, Chen Zhang, Shumin He, Yonglian Wang, Yujie Zhang, Hui Zhang, Zejun Huang, Yanmei Guo, Xiaoxuan Wang, Yongchen Du, Lei Liu, Junming Li
, Available online  , doi: 10.1016/j.jgg.2026.05.002
Abstract (314) PDF (0)
Abstract:
Tomato inflorescence architecture is an important determinant of yield potential and fruit distribution, with inflorescence length and branching influencing resource allocation and harvestability. Although several regulators of inflorescence development have been identified, the genetic basis of natural variation in inflorescence length remains insufficiently characterized. Using a mapping population derived from LA1256 and Moneymaker, two lines with contrasting inflorescence lengths, we identify a quantitative trait locus, qIFL2.1, for inflorescence length on chromosome 2. Fine mapping and functional analyses identify SlELP5 as a strong candidate gene underlying qIFL2.1. Natural variation in the SlELP5 promoter is associated with altered SlELP5 expression, which is negatively correlated with inflorescence length. Consistently, SlELP5 overexpression in the LA1256 background reduces inflorescence length, supporting a negative role of SlELP5 in regulating this trait. Transcriptomic, dual-luciferase, and EMSA assays further show that SlDOF9 is upregulated in OE-SlELP5 lines and directly activates SlELP5 expression by binding to its promoter, supporting a SlDOF9–SlELP5 regulatory module. These findings provide insights into the molecular regulation of tomato inflorescence architecture and identify a potential target for precision breeding of inflorescence traits.
Population genomic analysis reveals genomic variations associated with domestication and key agronomic traits in bitter gourd (Momordica charantia)
Yu Niu, Rui Wang, Liang Wang, Aizheng Huang, Xu Han, Renbo Yu, Shijun Chen, Juntao Wang, Jiaowen Cheng, Changming Chen, Chengjie Chen, Yan Yang, Yi Liao, Ming Hu
, Available online  , doi: 10.1016/j.jgg.2026.04.017
Abstract (199) PDF (10)
Abstract:
Bitter gourd (Momordica charantia) is a widely cultivated vegetable and medicinal crop for its nutritional and pharmaceutical properties. Yet, the genomic basis of its domestication and key agronomic traits remains largely unexplored. Here, we present a high-quality, chromosome-level genome assembly specifically representing a smooth bitter gourd cultivar (Y1745), with a total assembly size of 327.14 Mb, a contig N50 of 11.33 Mb, and a scaffold N50 of 23.47 Mb. Comparative genomics reveals lineage-specific expansions of gene families related to stress responses and secondary metabolism. Population genomic analysis of 192 globally representative accessions identifies three distinct genetic groups (China, Southeast Asia, and South Asia Subcontinent), reflecting complex demographic histories, and uncovers strong signatures of selection associated with domestication and regional adaptation. In the genome-wide association study (GWAS) of 35 traits, we identify 893 significant trait-associated loci. Notably, two candidate genes (McWRKY and McFPF1-like 1) are associated with flowering time regulation, and one candidate gene (McEXLB1) with fruit size determination. Our work provides valuable genomic resources for understanding bitter gourd domestication and offers potential genetic targets for breeding.
Knockout of TpNRAMP5 reduces grain cadmium concentration in Polish wheat
Yiran Cheng, Xing Chen, Hongzhu Zhou, Jian Ma, Rui Wang, Yongjun Dong, Xi Li, Dan Long, Jian Zeng, Xing Fan, Houyang Kang, Dandan Wu, Yonghong Zhou, Ke Wang, Yi Wang
, Available online  , doi: 10.1016/j.jgg.2026.04.021
Abstract (552) PDF (4)
Abstract:
Erratum to ’Deep insights and clinical benefits from the comprehensive cohort of fetal skeletal dysplasia in China’
Guozhuang Li, Kexin Xu, Jihao Cai, Yulin Jiang, Xiya Zhou, Yan Lv, Na Hao, Yiqing Yu, Sen Zhao, Qing Li, Lina Zhao, Zhengye Zhao, Zhihong Wu, Ying Zou, Terry Jianguo Zhang, Shuyang Zhang, Nan Wu, Qingwei Qi
, Available online  , doi: 10.1016/j.jgg.2026.02.012
Abstract (152) PDF (0)
Abstract:
Fibroblast-directed melanocyte recruitment via Cxcl12–Cxcr4 axis promotes post-inflammatory hyperpigmentation and skin barrier protection in zebrafish
Shizheng Zhao, Si Li, Chenyuliang Zhang, Zheng Tang, Ao Zhang, Cong Huang, Kefan Cheng, Tao Yu, Yan Yan, Zilong Wen
, Available online  , doi: 10.1016/j.jgg.2026.02.017
Abstract (427) PDF (0)
Abstract:
Post-inflammatory hyperpigmentation (PIH) is a common skin disorder characterized by brown or black macules. It can be categorized as transient, typically resolving within 6–12 months, or permanent, persisting for years. While the pathogenesis of PIH is commonly linked to localized melanocyte overactivation, the precise cellular and molecular basis for this dysregulation, as well as its physiological significance, remains poorly defined. Using an acetic acid-induced zebrafish model, we identify melanocyte migration as a critical driver of hyperpigmentation. This process is independent of immune cells but driven by fibroblasts, which secrete Cxcl12a to recruit melanocytes via the Cxcl12a–Cxcr4a axis. Fibroblast ablation irreversibly disrupts melanocyte patterning, indicating that aberrant fibroblast activity dictates the permanence of PIH. The recruited melanocytes form a dual protective barrier against both UV-induced DNA damage and microbial intrusion. The translational relevance of this mechanism is underscored by upregulated CXCL12 expression in fibroblasts from human PIH-related conditions such as keloids, acne, and atopic dermatitis. Therapeutically, the FDA-approved CXCR4 antagonist AMD3100 (Plerixafor) effectively prevents and treats PIH in our model. Our findings elucidate a fibroblast-mediated mechanism of melanocyte recruitment in PIH, uncover previously unappreciated barrier functions of melanocytes in skin repair, and propose a promising repurposed treatment strategy.
The HPCA1–VDAC3 module mediates the humidity adaptation trade-off by interfering with ROS homeostasis in Arabidopsis thaliana
Songyi Yang, Luna Tan, Zhen Yan, Meng Liu, Yongqi Hu, Yuhan Zou, Feifei Huang, Xuerui Li, Jin Yan, Wensen Fu, Ruyun Liang, Si Tang, Mengyun Guo, Qiusai Wu, Songfeng He, Wenli Chang, Wei Liu, Yu Han, Yan Song, Xuemeng Gao, Yingjun Yao, Kai Yang, Weitao Jia, Feihua Wu, Jianquan Liu, Huanhuan Liu
, Available online  , doi: 10.1016/j.jgg.2026.02.013
Abstract (85) PDF (2)
Abstract:
Distinct Arabidopsis thaliana ecotypes differ in humidity adaptation, with Tibet ecotype adapted to arid habitats exhibiting lower submergence tolerance than humidity-adapted counterparts. Here, we identify a unique 332-bp transposable element (TE) insertion in the promoter of HYDROGEN-PEROXIDE-INDUCED Ca2+ INCREASES 1 (HPCA1) specific to the Tibet ecotype. This insertion upregulates HPCA1 expression by the recruitment of active histone modifications. HPCA1, in turn, negatively regulates submergence tolerance through its interaction with VOLTAGE-DEPENDENT ANION CHANNEL 3 (VDAC3), a voltage-dependent ionic channel localized in the mitochondrial outer membrane. Transcriptomic analyses indicate that the HPCA1–VDAC3 module modulates submergence tolerance, at least in part, by regulating RBOHD, WRKY46, and MYC2 to maintain reactive oxygen species (ROS) homeostasis. Additionally, HPCA1 facilitates VDAC3 phosphorylation and inhibits antioxidant enzyme activities, potentially disrupting ROS balance. A negative correlation between HPCA1 expression levels and precipitation is observed across global A. thaliana ecotypes. Together, our results suggest that the HPCA1–VDAC3 module integrates H2O2 signaling and ROS homeostasis via regulation of RBOHD, WRKY46, and MYC2, thereby mediating a trade-off in humidity adaptation in A. thaliana and providing insights for breeding flooding-tolerant crops.
ATP6V1C1 deficiency impairs auditory and vestibular hair cell function and leads to sensorineural hearing loss in humans and mice
Qingyue Yuan, Hao Zheng, Jun Xu, Yun Lin, Jingyu Liu, Hu Peng, Ningning Yuan, Xiaofei Zheng, Hao Wu, Xiuhong Pang, Tao Yang
, Available online  , doi: 10.1016/j.jgg.2026.01.005
Abstract (83) PDF (0)
Abstract:
Multiple genes encoding v-ATPase subunits are associated with various forms of syndromic hearing loss. Their functions in hair cells, the key sensory cells required for hearing and balance, remain unclear. In this study, linkage analysis and exome sequencing of a large autosomal dominant family with non-syndromic deafness identify a pathogenic p.R281P variant in ATP6V1C1 that encodes the C1 subunit of the v-ATPase. Conditional knock-out (CKO) of Atp6v1c1 in mouse hair cells results in early-onset sensorineural hearing loss and vestibular malfunction. The CKO mice show synaptic defects in inner hair cells, evidenced by decreased wave I amplitude and prolonged latency of the auditory brainstem responses, loss of inner hair cell ribbon synapses, accumulation of endocytic compartments, and absence of F-actin mesh network surrounding the active zones. The cochlear and vestibular hair cells of the CKO mice also undergo disrupted autophagic flux and apoptosis. The Atp6v1c1 p.R281P knock-in mice develop late-onset, high-frequency hearing loss with normal hair cell morphology but degenerated spiral ganglion neurons due to disrupted autophagic flux and apoptosis. Our study reveals ATP6V1C1 as a causative gene for non-syndromic deafness and its important roles in maintenance and synaptic function of hair cells.
Whole-genome sequencing reveals Yunnan as the crossroads of East and Southeast Asia for human gene flow
Xiaobo Qian, Bo Li, Jianmei Liu, Yushan Huang, Wenxi Gu, Yuwen Zhou, Qiong Nan, Chao Wang, Le Cheng, Junkun Niu, Fengrui Zhang, Qian Li, Xiuqing Zhang, Jinlong Yang, Yinglei Miao, Mingyan Fang, Xin Jin, Yang Sun
, Available online  , doi: 10.1016/j.jgg.2026.01.003
Abstract (301) PDF (21)
Abstract:
Yunnan Province has long served as a key nexus facilitating economic and cultural exchanges between East Asia, Southeast Asia, and Qinghai-Tibet Plateau. However, previous genetic studies were largely limited by sparse marker density, low sequencing depth, or single-population designs, leaving the population genetic structure and demographic history insufficiently resolved. Here, we conduct a high-resolution population genetic study based on 366 high-depth whole-genome sequencing samples from six ethnic groups, including Bai, Dai, Hani, Miao, Tibetan, and Han. We identify approximately 3.51 million novel variants and reveal fine-scale population structure and complex demographic histories among Yunnan ethnic groups. Beyond the three ancestries proposed by the tri-genealogy hypothesis, we detect a Han Chinese-related lineage, Yan-Huang, within multiple Yunnan populations. We further demonstrate that Yunnan represents a major gene-flow hotspot across East and Southeast Asia despite geographic barriers. Finally, we identify genomic loci under positive selection with candidate genes enriched in immune regulation, energy metabolism, cardiac development, and dietary adaptation, highlighting the role of local environmental pressures in shaping the genetic diversity of Yunnan populations.
Multidisciplinary exploration of ancient atherosclerosis: paleo-genomic and paleo-nutritional analysis of a 13th century artificial mummy in China
Bangyan Wang, Rui Wang, Duo Zheng, Baoshuai Zhang, Yu Shao, Jianxue Xiong, Panxin Du, Zixi Jiang, Lobsang Dargye, Edward Allen, Chenshuang Sun, Yiwen Shen, Bowen Gong, Pengfei Sheng, Liugen Lin, Jiucun Wang, Chuan-Chao Wang, Shaoqing Wen
, Available online  , doi: 10.1016/j.jgg.2026.01.004
Abstract (249) PDF (0)
Abstract:
The 13th-century Changzhou Mummy, from the Lower Yangtze region in China, is the earliest known East Asian case of an artificially mummified body employing mercury and cinnabar enema without evisceration. This study conducts multidisciplinary research, integrating paleo-radiological, paleo-pathological, paleo-genetic, and paleo-nutritional analysis to investigate the phenotype, genotype, individual life history, and the process of deliberate mummification performed on this individual. We generate a whole genome with 12.7× coverage, revealing potential genetic predisposition for several atherosclerotic cardiovascular diseases (ASCVD). Stable C and N isotope analysis of bones, teeth and hairs indicates high animal protein consumption as well as terminal illness. Hereditary and dietary risk factors are consistent with the diagnosis of atherosclerosis determined via postmortem examination. Our study, leveraging high-quality ancient DNA, provides a unique opportunity to challenge and rethink the widely accepted consensus on the relationship between atherosclerosis and post-industrial age lifestyles, uncovering unrecognized genetic polymorphisms of ASCVD among ancient individuals, and improving our understanding of the role of genetic factors in the development and evolution of ASCVD.
SYMPK interacts with KIF20A and NUMA1 to coordinate spindle organization and safeguard oocyte meiotic maturation
Bei Chen, Mofan Zhou, Jiaqi Wang, Jinxin Xiao, Yirong Chen, Jinying Wang, Wenlin He, Tianbao Song, Jin Luo, Qingzhen Xie, Cong Liu
, Available online  , doi: 10.1016/j.jgg.2026.01.002
Abstract (200) PDF (0)
Abstract:
Mammalian oocyte maturation relies on the precise assembly of the acentrosomal spindle, and its disruption causes aneuploidy and developmental failure. Symplekin (SYMPK), a 3′-end processing scaffold with emerging functions in regulating chromosome dynamics, remains unexplored in oocytes. Here, we investigate whether SYMPK governs spindle dynamics and chromosome fidelity during meiotic maturation. We find SYMPK dynamically tracks spindle microtubules during oocyte maturation following germinal vesicle breakdown (GVBD). By generating oocyte-specific Sympk knockout mice, loss of SYMPK in oocytes yields complete female infertility and impaired oocyte quality. Sympk-deficient oocytes show a predominant metaphase I (MI) arrest, accompanied by disorganized spindle architecture and destabilized kinetochore-microtubule attachments. Furthermore, chromosome spreads indicate persistent spindle assembly checkpoint (SAC) activation, and pharmacologic SAC inhibition can partially restore meiotic progression but not spindle integrity in SYMPK-deficient oocytes. Mechanistically, immunoprecipitation-mass spectrometry in MI oocytes reveals that SYMPK interacts with the spindle regulators KIF20A and NUMA1, and is required for their proper localization to the spindle. Collectively, these findings establish that SYMPK supports KIF20A and NUMA1 to coordinate acentrosomal spindle organization, thereby safeguarding oocyte meiotic maturation and ensuring faithful female meiotic progression.
Tiny genome with big impact: mitochondrial DNA in cardiovascular health
Yafang Yang, Jiaoyu Li, Lu Qian, Yuyan Xiong, Yi Yu
, Available online  , doi: 10.1016/j.jgg.2025.12.009
Abstract (102) PDF (0)
Abstract:
Cardiovascular diseases remain the leading cause of mortality worldwide. Mitochondrion, a key cellular organelle, harbors its own mitochondrial DNA (mtDNA) fundamental to cellular energy production through oxidative phosphorylation (OXPHOS). Beyond its canonical bioenergetic function, mtDNA integrity, copy number, and genetic variation play critical roles in maintaining cardiovascular function. This review provides a comprehensive overview of the multifaceted contributions of mtDNA to cardiovascular health and disease. We summarize the structural features and core biological functions of mtDNA, as well as the regulatory mechanisms governing its replication, biogenesis, and turnover. Particular emphasis is focused on mtDNA abnormalities, including point mutations, large-scale deletions, copy number alterations, and epigenetic modifications, and how these disturbances drive key pathogenic processes such as oxidative stress, chronic inflammation, apoptosis, and cellular senescence within the cardiovascular system. Furthermore, we highlight accumulating evidence linking mtDNA dysregulation to major cardiovascular disorders, including heart failure, atherosclerosis, and hypertension. Finally, we discuss the emerging diagnostic potential of circulating cell-free mtDNA and related mtDNA-derived metrics as non-invasive biomarkers, and outline therapeutic strategies aimed at preserving mtDNA integrity, modulating mtDNA content, or applying gene-based interventions to mitigate cardiovascular pathology.
m6A-RegBase enables regulatory investigation of m6A RNA modification in plants
Beilei Lei, Pengjun Ding, Minggui Song, Yuhang Ma, Tingrui Shi, Wen Sun, Jing Yang, Chengchao Jia, Chujun Zhang, Mengmeng Yuan, Zenglin Li, Chuang Ma
, Available online  , doi: 10.1016/j.jgg.2025.12.010
Abstract (291) PDF (0)
Abstract:
HNRNPH2 variant linked to intellectual disability disrupts myelination by impairing oligodendrocyte differentiation
Yang Jiao, Xingyu Pan, Jingrong Zhao, Xiaoyu Teng, Xiaoyi Liao, Xinyu Hu, Qiu Wang, Dandan Zheng, Yuxiang Pan, Xiaohui Deng, Xinyi Tan, Yun Stone Shi, Xu Zhang, Lan Bao, Bin Wang
, Available online  , doi: 10.1016/j.jgg.2025.12.006
Abstract (228) PDF (0)
Abstract:
Intellectual disability (ID) arises from complex pathogenic mechanisms. Although myelin dysfunction and white matter damage have been implicated, the cellular and molecular mechanisms linking impaired myelination to cognitive deficits remain largely unknown. Here, we identify a de novo heterogeneous nuclear ribonucleoprotein H2 (HNRNPH2) variant, c.638C>T (p.Pro213Leu), in patients with ID. The Hnrnph2P213L knock-in mice display spatial learning deficits, representing a partial phenotypic overlap with HNRNPH2-related neurodevelopmental disorder. Notably, Hnrnph2P213L mice exhibit significant myelination defects, primarily due to the impaired differentiation of oligodendrocyte progenitor cells. Furthermore, the myelin-enhancing drug benztropine rescues myelination, restores myelin-related gene expression, and ameliorates cognitive deficits, highlighting the role of hnRNPH2 P213L-induced myelin abnormalities in the pathogenesis of ID. Mechanistically, the P213L mutation disrupts the interaction between hnRNPH2 and its target transcripts, leading to the downregulation of myelination-related genes. Collectively, these findings reveal a critical mechanistic connection between myelin dysfunction and ID, thereby offering potential therapeutic insights for X-linked neurodevelopmental disorders.
Dissecting the functional roles of CTCF and CTCF-s in human embryonic stem cells
Hongxin Zhong, Yuli Lu, Jie Zhang, Gongcheng Hu, Guangjin Pan, Hongjie Yao
, Available online  , doi: 10.1016/j.jgg.2025.12.007
Abstract (77) PDF (0)
Abstract:
CTCF is a highly conserved zinc finger protein that plays critical roles in transcriptional regulation and three-dimensional (3D) genome organization. An alternative splice isoform of CTCF (CTCF-s), lacking the N-terminal domain and 2.5 zinc fingers, competes with CTCF for genomic occupancy and reduces CTCF-mediated chromatin interactions. However, the functional differences between CTCF and CTCF-s remain unclear. In this study, by using an auxin-inducible degron (AID2) system with doxycycline-inducible transgene expression, we systematically investigate the roles of CTCF and CTCF-s in human embryonic stem cells (hESCs). Acute degradation of endogenous CTCF and CTCF-s, followed by isoform-specific rescue, reveals that CTCF is essential for cell morphology and proliferation, whereas CTCF-s exerts much weaker effects. Genome-wide ChIP-seq and Hi-C analysis uncover distinct binding landscapes for CTCF and CTCF-s, as well as their differential contributions to chromatin conformation. Notably, our data indicate that CTCF-s, like CTCF, could either act as a boundary insulator or bind to gene promoters to modulate expression of a fraction of genes. Overall, our study reveals that CTCF is dominant in regulating chromatin boundary stability and gene regulation, while CTCF-s contributes to a lesser degree.
An elite haplotype of the glutamine synthetase gene TaGS1.1-6A is associated with increased nitrogen use- and yield-related traits in wheat
Yazhou Wang, Wan Teng, Mingyang Lei, Yanpeng Wang, Xiang Ouyang, Caixia Gao, Ruilian Jing, Junming Li, Yiping Tong
, Available online  , doi: 10.1016/j.jgg.2025.12.008
Abstract (106) PDF (0)
Abstract:
Glutamine synthetase (GS) plays a crucial role in nitrogen (N) assimilation. Identifying elite alleles of GS genes can facilitate the breeding of wheat (Triticum aestivum) varieties with improved N use efficiency (NUE). Here, meta-quantitative trait loci (QTL) analysis based on five bi-parental linkage mapping populations reveals that TaGS1.1-6A co-localizes with a meta-QTL for N use- and yield-related traits. The promoter region of TaGS1.1-6A contains a variation caused by a miniature inverted-repeat transposable element (MITE) insertion. The MITE insertion induces DNA hypermethylation in the adjacent regions, thereby repressing TaGS1.1-6A transcription. The haplotype TaGS1.1-6AHapII without the MITE insertion has been subjected to selection during wheat breeding, and is associated with increased photosynthetic N use efficiency, N utilization efficiency, spike grain number, and grain yield per plant when a BC3F4 population is grown under varying N supply levels. Conversely, CRISPR/Cas9-mediated mutation of TaGS1.1 shows reduction in these traits. Furthermore, we develop a breeding strategy to enhance wheat grain yield under different N supply conditions by pyramiding TaGS1.1-6AHapII and the leaf senescence-delaying haplotype of TaNAM-A1. These findings suggest that TaGS1.1-6A contributes to N use- and yield-related traits, and TaGS1.1-6AHapII holds significant value for breeding wheat with improved NUE and yield.
Metabolite-QTL analysis reveals genetic basis of phenolic acid metabolism in barley grains
Ruilian Song, Siyu Zhang, Yixiang Wang, Yuhan Liu, Likun Liu, Chen Zhang, Yanjun Li, Qifei Wang, Genlou Sun, Xifeng Ren
, Available online  , doi: 10.1016/j.jgg.2025.12.004
Abstract (249) PDF (0)
Abstract:
Phenolic acid metabolites have important physiological functions, and many genetic loci affecting phenolic acid metabolism traits have been identified in several crops. Although barley is one of the important cereal crops, research on the genetic basis of phenolic acid synthesis in barley grains remains limited. Here, we analyze the 39 phenolic acid metabolites detected in mature grains of barley double haploid (DH) population and further identify 154 metabolite quantitative trait loci (mQTLs) related to 36 phenolic acids using four mapping methods. Subsequently, we identify 12 candidate genes that affect the content of phenolic acid metabolites, and overexpression of one candidate gene, HvCOMT-1, in barley reveals its involvement in the synthesis of phenolic acids. Moreover, we show that the transcription factor HvMYB-1 regulate the expression of HvCOMT-1. Functional analysis in Arabidopsis shows that HvCOMT-1 increases stem diameter and lignin deposition. Further analysis reveals that the expression level of HvCOMT-1 is closely related to the barley lodging-related traits. Overall, our findings enhance the understanding of the genetic basis for phenolic acid variations in mature barley grains and provide valuable reference for genetic improvement of barley nutritional quality.
MEI4 variations drive female reproductive disorders via impaired oocyte abundance and developmental potential
Yiyang Wang, Yu Qi, Keyan Xu, Shuyan Tang, Luyi Tan, Bingying Xu, Ying Wang, Shuxian Zhang, Yang Zou, Yuan Gao, Chunmei Zhang, Xin Liang, Xue Jiao, Shidou Zhao, Han Zhao, Shixuan Wang, Yingying Qin, Ting Guo, Zi-Jiang Chen
, Available online  , doi: 10.1016/j.jgg.2025.12.005
Abstract (232) PDF (0)
Abstract:

Meiotic DNA double-strand break (DSB) formation is pivotal for oocyte development, regulating both ovarian reserve and oocyte developmental potential. Mutations in DSB formation genes have been associated with premature ovarian insufficiency (POI) and adverse pregnancy outcomes in women. Whole exome sequencing in 1530 POI patients across two Chinese cohorts identifies loss-of-function variants in the DSB formation gene, MEI4, enriched in POI. These MEI4 variants impair DSB formation in vitro and reveal a previously unrecognized function of the MEI4 C-terminus in stabilizing the MEI4–REC114 subcomplex on the chromosome axes. Additionally, Mei4Arg356*/Arg356* mice display severe defects in DSB formation, leading to massive apoptosis in oocytes triggered by the HORMAD1-dependent synapsis checkpoint in late prophase I. The few mutant oocytes surviving past the checkpoint exhibit low developmental potential, characterized by complete early embryonic arrest due to aneuploidy. Notably, heterozygous Mei4+/Arg356* mice show intermediate follicle depletion and embryonic development arrest consistent with the phenotype of heterozygous POI and preimplantation embryonic arrest, suggesting a haploinsufficiency effect. This study defines the impacts of MEI4 mutation on oocyte quantity and quality, which can guide genetic diagnosis and intervention in patients with POI and early embryonic arrest, especially those with mutations in meiotic DSB formation genes.

A non-canonical bipartite NLS dictates nuclear import and function of the Drosophila Pc protein
Tao He, Yu Fan, Yongchang Zeng, Jie Zhou, Renjie Jiao, Alan Jian Zhu, Min Liu
, Available online  , doi: 10.1016/j.jgg.2025.12.003
Abstract (400) PDF (0)
Abstract:
Improving rice false smut resistance by editing grain-filling-related genes
Jie Liu, Ya-Dan Zhao, Peng-Yu Chen, Ting Zhan, Lu-Hao Liu, Jie Zhao, Guo-Bang Li, Zhi-You Gong, Yong Zhu, He Wang, Ji-Wei Zhang, Yan Li, Wen-Ming Wang, Jing Fan
, Available online  , doi: 10.1016/j.jgg.2025.12.002
Abstract (232) PDF (0)
Abstract:
The roles of strigolactones in plant resilience to environmental stresses
Yanting Wang, Mohammad Golam Mostofa, Feifei Yi, Lam-Son Phan Tran, Weiqiang Li
, Available online  , doi: 10.1016/j.jgg.2025.12.001
Abstract (210) PDF (0)
Abstract:
Strigolactones (SLs) are a group of phytohormones that enhance hyphal branching of arbuscular mycorrhizal fungi (AMF), promote seed germination of parasitic plants, and influence plant growth, development, and stress acclimation. SLs improve plant stress resilience by modulating shoot and root architecture, photosynthesis, nutrient homeostasis, and antioxidant defense. Under nutrient deficiency, SL accumulation enhances AMF colonization and triggers the expression of genes related to the nutrient starvation response. When subjected to drought, SLs mitigate water loss by modulating stomatal density and closure, cell membrane integrity, and anthocyanin biosynthesis. SLs also mitigate salinity and heavy metal stresses by maintaining ion homeostasis, while conferring thermotolerance and enhancing chilling tolerance. In this review, we highlight recent advances in SLs and their mechanisms in plant responses to environmental stresses, including nutrient deficiencies, drought, salinity, extreme temperatures, metal toxicity, herbivore attack, and pathogen infection. We further discuss how SLs interact with other phytohormones to coordinate the physiological, biochemical, and molecular regulatory events critical for plant acclimation to abiotic and biotic stresses.
Roles of ethylene in plant growth, development, and stress responses
Xun Zhang, Jianjun Tao, Yihua Huang, Shouyi Chen, Cuicui Yin, Jinsong Zhang
, Available online  , doi: 10.1016/j.jgg.2025.11.015
Abstract (463) PDF (30)
Abstract:
Ethylene, a pivotal gaseous phytohormone, regulates diverse processes in plant growth, development, and stress adaptation. In Arabidopsis, ethylene perception by endoplasmic reticulum (ER)-localized receptors initiates a canonical signaling cascade involving CONSTITUTIVE TRIPLE RESPONSE 1 (AtCTR1) and ETHYLENE INSENSITIVE 2 (AtEIN2). This pathway culminates in nuclear translocation of AtEIN2-CEND and activation of the transcription factor AtEIN3/EIN3-LIKE1 (AtEIL1). Rice employs conserved (OsEIN2, OsCTR2, OsEIL1/2) and unique (Mao Huzi 3 [MHZ3], MHZ11, MHZ1) components for ethylene signaling, reflecting adaptations to semi-aquatic environments. Ethylene regulates developmental processes including seed germination, apical hook formation, root architecture, flowering, and senescence, often via intricate crosstalk with auxin, abscisic acid (ABA), jasmonic acid (JA), gibberellins (GA), and brassinosteroids (BR). Ethylene signaling also influences rice yield-related traits such as grain filling, grain size, and starch biosynthesis. Moreover, ethylene modulates responses to abiotic stresses (such as submergence, hypoxia, salinity, drought, and temperature fluctuations) and nutrient imbalances. This review synthesizes current understanding of ethylene signaling and its functions, focusing on the model dicot Arabidopsis thaliana and the monocot rice (Oryza sativa). It highlights conserved and diverged mechanisms, underscoring ethylene’s potential as a target for enhancing crop resilience and productivity in changing environments.
The quantitative trait locus stiff2 controls stalk bending strength and root architecture in maize
Shuyang Zhong, Le Xu, Zhihai Zhang, Yan Li, Zhongwei Lin
, Available online  , doi: 10.1016/j.jgg.2025.11.014
Abstract (385) PDF (0)
Abstract:
Stalk lodging is a major threat to global maize production, which causes great annual yield losses. Stalk bending strength (SBS) is highly associated with resistance to stalk lodging in maize. However, the genetic basis of SBS remains largely unknown. In this study, we identify a quantitative trait locus (QTL), stiff2, corresponding to a known flowering-time gene ZmCCT, by integrating QTL mapping and association mapping. A 5-kilobase (kb) transposable element inserted in the promoter region of ZmCCT significantly reduces SBS, while upregulated expression of ZmCCT through transformation significantly enhances SBS. Gene regulatory network analysis reveals that ZmCCT may indirectly regulate a set of downstream genes, which contain nrt5 for nitrogen transport, Tu1, d8, and d9 for stalk elongation, ub2, ub3, and ch1 for stalk thickening, and myb69 and bm4 for lignin biosynthesis. These genes collectively increase stalk strength and improve lodging resistance aboveground. Additionally, stiff2 regulates not only aboveground resistance to stalk bending and breakage but also influences root system architecture, which enhances resistance to root lodging belowground. The identification of stiff2 and its downstream targets provides critical insights into the genetic control of maize lodging and offers powerful tools for breeding lodging-resistant cultivars.
Identification of a major locus Fhb.Er-3StS from Elymus repens conferring Fusarium head blight resistance in wheat
Fei Wang, Hanyu Wei, Hongjing Xu, Qianyi Guo, Wei Zhu, Lili Xu, Yiran Cheng, Yazhou Zhang, Yi Wang, Jian Zeng, Xing Fan, Yonghong Zhou, Dandan Wu, Yinghui Li, Houyang Kang
, Available online  , doi: 10.1016/j.jgg.2025.11.013
Abstract (500) PDF (0)
Abstract:
CellReasoner: a reasoning-enhanced large language model for cell type annotation
Guangshuo Cao, Yi Shen, Lingyu Zhao, Jianghong Wu, Haoyu Chao, Ming Chen, Dijun Chen
, Available online  , doi: 10.1016/j.jgg.2025.11.012
Abstract (447) PDF (0)
Abstract:
The role of primary cilia in physiological and pathological states of the central nervous system
Denghui Zhai, Xuebin Zhou, Zhangqi Xu, Xiying Chen, Ya Li, Jiabin Li, Tianhua Zhou, Shanshan Xie
, Available online  , doi: 10.1016/j.jgg.2025.11.011
Abstract (188) PDF (0)
Abstract:
Primary cilia, microtubule-based organelles widely existing on eukaryotic cells, play a critical, indispensable role in the development and functional maintenance of the central nervous system (CNS). Here, we summarize recent advances on the distribution of primary cilia across different regions of the CNS, providing a detailed map highlighting their presence on both neurons and glial cells. Furthermore, we elaborate on the roles of primary cilia in essential physiological functions, including progenitor cell proliferation, neurogenesis, neuronal migration, and synaptic connections within the CNS. We also discuss the emerging links between ciliary dysfunction and a range of CNS disorders, including cognitive impairment, metabolic disorders and hyperphagia-induced obesity, neurodegenerative diseases, and psychiatric conditions. Integrating existing findings, this review provides a panoramic perspective on ciliary roles in the CNS and lays a critical foundation for subsequent basic research, disease-mechanism studies, and therapeutic-target exploration.
Advances in monogenic female infertility
Hao Gu, Lei Wang, Qing Sang
, Available online  , doi: 10.1016/j.jgg.2025.11.009
Abstract (225) PDF (0)
Abstract:
Complete genome assembly of the Xian rice variety IR64 as a valuable source in genomics and breeding research
Min Li, Tingting Sheng, Linjun Yu, Shuyue Zheng, Shijiao Li, Shuran Zhou, Fengcai Wu, Fan Zhang, Chaopu Zhang, Erbao liu, Yingyao Shi, Xue Mi, Xueru Zhang, Zhikang Li, Yanru Cui, Wensheng Wang
, Available online  , doi: 10.1016/j.jgg.2025.11.010
Abstract (521) PDF (0)
Abstract:
Intrinsic NPRL2 and NPRL3 regulate the sensitivity of B-cell malignancies to CAR-T cell therapy
Fuxin Han, Yuting Lu, Yipeng Zhang, Xinran Ma, Chuan Tong, Jianshu Wei, Yelei Guo, Chun Liu, Zhiqiang Wu, Weidong Han, Yao Wang
, Available online  , doi: 10.1016/j.jgg.2025.11.007
Abstract (296) PDF (9)
Abstract:
Although chimeric antigen receptor (CAR) T-cell therapy has markedly improved outcomes for many patients with B-cell malignancies, a subset experiences limited benefit due to primary or secondary resistance. Building on CRISPR/Cas9 genome-wide screening in malignant B-cells, we identify NPRL2 and NPRL3 as key regulators of tumor sensitivity to CAR-T cytotoxicity. This study aims to investigate the impact and mechanisms of tumor-intrinsic NPRL2 and NPRL3 on the efficacy of CAR-T cell therapy. In a tandem CD19/20 CAR-T clinical trial for relapsed/refractory (R/R) B-cell lymphoma (NCT03097770), high tumor NPRL2 or NPRL3 expression correlates with therapeutic resistance in patients. Consistently, in vitro experiments confirm that tumor cells overexpressing NPRL2/NPRL3 exhibit resistance to CAR-T–mediated cytolysis. Mechanistically, NPRL2/NPRL3 suppresses mTORC1 activity within tumor cells, negatively regulating the conjugation between tumor cells and CAR-T cells, consequently impairing CAR-T cell activation and cytotoxic function, ultimately facilitating immune escape. As therapeutic strategies, either genetic ablation of tumor-intrinsic NPRL2/NPRL3 or pharmacological activation of mTORC1 enhances CAR-T cell activation, cytotoxic degranulation, and tumor clearance both in vitro and in vivo. In conclusion, targeting tumor NPRL2/NPRL3 or directly activating mTOR represents a promising combinational strategy to potentiate CAR-T efficacy and overcome resistance in clinical practice.
Ribosomal protein bL31c interacts with translation elongation factor RAB8D to regulate chloroplast translation elongation and PSI-LHCI-LHCII assembly in Arabidopsis
Yukun Wang, Xiangsheng Ke, Shuaihao Chen, Jiabei Sun, Mi Zhou, La Bo, Hongkang Zhou, Qin Lu, Xin Hou
, Available online  , doi: 10.1016/j.jgg.2025.11.006
Abstract (296) PDF (9)
Abstract:
Chloroplast translation systems have evolved specialized regulatory mechanisms distinct from those of their prokaryotic ancestors. However, critical gaps persist in understanding how these systems coordinate translation efficiency with photosynthetic apparatus assembly—a process central to plant development. Here, we identify Arabidopsis BACTERIAL LARGE RIBOSOMAL SUBUNIT PROTEIN 31 (bL31c) as a critical chloroplast ribosomal protein that interacts with the translation elongation factor RAB GTPASE HOMOLOG 8D (RAB8D) to ensure translation elongation efficiency. Knocking down bL31c disrupts chloroplast translation, causing preferential depletion of photosystem I (PSI) subunits, a functional imbalance between PSI and PSII, and paradoxical accumulation of the PSI-LHCI-LHCII supercomplex. Comparative analysis reveals evolutionary conservation of the bL31c-EF-Tu functional module in Cyanobacteria but not in E. coli, demonstrating lineage-specific adaptation of translation surveillance mechanisms. Crucially, pharmacological inhibition of translation elongation in wild-type plants phenocopies the photosystem stoichiometry defects observed in bl31c mutants, establishing defective ribosome processivity as the primary driver of photosystem imbalance. Our findings uncover a plant-specific ribosomal checkpoint mechanism that dynamically coordinates protein synthesis with photosynthetic complex assembly, providing important insights into the evolutionary rewiring of organellar gene expression systems in eukaryotes.
hfCas12Max-mediated targeted integration at accessible chromatin regions with a goat-derived UCOE enhances stable recombinant lactoferrin expression
Zhenliang Zhu, Jing Han, Jianglin Zhao, Yangyi Jian, Yong Zhang, Yayi Liu, Peipei Bian, Zhenyu Wei, Xiaodan Ma, Rui Feng, Yuanpeng Gao, Jun Liu
, Available online  , doi: 10.1016/j.jgg.2025.11.008
Abstract (159) PDF (0)
Abstract:
Stable transgene expression in the mammary gland is crucial for recombinant protein production in livestock, yet it is frequently hampered by transgene silencing and random integration. To address this, we profile chromatin accessibility in goat mammary epithelial cells (GMECs) using ATAC-seq and identify 15 highly accessible genomic regions. Three of these regions are confirmed to support stable transgene expression. Notably, we identify a goat-derived ubiquitous chromatin opening element (UCOE) in the SF3B1-COQ10B intergenic region, with a high GC content (65%) and CpG island enrichment. This UCOE improves hfCas12Max-mediated integration of large DNA fragments and maintains high-level expression of human lactoferrin (hLTF) in GMECs. Subsequently, we precisely integrate the UCOE-hLTF cassette into the highly accessible loci and generate a transgenic goat via somatic cell nuclear transfer, without detectable off-target effects. Our pipeline, which integrates chromatin accessibility profiling, UCOE discovery, and precision editing, demonstrates the role of CpG island-containing UCOEs in preventing transgene silencing. The study provides valuable tools for enhancing recombinant protein production and supports the breeding of dairy goats for milk with high lactoferrin content, while advancing the understanding of the interactions between chromatin, regulatory elements, and transgenes in molecular breeding.
Genome-wide association study of HBV-related hepatocellular carcinoma identifies a functional variant at the FAM114A1 locus
Hong-Ping Yu, Bang-De Xiang, Ji Qian, Hongliang Liu, Yuan-Feng Li, Qiuling Lin, Shun Liu, Junjie Wei, Shicheng Zhan, Binbin Jiang, Juncheng Dai, Liang Ma, Litu Zhang, Yingchun Liu, Qiuping Wen, Wenfeng Gong, Shengping Li, Yanji Jiang, Ji Zheng, Tianyi Zhu, Zihan Zhou, Xiaoyun Zeng, Ziliang Wang, Ji-Ao Wang, Rui Guo, Yuan Yang, Qingyi Wei, Gangqiao Zhou, Xiao-Qiang Qiu, Weizhong Tang, Mengyun Wang, Ruoxin Zhang
, Available online  , doi: 10.1016/j.jgg.2025.11.003
Abstract (186) PDF (0)
Abstract:
Liver cancer ranks sixth in cancer incidence and third in cancer-related deaths worldwide. Hepatocellular carcinoma (HCC) is the primary histological subtype, and hepatitis B virus (HBV) carriers have a higher risk of HCC. Although several susceptibility loci for HCC have been identified in East Asian populations through genome-wide association studies (GWAS), the underlying biological mechanisms of this malignancy remain incompletely understood. Here, we conduct a two-stage GWAS including 2413 cases and 2794 HBV-positive controls from a high-incidence region in Southern China. The function of the susceptibility locus is investigated by bioinformatic and experimental approaches, supported by a xenograft model. We identify a 4p14 locus significantly associated with the risk of HCC (rs55718051, OR [95% CI] = 0.73 [0.67–0.80], Pmeta = 9.14 × 10⁻¹¹), and 18q23 locus with borderline significance (rs12964643: OR [95% CI] = 0.75 [0.67–0.83], Pmeta = 1.11 × 10⁻⁷). Functional experiments indicate the role of rs55718051 in FAM114A1 expression regulation, possibly through interaction of FOXA1. Knockdown of FAM114A1 significantly enhances the oncogenic phenotypes in liver cancer cells, suggesting its potential tumor suppressor role. Our findings expand the understanding of HCC susceptibility and suggest FAM114A1 as a potential suppressor in HBV-related HCC carcinogenesis.
Key divisions and cell specification during embryo pattern formation require SMU1-mediated splicing of CAK genes in Arabidopsis
Xiaoyi Huang, Yue Liu, Yajun Cai, Xiaogang Long, Emily Xu, Xiaoming Zhong, Jingyuan Zhang, Man Wang, Guoyong Xu, Peng Zhao, Kun Wang, Limin Pi, Arp Schnittger, Hongchun Yang
, Available online  , doi: 10.1016/j.jgg.2025.10.008
Abstract (313) PDF (0)
Abstract:
Embryonic pattern formation and cell specification require precise cell division and cell cycle regulation. Splicing factors and the splicing of precursor mRNA (pre-mRNA) play significant roles in embryo development. However, how splicing factors control embryonic patterning via RNA splicing remains unclear. Here, we show that the mutation of SUPPRESSORS OF MEC-8 AND UNC-52 1 (SMU1), a conserved subunit of the spliceosomal B complex, causes compromised cell fate of the hypophysis and quiescent center (QC), failed embryonic root apical meristem (RAM) formation, as evidenced by altered WUSCHEL-RELATED HOMEOBOX 5 (WOX5) expression and perturbed auxin signaling. This results in smu1 embryo lethality. The splicing efficiency of three out of four CYCLIN-DEPENDENT KINASE ACTIVATOR (CAK) genes is decreased, leading to reduced protein levels in smu1 embryos. These CAK genes are required for hypophysis specification and embryonic RAM formation. SMU1 binds CAK transcripts in vitro and in vivo. Restoring the expression of either CAK gene partially rescues the defects in smu1 embryos, leading to the formation of QC-like cells, continued embryo development, and even the production of viable seeds. Our data suggest that SMU1 binds to CAK transcripts and promotes their splicing, enabling cell cycle progression to promote embryonic RAM formation.
Transcriptomic landscape of Marchantia polymorpha sexual organs at single-nucleus resolution
Yuying Zeng, Yuqing Cai, Zhencheng Tu, Jingyi Liao, Xiayi Chen, Xing Guo, Sibo Wang, Linzhou Li, Yan Xu, Shanshan Dong, Huanming Yang, Tong Wei, Yang Liu
, Available online  , doi: 10.1016/j.jgg.2025.11.002
Abstract (313) PDF (1)
Abstract:

Marchantia polymorpha, a model liverwort, provides a valuable system for investigating the evolution of plant sexual reproduction. To explore the cellular landscape of its reproductive structures, we generate a single-nucleus transcriptomic atlas of the antheridiophore, archegoniophore, and sporophyte. Using single-nucleus RNA sequencing (snRNA-seq), we capture over 30,000 high-quality nuclei and identify distinct cell populations. In the male organ, we characterize stages of spermatogenesis from early antheridium cells to mature sperm, revealing dynamic transcriptional programs including cell cycle regulation, chromatin remodelling, and calcium signalling. In the female organ, we define cell types including archegonial layers and secondary central cells. Sporophyte clusters are annotated as spores, elaters, capsule wall, foot, and seta cells, with transcriptional signatures related to structural support, stress response, and reproductive functions. Cross-species analysis indicates that capsule wall cells in liverworts are similar to tapetum cells. Notably, foot cells exhibit high expression of genes involved in sporopollenin biosynthesis and signaling pathways, serving as a central hub that mediates communication between the maternal gametophyte and the developing sporophyte. This study provides a comprehensive cellular and molecular map of M. polymorpha reproductive organs and sporophyte, establishing a framework for investigating the development and evolution of sexual reproduction in early land plants.

Epigenetic regulation of cilia stability and kidney development by the chromatin remodeling SWI/SNF complex in zebrafish
Xiaoyu Cheng, Qianshu Zhu, Shilin Ma, Xiaoyu Peng, Guanliang Huang, Guifen Liu, Wentao Zhang, Yong Zhang, Cizhong Jiang, Andong Qiu, Ying Cao
, Available online  , doi: 10.1016/j.jgg.2025.11.001
Abstract (73) PDF (0)
Abstract:

Cilia are vital subcellular organelles whose assembly is regulated by master transcription factors, such as Foxj1 and Rfx. However, the mechanisms of epigenetic regulation over cilia stability remain largely unclear. Here, we investigate the epigenetic control by manipulating chromatin remodeling genes in zebrafish. We demonstrate that the depletion of multiple components of the switch/sucrose non-fermentable (SWI/SNF) chromatin remodeling complex induces ciliopathy-like phenotypes in zebrafish embryos. Specifically, the loss of Actl6a, an essential component of the SWI/SNF complex, leads to cilia disassembly and cystic kidney defects, without affecting cilia motility. Our multi-omics analyses (RNA-seq, ATAC-seq, and FitCUT&RUN) consistently reveal that in Actl6a-depleted pronephros or embryos, a critical set of cilia genes, including the master regulators foxj1a and rfx2, exhibit concordant downregulation across the transcriptional level, chromatin accessibility, and SWI/SNF binding. Consistently, the depletion of foxj1a or rfx2 causes cilia assembly defects and cystic kidney formation in zebrafish. Furthermore, overexpression of either foxj1a or rfx2 mRNA substantially rescues the cystic kidney and cilia disassembly defects observed in actl6a-/- mutant embryos. Collectively, these findings reveal that the SWI/SNF complex maintains cilia stability and kidney homeostasis by directly modulating the expression of the key ciliogenesis transcription factors foxj1a and rfx2.

ZmPRX38 is required for improving stalk strength and yield in maize
Xiaqing Wang, Tianyi Wang, Ruyang Zhang, Min Deng, Xuan Sun, Jinghuan Li, Dongmei Chen, Yanxin Zhao, Jidong Wang, Shuai Wang, Zhiyong Li, Ruibin Xu, Ronghuan Wang, Wei Song, Jiuran Zhao
, Available online  , doi: 10.1016/j.jgg.2025.10.009
Abstract (290) PDF (0)
Abstract:
Stalk lodging is a major problem in maize production, usually causing significant yield losses due to weak stalk strength. Understanding the genetic basis of stalk strength is crucial for improving maize lodging resistance. In this study, we identify 31 quantitative trait loci (QTLs) related to maize stalk strength and clone ZmPRX38 (encoding peroxidase 38) responsible for a hotspot QTL region of stalk strength. ZmPRX38 is highly expressed in maize stalk during vegetative growth stage, and its protein is localized in the cell membrane, cytoplasm and apoplast. Knockout of ZmPRX38 decreases stalk strength and yield in maize, while overexpressing ZmPRX38 increases stalk strength and yield. ZmPRX38 in phenylpropanoid pathway is involved in the biosynthesis of guaiacyl lignin, p-hydroxy-phenyl lignin, and syringyl lignin. Additionally, we identify a favorable haplotype of ZmPRX38, which enhances stalk strength, containing 3 loci distributed in the 5’ untranslated region (UTR), exon 1, and 3’UTR of ZmPRX38, respectively. Although 91.46% of maize natural lines contain this favorable haplotype, most of the Huang-gai (HG) lines, a backbone maize germplasm, contain the unfavorable haplotypes. Therefore, targeted improvement of ZmPRX38 by editing unfavorable haplotypes may be an effective strategy for increasing maize stalk strength, thereby improving maize lodging resistance and yield.
Dual-target CRISPR-Cas12 diagnostics based on asymmetrically chemical-modified DNA probe
Xinge Wang, Yangcan Chen, Yanping Hu, Shengqiu Luo, Siqi Wang, Bangwei Mao, Changxian Peng, Chongjian Chen, Weiye Pan, Haiyan Yan, Jianyou Liao, Qi Zhou, Wei Li
, Available online  , doi: 10.1016/j.jgg.2025.10.007
Abstract (59) PDF (1)
Abstract:
CRISPR-based nucleic acid detection technologies have revolutionized infectious disease detection and environmental monitoring by leveraging RNA–DNA complementarity to enable rapid, precise, and cost-effective detection of targets. However, achieving multitarget detection in one tube still presents challenges that necessitate further research. Here, we develop a nucleic acid detection module based on the CRISPR-Cas12i system. Importantly, we find that Cas12i and AapCas12b exhibit opposite trans-cleavage preferences for asymmetrically phosphorothioate-modified single-strand DNA probes, enabling the development of an effective dual-target nucleic acid detection platform by combining these two Cas12 nucleases in one tube. Moreover, this dual-target detection platform exhibits high specificity and sensitivity in genotyping the nucleic acid targets of human papillomavirus (HPV) 16 and HPV18, as well as Influenza A virus (FluA) and Respiratory syncytial virus. Notably, combined with loop-mediated isothermal amplification, this platform achieves high detection rates for clinical samples (18/18 FluA and 18/18 GAPDH internal reference detection rate). Taken together, these results can broaden the application of CRISPR-based Cas12 proteins for multi-target nucleic acid detection in one tube.
E75-induced Toll/NF-κB signaling cooperates with Notch and Hippo pathways to promote tumor malignancy
Xianping Wang, Yifan Guo, Chenglong Wang, Jingjie Mu, Xianjue Ma
, Available online  , doi: 10.1016/j.jgg.2025.10.006
Abstract (35) PDF (0)
Abstract:
A universal cost-efficient sample labeling approach for multiplexed single cell RNA-seq based on recombinant HUH-endonuclease-agglutinin tagging
Quanyong Zhang, Maorong Li, Luemou Shen, Lujia Chen, Zhiheng Yuan, Xu Zhang, Hao Wang, Yang Yu, Fucheng Luo, Guangdun Peng, Jingjie Hu, Zhenmin Bao, Ning Song, Kai Chen
, Available online  , doi: 10.1016/j.jgg.2025.10.004
Abstract (408) PDF (0)
Abstract:
Recent advances in single-cell transcriptomics have revolutionized our understanding of cellular diversity and tissue heterogeneity, providing unprecedented insights into biological and medical research. However, the high per-assay cost limits broader applications of this technology. Although sample-labeling strategies enabling multiplexing have emerged, current methods suffer from either impractical complexity for barcoding or high cost for preparing the index labeling reagents. To address these challenges, here we present HEATag, a universal cell membrane labeling approach that combines Duck circovirus HUH endonuclease (DCV) with wheat germ agglutinin (WGA) to efficiently tag cell membranes with indexed single-stranded DNA (indexed ssDNA). The DCV domain enables rapid, sequence-specific conjugation of indexed ssDNA, while the WGA domain ensures robust labeling of fresh or fixed cells across diverse species. This method is compatible with both commercial platforms and custom systems, readily adaptable to various single cell omics workflows. Therefore, HUH-endonuclease-agglutinin tagging (HEATag) provides a universal, cost-effective and scalable solution for high-throughput single-cell studies, enhancing library preparation efficiency and minimizing batch effects for single-cell researchers.
To see and to know: the power of live imaging in illuminating and decoding biological complexity
Miaoling Yang, Zhuo Du
, Available online  , doi: 10.1016/j.jgg.2025.10.003
Abstract (287) PDF (0)
Abstract:
Live imaging enables direct observation of dynamic biological processes, capturing their progression from molecular to organismal scales in space and time. Through high-resolution observation, it provides a powerful means to decode biological complexity by revealing dynamic behaviors, spatial patterns, and regulatory changes. This review illustrates the application of live imaging in investigating complex biological processes with spatiotemporal resolution and mechanistic insight. We first highlight the analytical power and integrative strategies of live imaging, and then summarize recent advances that further extend its capacities. We then focus on four complex processes—cell proliferation, lineage regulation, morphogenesis, and atlas construction—to elucidate how live imaging contributes to their decoding through representative studies. We also discuss the conceptual and practical limitations that currently constrain the full interpretive potential of live imaging, underscoring the need for deeper integration between observation, perturbation, and modeling. Looking ahead, live imaging will benefit from both technical refinement and advances in data standardization and visualization, functional quantification, multiscale integration, and the discovery of generalizable principles. Together, these directions advance a more integrative and mechanistic understanding of complex biological processes.
Dissecting the genetic basis of reproductive transition and reproductive growth in wheat by considering accumulated temperature
Liujie Jin, Kening Duo, Chao Fu, Yunzhen Li, Chao He, Xin Gong, Wenhao Yan
, Available online  , doi: 10.1016/j.jgg.2025.09.012
Abstract (164) PDF (0)
Abstract:
Maize Chlorotic Leaf Spot1 encodes a fumarylacetoacetate hydrolase essential for carbohydrate partitioning
Ruchang Ren, Sihang Zhao, Hong Jia, Hao Li, Lishuan Wu, Jinge Tian, Yifan Zhu, Junxiang Tang, Xiangyang Guo, Chenglong Wang, Feng Tian
, Available online  , doi: 10.1016/j.jgg.2025.10.002
Abstract (99) PDF (0)
Abstract:
Carbohydrate partitioning from photosynthetic sources to non-photosynthetic sinks is essential for plant development and crop yield. Using a maize-teosinte BC2S3 population, we identify Chlorotic Leaf Spot1 (CLS1), a fumarylacetoacetate hydrolase (FAH) in the tyrosine degradation pathway that plays an essential role in carbohydrate partitioning in maize. CLS1 localizes to the plasma membrane, cytoplasm, and nucleus. Allelic tests and sequence analysis reveal that the teosinte parent CIMMYT8759 carries a weak allele of CLS1, likely due to rare amino acid substitutions at residues 175 and 355. Loss-of-function mutants of CLS1 develop chlorotic leaf spots accompanied by carbohydrate hyperaccumulation, reduced photosynthetic efficiency, chloroplast damage, and impaired transient starch conversion. Critically, cls1 mutants exhibit ectopic callose accumulation and aberrant plasmodesmata ultrastructure at the mesophyll-bundle sheath and bundle sheath-vascular parenchyma interfaces. This defect causes starch granule and soluble sugar accumulation in chlorotic leaf tissues, indicating a disruption of the symplastic transport pathway. Collectively, our results uncover an important role for FAH in plant development and identify CLS1 as a key regulator of symplastic carbohydrate partitioning.
circRNAs derived from a nuclear hormone receptor act differentially on insect metamorphosis and reproduction
Lulu Gao, Qiang Yan, Ying Qu, Wanwan Li, Jiasheng Song, Shutang Zhou
, Available online  , doi: 10.1016/j.jgg.2025.10.001
Abstract (284) PDF (0)
Abstract:
Insects are the most diverse group on earth, partially owing to their metamorphosis and strong fecundity. Circular RNAs (circRNAs) are stable molecules implicated in a broad range of biological processes. However, the regulatory roles of circRNAs in insect metamorphosis and reproduction are unclear. Methoprene-tolerant (Met) is the nuclear receptor of juvenile hormone (JH) that plays dual roles of inhibiting precocious metamorphosis and promoting reproduction. Here, we report that locust Met generates two circRNAs, circMet1 and circMet2, respectively. While circMet1 is highly expressed in the cuticle of late final instar, circMet2 is more abundant in the corpora allata, brain, and fat body of early vitellogenic adults. Interestingly, circMet2 is generated by complementary pairing of Penelope-like remnants across the introns of Met. Moreover, circMet2 functions as a miRNA sponge of four species-specific miRNAs that downregulate Met translation. siRNA-mediated knockdown of circMet1 causes the delay of metamorphosis and retarded vitellogenesis. Loss of circMet2 results in significantly decreased vitellogenin synthesis, along with blocked ovarian growth. These results reveal the differential roles of circMet1 and circMet2 in modulating insect metamorphosis and female reproduction. This study advances our understanding of how circRNAs derived from a single gene exert distinct roles in insect life history.
Single-molecule chromatin profiling reveals cell type-specific A/B compartment alteration and multi-enhancer transcriptional coordination
Luo-Ran Liu, Jia-Yong Zhong, Xin Bai, Chen-Liang Ye, Chunhui Hou, Junjun Ding, Wei Chi, Chuan-Le Xiao, Longjian Niu
, Available online  , doi: 10.1016/j.jgg.2025.09.011
Abstract (69) PDF (0)
Abstract:
In eukaryotic organisms, the three-dimensional organization and epigenomic landscape of chromatin are fundamental to the regulation of gene expression. Previous studies have provided significant insights into CpG methylation, chromatin accessibility, and the dynamics of 3D architecture. However, a systematic delineation of how these epigenomic features regulate transcriptional activity remains limited. In this study, we develop nanoCAM-seq, a single-molecule sequencing technique designed to simultaneously profile higher-order chromatin interactions, chromatin accessibility, and endogenous CpG methylation. This approach provides an integrative view of chromatin features associated with cis-regulatory elements and reveals their coordinated dynamics during transitions of A/B compartments. Single-molecule analyses using nanoCAM-seq further reveal that promoters characterized by low CpG methylation and high chromatin accessibility more frequently interact with multiple enhancers. Collectively, our findings establish nanoCAM-seq as a powerful approach for resolving the coordinated dynamics of chromatin architecture and epigenetic modifications, offering critical insights into the regulatory mechanisms underlying gene expression.
Genomic insights into population structure, adaptation, and archaic introgression at the Himalayan–East Asian crossroads
Mengge Wang, Shuhan Duan, Qiuxia Sun, Yuntao Sun, Lintao Luo, Yunhui Liu, Renkuan Tang, Libing Yun, Chao Liu, Guanglin He
, Available online  , doi: 10.1016/j.jgg.2025.09.010
Abstract (64) PDF (2)
Abstract:
Tibetan-Yi Corridor (TYC) is a crucial agro-pastoral region in the eastern Himalayas, linking Qinghai‒Xizang Plateau with the lowlands of East Asia and facilitating human migration for millennia. However, genomic research on TYC populations remains limited, which limits the understanding of their origins and health. We provide genomic data from 1031 individuals belonging to Austroasiatic and Sino-Tibetan groups, including 147 whole-genome sequences from 13 underrepresented Tibeto-Burman and Austroasiatic communities. Our analysis reveals approximately 3.3 million new genetic variants and 4 distinct genetic backgrounds within TYC populations. Demographic reconstructions reveal strong genetic connections among Tibeto-Burman groups, Central Plain Sinitic populations, and Yangshao farmers, supporting a common origin for Sino-Tibetan speakers. We identify signatures of high-altitude adaptations typical of Tibetans and TYC-specific variants linked to pigmentation and hypoxia responses. Differentiation involves mechanisms such as HLA-DQB1, which are related to immune function. Several rare pathogenic variants, like CYP21A2 and PRX, are notably frequent. Variants influencing warfarin sensitivity show significant variation. Archaic human introgression further promotes genomic complexity, impacting cardiovascular and immune-related genes, which suggests adaptation through ancient human interactions. These findings refine the evolutionary history of TYC populations and underscore the need for broader genomic research to capture regional diversity and inform precision medicine.
Genetic innovations underlying the evolution of root nodule symbiosis in Leguminosae
Tengfei Liu, Hao Lin, Zhixi Tian
, Available online  , doi: 10.1016/j.jgg.2025.09.008
Abstract (44) PDF (0)
Abstract:
Root nodule symbiosis (RNS) is a mutualistic association formed between nitrogen-fixing rhizobia or Frankia and host plants limited to four orders within Rosid I—Fabales, Fagales, Cucurbitales and Rosales—which comprise the so-called ‘Nitrogen Fixing Nodulation Clade’ (NFNC). The majority of nodulation studies have focused on Leguminosae, given their agricultural and environmental importance, as well as the widespread occurrence of nodulation among members of this family. Endowing cereal crops with nitrogen fixation, like Leguminosae, presents a strategy to reduce the detrimental effects of synthetic fertilizer overuse. Different hypotheses on the origin of RNS have been proposed, however key genetic innovations underlying the evolution of RNS, even in Leguminsoae, have been rarely reported. In this review, we begin by examining current knowledge of genetic innovations—including gene gain, gene loss, and the acquisition or loss of conserved noncoding sequences (CNS) in preexisting genes. We explore the available evidence supporting these genetic innovations underlying the evolution of RNS in Leguminosae and offer the phylogenomics approach that could be applied to uncover these genetic innovations. Finally, we conclude by proposing a model of genetic innovations underlying the evolution of RNS in Leguminsoae and consider the potential implications for the development of nitrogen-fixing crops.
Application of an optimized non-invasive prenatal testing for thalassemia based on change of haplotype doses
Fei Sun, Yao Zhou, Xing Zhao, Qiuling Jie, Linna Ma, Dan Lin, Yaxuan Li, Yangqing Mai, Ge Gao, Yongfang Zhang, Qi Li, Yanlin Ma
, Available online  , doi: 10.1016/j.jgg.2025.09.007
Abstract (264) PDF (14)
Abstract:

Patients affected by monogenic diseases impose a substantial burden on both themselves and their families. The primary preventive measure, i.e., invasive prenatal diagnosis, carries a risk of miscarriage and cannot be performed early in pregnancy. Hence, there is a need for non-invasive prenatal testing (NIPT) for monogenic diseases. By utilizing enriched cell-free fetal DNA (cffDNA) from maternal plasma, we refine the NIPT method, which combines targeted region capture technology, haplotyping, and analysis of informative site frequency. We apply this method to 93 clinical families at genetic risk for thalassemia, encompassing various genetic variant types, to establish a workflow and evaluate its efficiency. Our approach requires only 3 ng of DNA input to generate 0.1 GB of informative target genomic data and leverages a minimum of 3% cffDNA. This method has a 98.16% success rate and 100% concordance with conventional invasive methods. Furthermore, we demonstrate the ability to analyze fetal genotypes as early as eight weeks of gestation. This study establishes an optimized NIPT method for the early detection of various thalassemia disorders during pregnancy. This technique demonstrates high accuracy and potential for clinical application in prenatal diagnosis.

The application and prospects of spatial omics technologies in clinical medical research and molecular diagnostics
Xiaofeng Wu, Weize Xu, Da Lin, Leqiang Sun, Lit-Hsin Loo, Jinxia Dai, Gang Cao
, Available online  , doi: 10.1016/j.jgg.2025.09.003
Abstract (406) PDF (0)
Abstract:
While conventional FISH and IHC methods struggle to decode complex tissue heterogeneity and comprehensive molecular diagnosis due to low-throughput spatial information, spatial omics technologies enable high-throughput molecular mapping across tissue microenvironments. These technologies are emerging as transformative tools in molecular diagnostics and medical research. By integrating histopathological morphology with spatial multi-omics profiling (genome, transcriptome, epigenome, and proteome), spatial omics technologies open an avenue for understanding disease progression, therapeutic resistance mechanisms, and precise diagnosis. It particularly enhances tumor microenvironment analysis by mapping immune cell distributions and functional states, which may greatly facilitate tumor molecular subtyping, prognostic assessment, and predicting the efficacy of radiotherapy and chemotherapy. Despite the substantial advancements in spatial omics, the translation of spatial omics into clinical applications remains challenging due to robustness, efficacy, clinical validation, and cost constraints. In this review, we will summarize the current progress and prospects of spatial omics technologies, particularly in medical research and diagnostic applications.
Genome of aerial alga Trentepohlia odorata reveals insights into the evolution of terrestrial lifestyle in green algae
Yuanhao Li, Jiao Fang, Xiaoya Ma, Hao Wang, Shiyu Zhang, Yu Peng, Lin Chen, Guoxiang Liu, Huan Zhu, Zhenhua Zhang, Bojian Zhong
, Available online  , doi: 10.1016/j.jgg.2025.09.004
Abstract (72) PDF (0)
Abstract:
Trentepohliales is a completely terrestrial order within Ulvophyceae (the core Chlorophyta), and its closely related lineages are mainly marine macroalgae (green seaweeds). Despite the considerable interest in their biotechnological potential, little is known about their adaptations to challenging terrestrial habitats. Here, we assemble the high-quality reference genome of Trentepohlia odorata. This alga shows duplications of key genes associated with lipid metabolism and carotenoid synthesis, potentially facilitating intracellular accumulation of lipid droplets and carotenoids. We further reveal positive selection and expansion of gene families involved in vesicle trafficking and cell division regulation in T. odorata compared to other algae (cleavage furrow-mediated cell division) in Ulvophyceae, providing a genetic foundation for the evolution of phragmoplast-mediated cell division. The combined C4-like and biophysical carbon-concentrating mechanisms (CCMs) of T. odorata enable adaptation to fluctuating CO2 environments, and support efficient photosynthesis under CO2-limited conditions. Adaptive strategies of T. odorata to terrestrial stressors, such as drought, intense light, and UV-B radiation, include horizontally acquired genes involved in cell wall synthesis and remodeling, homeostasis of aldehydes, and expanded genes associated with reactive oxygen species (ROS), DNA repair, and photoprotection. Our study provides a valuable genomic resource for studying aerial algae and improves understanding of plant terrestrialization.
Genomic sequencing as a key primary recommendation for neonatal hyperbilirubinemia: a population-based multicenter study
Dabin Huang, Xia Gu, Weizhong Li, Hongying Mi, Haiquan Zeng, Guiying Zhuang, Sitao Li, Congcong Shi, Tao Wei, Wei Zhou, Xin Xiao, Wenhao Zhou, Hu Hao
, Available online  , doi: 10.1016/j.jgg.2025.09.002
Abstract (292) PDF (0)
Abstract:
Genetic variations are risk factors for neonatal hyperbilirubinemia (NHB), a common cause of infant hospitalization in the first postnatal week, but their contribution and long-term impacts remain unclear. This population-based multicenter study enrolls 1,780 hospitalized NHB newborns and 38,158 genetically screened newborns across 20 hospitals (2019–2022). Excluding cases with clear clinical causes, 977 NHB cases are categorized into genetic variation-positive and -negative groups. Results show significantly higher NHB-related gene variants (81.63% vs. 65.62%) and positive variation rates (36.29% vs. 9.4%) in NHB cases than in the general newborn population (all P < 0.001). Among the 977 NHB cases, 325 (33.3%) have positive variants, with higher rates of severe hyperbilirubinemia (16.9% vs. 9.7%, P = 0.001), prolonged jaundice (36.3% vs. 27.6%, P = 0.005), and cholestasis/hypercholanaemia (23.7% vs. 14.7%, P < 0.001) in the positive group. Cumulative genetic variants in bilirubin metabolism pathways exhibit dose-dependent associations with increased risks of complications. Long-term follow-up reveals that UGT1A1 variants prolong jaundice up to 1 month, while severe SLC10A1 variants cause persistent cholestasis/hypercholanaemia beyond 9 months. This large-scale evidence highlights genetic factors as key NHB determinants, with implications for neonatal care protocols to integrate genetic testing and establish long-term surveillance for variant carriers.
Integrative multi-omics and genomic prediction reveal genetic basis of early salt tolerance in alfalfa
Fei He, Ming Xu, Ruicai Long, Kai Zhu, Mengrui Du, Wenqi Ma, Hui Xue, Yanling Peng, Lin Chen, Junmei Kang, Yongfeng Zhou, Qingchuan Yang, Fan Zhang
, Available online  , doi: 10.1016/j.jgg.2025.09.001
Abstract (408) PDF (0)
Abstract:

The genetic basis of early-stage salt tolerance in alfalfa (Medicago sativa L.), a key factor limiting its productivity, remains poorly understand. To dissect this complex trait, we integrate genome-wide association study (GWAS) and transcriptomics (RNA-seq) from 176 accessions within a machine learning based genomic prediction framework. Analysis reveals weak genetic correlations among four salt-tolerance traits and a gradual decline in performance under increasing salt stress. GWAS identify 60 significant associated SNPs, with the highest number detected under 100 mM salt stress. Salt tolerance exhibits an additive effect from favorable haplotypes, which are most abundant in Chinese accessions. GWAS-associated genes are related to key regulators of hormone signaling and osmotic adjustment, while transcriptome analysis indicates a global repression of stress-responsive transcription factors. Integrating these multi-omics datasets allow us to identify 14 candidate genes, including MsHSD1 (seed dormancy) and MsMTATP6 (energy metabolism). Crucially, incorporating these markers into genomic prediction models improve cross-population predictive accuracy to an average of 54.4%. This study provides insights into the genetic architecture of salt tolerance in alfalfa and offers valuable markers to facilitate molecular breeding.

ATPase-deficient CHD7 disease variant disrupts neural development via chromatin dysregulation
Guangfu Wang, Zhuxi Huang, Chenxi He, Ze Wang, Shuhua Dong, Ming Zhu, Fei Lan, Wenhao Zhou, Weijun Feng
, Available online  , doi: 10.1016/j.jgg.2025.08.012
Abstract (241) PDF (0)
Abstract:
Chromodomain helicase DNA binding protein 7 (CHD7), an ATP-dependent chromatin remodeler, plays versatile roles in neurodevelopment. However, the functional significance of its ATPase/nucleosome remodeling activity remains incompletely understood. Here, we generate genetically engineered mouse embryonic stem cell lines harboring either an inducible Chd7 knockout or an ATPase-deficient missense variant identified in individuals with CHD7-related disorders. Through in vitro neural induction and differentiation assays combined with mouse brain analyses, we demonstrate that CHD7 enzymatic activity is indispensable for gene regulation and neurite development. Mechanistic studies integrating transcriptomic and epigenomic profiling reveal that CHD7 enzymatic activity is essential for establishing a permissive chromatin landscape at target genes, marked by the open chromatin architecture and active histone modifications. Collectively, our findings underscore the pivotal role of CHD7 enzymatic activity in neurodevelopment and provide critical insights into the pathogenic mechanisms of CHD7 missense variants in human disease.
Deciphering the genetic regulation of flowering time in rapeseed for early-maturation breeding
Minghao Zhang, Wei Chang, Ruicheng Hu, Yuxuan Ruan, Xiaodong Li, Yonghai Fan, Boyu Meng, Shengting Li, Mingchao Qian, Yuling Chen, Yuanyi Mao, Daifei Song, Haikun Yang, Luxiang Niu, Guangyuan Cao, Zhixia Deng, Zhixuan Qin, Hui Wang, Kun Lu
, Available online  , doi: 10.1016/j.jgg.2025.08.011
Abstract (352) PDF (0)
Abstract:
Flowering time is a critical agronomic trait with a profound effect on the productivity and adaptability of rapeseed (Brassica napus L.). Strategically advancing flowering time can reduce the risk of yield losses due to extreme climatic conditions and facilitate the cultivation of subsequent crops on the same land, thereby enhancing overall agricultural efficiency. In this review, we synthesize current information on flowering time regulation in rapeseed through an integrated analysis of its genetic, hormonal, and environmental dimensions, emphasizing their crosstalk and implications for yield. We consolidate multi-omics evidence from population genetics, functional genomics, and systems biology to create a haplotype-based framework that overcomes the trade-off between flowering time and yield, providing support for the precision breeding of early-maturing cultivars. The insights presented here could inform future research on flowering time regulation and guide strategies for increasing rapeseed productivity.
Estimating genetic load from 5000 Chinese exomes
Xiaoyue Du, Xiaoxi Zhang, Jiucun Wang, Li Jin, Shuhua Xu
, Available online  , doi: 10.1016/j.jgg.2025.08.009
Abstract (347) PDF (1)
Abstract:
Recent advancements in genome sequencing have enabled the estimation of genetic load through deleterious mutation profiling. However, Chinese populations remain underexplored in this context. We analyze whole-exome sequencing data from 5002 individuals, encompassing major Han subgroups—North Han (N-Han), South Han (S-Han), and Guangxi Han (G-Han)—as well as 13 ethnic minorities. Notably, G-Han exhibits significant genetic affinity with the Zhuang population. Systematic curation of 2110 ClinVar pathogenic or likely pathogenic variants reveals 93.4% are ultra-rare. Exceptions include GJB2 rs72474224-A (hearing loss), which shows higher frequencies in Zhuang and G-Han, and β-thalassemia-associated HBB variants (rs33986703-A and rs33950507-T), which are elevated in G-Han compared to other Han subgroups. Among 96 autosomal dominant mutation carriers, LDLR variants are predominant (∼25%), with comparable frequencies across Han subgroups. Adaptive signatures highlight gene-environment interactions: MTHFR rs1801133-A (UV adaptation) declines southward, while ALDH2 rs671-A (alcohol metabolism) displayed the opposite trend. ABCC11 rs17822931-A, associated with tropical adaptation, is particularly prevalent in G-Han. Gene-based rare-variant collapsing analyses identify an elevated risk of retinitis pigmentosa in S-Han (PRPF4, TUB). Our findings demonstrate that genetic load in Chinese populations is influenced by demographic history, population structure, and regional adaptation, emphasizing the importance of population-specific frameworks in precision medicine.
Rapamycin alleviates neurodegeneration in a Drosophila model of spinocerebellar ataxia type 51
Cuijie Wei, Taoyun Ji, Jin Xu, Yilei Zheng, Fuze Zheng, Suxia Wang, Chao Gao, Yalan Wan, Zhenyu Li, Jianwen Deng, Hui Xiong
, Available online  , doi: 10.1016/j.jgg.2025.08.010
Abstract (41) PDF (0)
Abstract:
Spinocerebellar ataxia (SCA) type 51 is a neurodegenerative disease caused by CAG repeat expansions in exon 1 of the THAP11 gene. These repeats are translated into a glutamine-rich protein, THAP11-polyQ, which forms protein aggregates and exhibits toxicity in cell models; however, the underlying mechanism remains unclear. In this study, we generate transgenic Drosophila models expressing varying lengths of THAP11-polyQ using the UAS-GAL4 system and assess neurodegeneration through pathological and behavioral analyses. Our results demonstrate that expression of THAP11-polyQ in transgenic flies leads to progressive neuronal cell loss, locomotor deficiency, and reduced survival. RNA sequencing of patient-derived skin fibroblasts reveals significant enrichment of the PI3K–Akt–mTOR pathway, and electron microscopy of transgenic flies shows an increase in multilamellar bodies, suggesting involvement of autophagy in SCA51. Consequently, we treat the fly model with rapamycin, an mTOR inhibitor known to enhance autophagy. This treatment reduces toxic THAP11-polyQ protein aggregates, significantly alleviates neuronal degeneration, and improves locomotor function, consistent with the rescue effects observed upon overexpression of Atg8a. Overall, these findings suggest that the Drosophila model, which recapitulates the neurodegenerative features of SCA51, can be used to investigate pathogenic mechanisms, and that rapamycin holds promising potential as a therapeutic approach for this disease.
MitoQ alleviates m.3243A>G-induced mitochondrial dysfunction by stabilizing PINK1 and enhancing mitophagy
Baige Cao, Lei Fang, Yinan Zhang, Chuwen Lin, Peng Liu, Huina Zhang, Orion Fan, Ming Xu, Zhao Qin, Congrong Wang
, Available online  , doi: 10.1016/j.jgg.2025.08.007
Abstract (98) PDF (0)
Abstract:
The mitochondrial 3243A>G mutation (m.3243A>G) is associated with diverse clinical phenotypes. To elucidate the underlying mechanisms and explore intervention strategies in m.3243A>G patients, urine-derived stem cells (USCs) and a mitochondrial leucyl-tRNA synthetase (lars-2) deficient Caenorhabditis elegans (C. elegans) model are used to assess mitochondrial homeostasis and neuromuscular dysfunction. Patient-derived USCs with high levels of m.3243A>G heteroplasmy exhibit impaired mitochondrial function, disrupted mitochondrial dynamics, and inhibited mitophagy, which are reversed by MitoQ through suppression of OMA1 zinc metallopeptidase (OMA1)-induced mitochondrial phosphatase and tensin (PTEN) induced kinase 1 (PINK1) degradation. Furthermore, lars-2 knockdown in C. elegans induces mitochondrial stress and mimics the loss of neural and muscle functions observed in patients with the m.3243A>G mutation. MitoQ treatment partially improves neurobehavioral function by promoting the PINK1 pathway. These findings suggest that MitoQ has therapeutic potential in the context of the m.3243A>G mutation.
Assembly of a high-quality reference genome and characterization of a chemical-mutagenized library of an elite soybean cultivar Tianlong 1
Yinghua Sheng, Yicheng Huang, Zilun Jin, Xuyan Wang, Chenghui Liu, Jingwen Zhang, Zhipeng Zhou, Chuang Ma, Jianwei Zhang, Min Chen
, Available online  , doi: 10.1016/j.jgg.2025.08.006
Abstract (125) PDF (1)
Abstract:
Soybean (Glycine max L.) is a globally vital crop for oil production and food security. High-quality genomic resources are instrumental for both functional genomics and breeding. Here, we report a near-complete, high-quality genome assembly of the elite cultivar Tianlong 1 (TL1), featuring fully resolved telomeres and centromeres, as well as a gap-free assembly of 14 of its 20 chromosomes. On the basis of the genome assembly, we generate an ethyl methanesulfonate (EMS)-mutagenized population comprising 2,555 M7 plants. Whole-genome re-sequencing of 288 EMS mutants uncovers 1,163,869 high-confidence single-nucleotide polymorphisms (SNPs) and 542,709 insertions/deletions (InDels), achieving 91.89% coverage of predicted protein-coding genes. Phenotypic screening demonstrates robust genotype–phenotype associations, with two nonsynonymous mutants displaying pronounced defects in seed and leaf development. Collectively, the chromosome-scale TL1 genome assembly and the extensively characterized mutant population establish valuable resources for functional genomics and precision breeding in soybean and related legume species.
Recapitulation of key phenotypes and pathological features of primary familial brain calcification (PFBC) in a mouse Slc20a2S602W/S602W model
Junyu Luo, Man Jiang, Jun Liu, Xuan Xu, Jing-Yu Liu
, Available online  , doi: 10.1016/j.jgg.2025.08.005
Abstract (353) PDF (2)
Abstract:
Gene traffic mediated by transposable elements shaped the dynamic evolution of ancient sex chromosomes of varanid lizard
Zexian Zhu, Jason Dobry, Erik Wapstra, Qi Zhou, Tariq Ezaz
, Available online  , doi: 10.1016/j.jgg.2025.08.002
Abstract (116) PDF (0)
Abstract:
Lizards usually exhibit frequent turnovers and a much greater diversity of sex determination mechanisms compared to birds and mammals, with the conserved ZW sex chromosomes of anguimorph lizards originating over 115 million years ago a seeming exception. We previously discovered in an anguimorph lizard Varanus acanthurus (Vac) whose entire chrW, but not chrZ is homologous to part of the chr2 by cytogenetic mapping, suggesting its complex history of sex chromosome evolution yet to be elucidated. To address this, we assemble a chromosome-level genome, and provide evidence that the Vac sex chromosome pair has undergone at least two times of recombination loss, producing a pattern of evolutionary strata like that of birds and mammals. Comparison to other lizard genomes date the stepwise propagation of specific retrotransposon subfamilies enriched near the duplicated gene pairs on the chrW and chr2 to the varanid ancestor. These retrotransposons probably have mediated the recruitment and amplification of autosomal genes on the chrW, including members of a large vomeronasal chemosensory receptor gene family V2R. Our results suggest that the W or Y chromosome as a refugium of repetitive elements, may recurrently recruit short-lived functional genes responsible for sexual dimorphisms during its long-term course of degeneration.
Spatiotemporal dynamics of neuron differentiation and migration in the developing human spinal cord
Yuan Yu, Mengjie Pan, Quanyou Cai, Ziyu Feng, Baomei Cai, Kaixuan Lin, Shangtao Cao, Mingwei Min, Lihui Lin, Yanlin Ma, Jiekai Chen
, Available online  , doi: 10.1016/j.jgg.2025.08.004
Abstract (250) PDF (0)
Abstract:
Precise formation of complex neural circuits in the spinal cord, achieved through the integration of diverse neuronal populations, is essential for central nervous system function. However, the specialization and migration of human spinal cord neurons remain poorly understood. In this study, we perform single-cell transcriptome sequencing of human spinal cord from Carnegie Stage (CS) 16-21 and mouse spinal cord from embryonic day (E) 8.0-11.5, complemented by in situ sequencing of human spinal cord (CS 16-20). Our results reveal the critical role of the precursor state in neuronal differentiation and migration, identifying key transcription factors that regulate these processes across species. Notably, each neuronal lineage expresses unique markers as early as the progenitor stage at the spinal cord midline, and subsequently undergoes a shared transcriptional program during precursor commitment that guides migration. This synchronized migration, validated by spatial transcriptomics, occurs in both dorsal and ventral regions. Our findings offer important insights into the migration patterns and regulatory factors that guide spinal cord neuron subtype specification during embryogenesis.
A telomere-to-telomere genome assembly of radish (Raphanus sativus L.) provides insights into QTL mapping of bolting traits
Feng Yang, Sihan Peng, Shuai Yuan, Maolin Ran, Xiaomei Li, Yuejian Li, Bin Liu, Ming Li, Chuibao Kong, Xiao Yang, Guohui Pan, Xiaoping Yong, Ke Ran, Na Kuang, Dawei Zhang, Honghui Lin
, Available online  , doi: 10.1016/j.jgg.2025.07.014
Abstract (429) PDF (1)
Abstract:
Radish (Raphanus sativus L.) is an important cruciferous root vegetable, with bolting regulated by multiple genes; however, the genetic mechanisms underlying bolting regulation remain unclear. Here, the genome of the cultivar C60213 is assembled into a high-quality, gap-free telomere-to-telomere structure, spanning nine chromosomes and totaling 472.71 Mb, using a combination of Oxford Nanopore, PacBio, and Hi-C sequencing technologies. It identifies 49,768 protein-coding genes, 97.38% of which are functionally annotated. Repetitive sequences constitute 59.72% of the genome, primarily comprising long terminal repeats. A high-density genetic linkage map is constructed using an F2 population derived from a cross between early- and late-bolting radishes, identifying seven major quantitative trait loci associated with bolting and flowering. RNA-seq and quantitative real-time PCR analysis reveal that the RsMIPS3 gene is found to be associated with bolting, with its expression decreasing during this process. Notably, RsMIPS3 overexpression in Arabidopsis delays bolting, confirming its role in regulating bolting time. These findings advance radish genome research and provide a valuable target for breeding late-bolting varieties.
Chromosome-level genome assembly of finger millet (Eleusine coracana) provides insights into drought resistance
Jiguang Li, Chaonan Guan, Xiaobo Li, Yanlan Wang, Jiayue He, Liqin Hu, Yaliang Shi, Yuqi He, Kaixuan Zhang, Rui Tang, Meiliang Zhou
, Available online  , doi: 10.1016/j.jgg.2025.07.012
Abstract (231) PDF (1)
Abstract:
Finger millet (Eleusine coracana Gaertn.), a nutritionally rich and drought-resilient C4 cereal, possesses exceptional grain storage longevity (up to 50 years). Here, we report a high-quality genome assembly of the allotetraploid cultivar C142, revealing extensive structural rearrangements between its two subgenomes (subA and subB), which are associated with asymmetric gene expression and subgenome dominance favoring subA. SubB diverged from its presumed progenitor E. indica approximately 6.8 million years ago, and two whole-genome duplication events shaped the current genome architecture, contributing to gene redundancy and adaptive potential. Notably, expansion of stress-related gene families, such as aldo-keto reductases, suggests a role in oxidative stress response and drought adaptation. Using genome-wide association studies (GWAS), we identify several candidate genes associated with key agronomic traits. Among them, EcMDHAR, encoding monodehydroascorbate reductase, plays a critical role in enhancing drought tolerance. Different EcMDHAR haplotypes exhibit distinct expression profiles, supporting their functional relevance in drought adaptation. This genomic resource not only advances our understanding of polyploid genome evolution in millets but also provides a foundation for genome-assisted improvement of drought resistance and nutritional quality in finger millet.
Massively parallel characterization of non-coding de novo mutations in autism spectrum disorder
Congcong Chen, Songwei Guo, Yanan Shi, Xinyu Gu, Ziye Xu, Yingjia Chen, Yayun Gu, Na Qin, Yue Jiang, Juncheng Dai, Yuanlin He, Xiao Han, Yan Liu, Zhibin Hu, Xiaoyan Ke, Cheng Wang
, Available online  , doi: 10.1016/j.jgg.2025.07.008
Abstract (293) PDF (0)
Abstract:
Autism spectrum disorder (ASD) is a neurodevelopmental disorder where de novo mutations play a significant role. Although coding mutations in ASD have been extensively characterized, the impact of non-coding de novo mutations (ncDNMs) remains less understood. Here, we integrate cortex cell-specific cis-regulatory element annotations, a deep learning-based variant prediction model, and massively parallel reporter assays to systematically evaluate the functional impact of 227,878 ncDNMs from Simons Simplex Collection (SSC) and Autism Speaks MSSNG resource (MSSNG) cohorts. Our analysis identifies 238 ncDNMs with confirmed functional regulatory effects, including 137 down-regulated regulatory mutations (DrMuts) and 101 up-regulated regulatory mutations (UrMuts). Subsequent association analyses reveal that only DrMuts regulating loss-of-function (LoF) intolerant genes rather than other ncDNMs are significantly associated with the risk of ASD (Odds ratio = 4.34; P = 0.001). A total of 42 potential ASD-risk DrMuts across 41 candidate ASD-susceptibility genes are identified, including 12 recognized and 29 unreported genes. Interestingly, these noncoding disruptive mutations tend to be observed in genes extremely intolerant to LoF mutations. Our study introduces an optimized approach for elucidating the functional roles of ncDNMs, thereby expanding the spectrum of pathogenic variants and deepening our understanding of the complex molecular mechanisms underlying ASD.
Universal single-copy ortholog benchmark gene set for bryophytes
Xuping Zhou, Tao Peng, Jin Yu, Shanshan Dong, Yang Liu
, Available online  , doi: 10.1016/j.jgg.2025.07.009
Abstract (533) PDF (0)
Abstract:
Functional characterization of OsLT9 in regulating rice leaf thickness
Jian Wang, Dagang Chen, Haifei Hu, Yamei Ma, Tifeng Yang, Jie Guo, Ke Chen, Chanjuan Ye, Juan Liu, Xinqiao Zhou, Chuanguang Liu, Junliang Zhao
, Available online  , doi: 10.1016/j.jgg.2025.07.010
Abstract (74) PDF (0)
Abstract:
Leaf thickness in rice critically influences photosynthetic efficiency and yield, yet its genetic basis remains poorly understood, with few functional genes previously characterized. In this study, we employ a pangenome-wide association study (Pan-GWAS) on 302 diverse rice accessions from southern China, identifying 49 quantitative trait loci (QTLs) associated with leaf thickness. The most significant locus, qLT9, is fine-mapped to a 79 kb region on chromosome 9. Transcriptomic and genomic sequence analyses identify LOC_Os09g33480, which encodes a protein belonging to Multiple Organellar RNA Editing Factor (MORF) family, as the key candidate gene. Overexpression and complementation transgenic experiments confirm LOC_Os09g33480 (OsLT9) as the functional gene underlying qLT9, demonstrating a 24-bp Indel in its promoter correlates with the expression levels and leaf thickness. Notably, OsLT9 overexpression lines show not only thicker leaf, but also significantly enhanced photosynthetic efficiency and grain yield, establishing a link between leaf thickness modulation and yield enhancement. Population genomic analyses indicate strong selection for OsLT9 during domestication and breeding, with modern cultivars favoring thick leaf haplotype of OsLT9. This study establishes OsLT9 as a key regulator controlling leaf thickness in rice, and provides a valuable genetic resource for molecular breeding of high-yielding rice through optimization of plant architecture.
Postnatal critical-period brain plasticity and neurodevelopmental disorders: revisited circuit mechanisms
Ziwei Shang, Xiaohui Zhang
, Available online  , doi: 10.1016/j.jgg.2025.07.006
Abstract (267) PDF (0)
Abstract:
Critical periods (CPs) are defined as postnatal developmental windows during which brain circuits exhibit heightened sensitivity to altered experiences or sensory inputs, particularly during brain development in humans and animals. During the CP, experience-induced refinements of neural connections are crucial for establishing adaptive and mature brain functions, and aberrant CPs are often accompanied by many neurodevelopmental disorders (NDDs), including autism spectrum disorders and schizophrenia. Understanding neural mechanisms underlying the CP regulation is key to delineating the etiology of NDDs caused by abnormal postnatal neurodevelopment. Recent evidence from studies using innovative experimental tools has continuously revisited the inhibition-gating theory of CP to systematically elucidate the differential roles of distinct inhibitory circuits. Here, we provide a comprehensive review of classical experimental findings and emerging inhibitory-circuit regulation mechanisms of the CP, and further discuss how aberrant CP plasticity is associated with NDDs.
Receptor-like protein 51 regulates brassinosteroid signaling by promoting the abundances of BRI1 and BAK1
Yuan Fang, Pengcheng Li, Bingqing Tao, Yujun Wu, Beibei Liu, Chengbin Xiao, Jia Li, Kai He
, Available online  , doi: 10.1016/j.jgg.2025.07.005
Abstract (353) PDF (0)
Abstract:
Brassinosteroids (BRs) are essential phytohormones that broadly regulate plant growth, development, and adaptation to biotic and abiotic stresses. In Arabidopsis, apoplastic BR molecules are perceived by a plasma membrane-localized receptor complex comprising the ligand-binding receptor BRI1 and the co-receptor BAK1. While negative regulators of the BR receptor complex, such as BKI1, BIR3, and PUB12/13, have been well characterized, how BRI1 and BAK1 are positively modulated in the BR pathway remains largely unknown. In this study, a genetic screen involving overexpression of RLP genes in the bak1-3 bkk1-1 double mutant reveals that enhanced RLP51 expression partially suppresses the BR-deficient phenotypes of bak1-3 bkk1-1. RLP51 overexpression also partially rescues the weak bri1 mutant allele, bri1-301. Although the rlp51 single mutant exhibits wild-type-like phenotypes, it enhances BR-defective phenotypes in bri1-301 and bak1 serk1 mutants. RLP51 is next found to interact with both BRI1 and BAK1 without affecting BRI1-BAK1 interaction. Critically, co-expression of RLP51 with BRI1 or BAK1 significantly increases BRI1 and BAK1 protein abundances. RLP51 appears to promote protein synthesis rather than stabilize BRI1 and BAK1 proteins. Thus, our study identifies RLP51 as a positive regulator of BR signaling that enhances the protein levels of BRI1 and BAK1.
Whole-genome sequencing provides insights into the evolutionary adaptation and conservation of gibbons
Wang Guiqiang, Wu Yajiang, Wang Song, Jia Ting, Yang Peng, Xu Zhongshi, Niu Wenhui, Shan Fen, Wang Chen, Chen Wu, Sun Ting
, Available online  , doi: 10.1016/j.jgg.2025.07.004
Abstract (98) PDF (0)
Abstract:
Gibbons are small, arboreal apes that play a critical role in tropical biodiversity and ecosystem ecology. However, nearly all species of gibbons are threatened by habitat loss, illegal trade, hunting, and other human activities. Long-term poor understanding of their genetics and evolution undermines effective conservation efforts. In this study, we analyse comparative population genomic data of four Nomascus species. Our results reveal strong genetic differentiation among these species as well as gene flow among Nomascus species. Additionally, we identify genomic features that are potentially related to natural selection linked to vocalization, fructose metabolism, motor balance, and body size, consistent with the unique phenotype and adaptability of gibbons. Inbreeding coupled with population declines due to climate change and historical human activities leads to reduced genetic diversity and the accumulation of deleterious variation that likely affects cardiovascular disease and the reproductive potential of gibbons and further reduces their fitness, highlighting the urgent need for effective conservation strategies.
Genetic landscape of hereditary cardiomyopathies and arrhythmias in China
Yang Lu, Zeyuan Wang, Shuyuan Zhang, Yaping Liu, Ye Jin, Zhuang Tian, Shuyang Zhang
, Available online  , doi: 10.1016/j.jgg.2025.07.003
Abstract (234) PDF (0)
Abstract:

Hereditary cardiomyopathies and arrhythmias are major contributors to cardiovascular morbidity and mortality. The advent of next-generation sequencing (NGS) has made genetic testing more accessible, which is crucial for precise diagnosis and targeted therapeutic strategies. The aim of this study is to explore the landscape of genetic variants, the relationship between specific variants and clinical phenotypes, and the impact on clinical decision-making in China. A total of 1,536 probands (median age, 37 years; 1,025 males [66.7%]) with suspected hereditary cardiomyopathy or arrhythmia (covering 15 clinical phenotypes) are recruited from 146 hospitals across 30 provinces and cities in China. Positive results are confirmed in 390 of 1536 probands, leading to a diagnostic yield of 25.4%. Forty-two and three tenths percent (n = 169) of family members carry the same variants as positive probands. Hypertrophic cardiomyopathy (HCM) and dilated cardiomyopathy (DCM) are the predominant phenotypes, with MYBPC3 variants having the highest frequency in HCM and TTN variants in DCM. In 76.9% of the positive probands, the identified variants are helpful in clinical management, family screening and fertility. This large-scale study provides significant insights into the genetic landscape of hereditary cardiomyopathies and arrhythmias in China.

Coiled-coil domain-containing 38 is required for acrosome biogenesis and fibrous sheath assembly in mice
Yaling Wang, Xueying Huang, Guoying Sun, Jingwen Chen, Bangguo Wu, Jiahui Luo, Shuyan Tang, Peng Dai, Feng Zhang, Jinsong Li, Lingbo Wang
, Available online  , doi: 10.1016/j.jgg.2023.09.002
Abstract (330) PDF (18)
Abstract:
During spermiogenesis, haploid spermatids undergo dramatic morphological changes to form slender sperm flagella and cap-like acrosomes, which are required for successful fertilization. Severe deformities in flagella cause a male infertility syndrome, multiple morphological abnormalities of the flagella (MMAF), while acrosomal hypoplasia in some cases leads to sub-optimal embryonic developmental potential. However, evidence regarding the occurrence of acrosomal hypoplasia in MMAF is limited. Here, we report the generation of base-edited mice knocked out for coiled-coil domain-containing 38 (Ccdc38) via inducing a nonsense mutation and find that the males are infertile. The Ccdc38-KO sperm display acrosomal hypoplasia and typical MMAF phenotypes. We find that the acrosomal membrane is loosely anchored to the nucleus and fibrous sheaths are disorganized in Ccdc38-KO sperm. Further analyses reveal that Ccdc38 knockout causes a decreased level of TEKT3, a protein associated with acrosome biogenesis, in testes and an aberrant distribution of TEKT3 on sperm. We finally show that intracytoplasmic sperm injection overcomes Ccdc38-related infertility. Our study thus reveals a previously unknown role for CCDC38 in acrosome biogenesis and provides additional evidence for the occurrence of acrosomal hypoplasia in MMAF.
PHYTOCHROME-INTERACTING FACTOR3 (PIF3) orthologs orchestrate stem elongation and wood formation in Populus
Xingyue Xiao, Hongli Cheng, Jianghai Mo, Fan Sun, Qin Song, Chengshan Zhang, Pan Yang, Keming Luo, Hongbin Wei
, Available online  , doi: 10.1016/j.jgg.2025.07.002
Abstract (328) PDF (0)
Abstract:

Phytochrome-interacting factors (PIFs) have been established as negative regulators of vascular patterning and xylem differentiation in the herbaceous plant Arabidopsis thaliana, however, PIFs’ regulatory role in secondary growth in woody species remains unclear. Here, we examined the expression patterns and involvement of PtoPIF3.1 and PtoPIF3.2 during stem growth and secondary xylem development in Populus tomentosa. Overexpression of either PtoPIF3.1 or PtoPIF3.2 significantly enhances both longitudinal stem growth and radial wood development. Conversely, CRISPR-generated Ptopif3.1 and Ptopif3.2 mutants exhibit reciprocal phenotypic defects. Exogenous auxin application partially restores the phenotypes of Ptopif3.1 and Ptopif3.2 mutants, and the auxin biosynthesis-deficient mutant Ptoyuc8 exhibits developmental abnormalities similar to those observed in Ptopif3 mutants. Further analysis revealed that PtoPIF3s directly bind to and activate expression of PtoYUC8 and cell expansion-related genes PtoEXPA1.1/1.2, while modulating cambial division and secondary xylem development marker genes (PtoWOX4, PtoANT, PtoCYCD3s, and PtoHB7/8) through auxin-mediated signaling. Together, our findings establish PtoPIF3.1/3.2 as key regulators that coordinate stem elongation and secondary growth in Populus, highlighting the functional divergence of PIF homologs between herbaceous and woody species.

PenCards: a global and community-contributed public archive of variant penetrance
Zhaopo Zhu, Ling Shang, Chuhan Shao, Zheng Wang, Xinxin Mao, Yuanfeng Huang, Pei Yu, Bin Li, Jinchen Li, Guihu Zhao
, Available online  , doi: 10.1016/j.jgg.2025.07.001
Abstract (96) PDF (0)
Abstract:

Penetrance is a crucial indicator for accurately assessing disease risk and plays a vital role in disease research, gene therapy, and genetic counseling. However, with penetrance data dispersed across various sources, efficiently accessing and consolidating this information becomes a challenge. A comprehensive platform that integrates penetrance is urgently needed. Here, we present PenCards, a global, community-contributed public archive of variant penetrance, by first collecting penetrance data from all published literature and then using large international cohorts to specifically calculate the penetrance of autism-related variants. PenCards contains a total of 244,531 variants—including 239,244 single nucleotide variants, 4,994 insertions and deletions, and 293 copy number variants, covering approximately 300 phenotypes. We also provide a submission portal for the dynamic updating of penetrance. Additionally, to help users efficiently access genetic information, we comprehensively integrate over 150 variant- and gene-level resources. In summary, PenCards is a powerful platform designed to advance genetic research and diagnostics. PenCards is publicly available at https://genemed.tech/pencards/.

USP21 deubiquitinates DPYSL2 and enhances its centrosomal abundance to promote cilium formation
Ting Song, Peng Zhou, Fengguo Zhang, Chunli Liu, Xueqing Han, Yiyang Yue, Mingzheng Hu, Shaodong Yan, Qingchao Li, Min Liu, Jun Zhou, Huijie Zhao
, Available online  , doi: 10.1016/j.jgg.2025.06.006
Abstract (57) PDF (0)
Abstract:

Cilia are microtubule-based organelles projecting from the cell surface with important sensory and motility functions. Ciliary defects are associated with diverse diseases collectively known as ciliopathies. However, the molecular mechanisms that govern ciliogenesis remain not fully understood. Herein, we demonstrate that ubiquitin-specific protease 21 (USP21) is indispensable for cilium formation through its deubiquitinating activity. Usp21 knockout mice exhibit ciliary defects in multiple organs, such as the kidney, liver, and trachea. Our data also reveal a constant localization of USP21 at the centrosome and basal body during ciliogenesis. Mechanistically, USP21 interacts with dihydropyrimidinase-like 2 (DYPSL2) at the centrosome and removes lysine 48-linked ubiquitination from DYPSL2. Loss of USP21 leads to the proteasomal degradation of DPYSL2 and causes a significant reduction in its centrosome abundance, ultimately resulting in ciliary defects. These findings thus identify a critical role for the USP21–DPYSL2 axis in ciliogenesis and have important implications for health and disease.

The auxin response factor OsARF12 modulates rice leaf angle via affecting shoot gravitropism
Fengjun Xian, Shuya Liu, Bin Xie, Jishuai Huang, Qiannan Zhang, Yimeng Xu, Xinrong Zhang, Chen Lv, Lin Zhu, Jun Hu
, Available online  , doi: 10.1016/j.jgg.2025.06.005
Abstract (369) PDF (3)
Abstract:

Leaf angle is a pivotal agronomic trait that significantly influences crop architecture and yield. Plant hormones, such as auxin, play a critical role in regulating leaf angle; however, the underlying molecular mechanisms remain to be fully elucidated. Here, we reveal that the auxin response factor gene, OsARF12, which is highly expressed in the leaf lamina joint, negatively regulates rice (Oryza sativa) leaf angle via affecting shoot gravitropism. Overexpression lines of OsARF12 exhibit more erect leaf angles, while the osarf12 mutants display enlarged leaf angles compared to the wild type. Further studies demonstrate that OsARF12 directly activates the expression of Loose Plant Architecture1 (LPA1) and LAZY1 by binding to their promoters. The osarf12 mutant presents impaired shoot gravitropism, a phenotype consistent with that of the lpa1 or lazy1 mutant. Collectively, we elucidate the biological functions of OsARF12, which modulates leaf angle through its impact on shoot gravitropism by regulating the expression levels of LPA1 and LAZY1. This study provides insight into the role of auxin in determining rice leaf angle, potentially holding profound effects for the optimization of crop architecture.

Cohesin variants and meiotic timing shape chromosome segregation accuracy
Yuanyuan Liu, Bohan Liu, Shuo Wang, Li Zhao, Qian Li, Feifei Qi, Ruoxi Wang, Jun Zhou, Jinmin Gao
, Available online  , doi: 10.1016/j.jgg.2025.06.003
Abstract (113) PDF (0)
Abstract:

The frequency of aneuploid gamete formation increases with maternal age, yet the effects of genetic variants on meiotic chromosome segregation accuracy during aging remain poorly understood. Using the multicellular organism Caenorhabditis elegans, we investigate the impact of mutations in the conserved cohesin complex on age-associated meiotic errors. Point mutations in the head domain of the cohesin component SMC-1, which alter local hydrophobicity, cause meiotic defects that vary with age. A severe mutation causes incomplete synapsis and defective crossover formation, and a minor one causes age-related diakinesis bivalent abnormalities. Notably, while the mild mutation causes defects only in aged worms, worms with the severe mutation exhibit significantly alleviated phenotypes with age. Genetic and cytological analyses suggest that this alleviation results from a slowed meiotic progression during early prophase, which restores impaired cohesin loading. These findings reveal that cohesin variants, meiotic progression speed during early prophase, and the overall duration of meiosis collectively shape the accuracy of meiotic chromosome segregation.

The nucleoporin CPR5 and histone chaperone NAP1 form a complex to negatively modulate plant immunity
Fenghui Xu, Xinyi Cai, Yi Yang, Yuehui Zhang, Weiyi Dou, Leiwen Pan, Shui Wang
, Available online  , doi: 10.1016/j.jgg.2025.06.002
Abstract (58) PDF (0)
Abstract:
3vGCIM: a compressed variance component mixed model for detecting QTL-by-environment interactions in RIL population
Mei Li, Yuan-Ming Zhang
, Available online  , doi: 10.1016/j.jgg.2025.05.011
Abstract (27) PDF (0)
Abstract:

Existing quantitative trait locus (QTL) mapping had low efficiency in identifying small-effect and closely linked QTL-by-environment interactions (QEIs) in RILs, especially in the era of global climate change. To address this challenge, here we integrate the compressed variance component mixed model with our GCIM to propose 3vGCIM for identifying QEIs in RILs, and extend 3vGCIM-random to 3vGCIM-fixed. 3vGCIM integrates genome-wide scanning with machine learning, significantly improving power. In the mixed full model, we consider all possible effects and control for all possible polygenic backgrounds. In simulation studies, 3vGCIM exhibits higher power (∼92.00%), higher accuracy of the estimates for QTL position (∼1.900 cM2) and effect (∼0.050), and lower false positive rate (∼0.48‰) and false negative rate (<8.10%) in three environments of 300 RILs each than ICIM (47.57%; 3.607 cM2, 0.583; 2.81‰; 52.43%) and MCIM (60.30%; 5.279 cM2, 0.274; 2.17‰; 39.70%). In the real data analysis of yield-related in 240 RILs, 3vGCIM mines more known genes (57∼60) and known gene-by-environment interactions (GEIs) (14∼19) and candidate GEIs (21∼23) than ICIM (27, 2, and 7), and MCIM (21, 1, and 3), especially in small-effect and linked QTLs and QEIs. This makes 3vGCIM a powerful and sensitive tool for QTL mapping and molecular QTL mapping.

Harness the wild: progress and perspectives in wheat genetic improvement
Xiubin Tian, Ziyu Wang, Wenxuan Liu, Yusheng Zhao
, Available online  , doi: 10.1016/j.jgg.2025.05.010
Abstract (50) PDF (2)
Abstract:

Common wheat (Triticum aestivum L.) is a staple hexaploid crop with numerous wild relatives. However, domestication and modern breeding have significantly narrowed its genetic diversity, diminishing its capacity to adapt to climate change. Wild relatives of wheat serve as a vital reservoir of genetic diversity, offering traits that enhance its resistance to various biotic and abiotic stresses. Over recent decades, remarkable progress has been made in utilizing superior genes from wild relatives to bolster wheat's defenses against diseases and pests, though the exploration of genes conferring abiotic stress tolerance has lagged behind. In this review, we summarize key advancements in the utilization of wild relatives for wheat enhancement over the past century, emphasizing both theoretical and technological innovations. Furthermore, we evaluate the potential contributions of wild relatives to address production challenges posed by climate changes. We also explore strategies for isolating superior genes and developing pre-breeding germplasm to support the future development of climate-resilient wheat varieties.

Single-cell transcriptome analysis revealed critical causative candidates for down syndrome-related lung diseases
Chunchun Zhi, Xucong Shi, Siqi Chen, Zhaowei Cai, Xiaoling Jiang
, Available online  , doi: 10.1016/j.jgg.2025.05.009
Abstract (210) PDF (0)
Abstract:

Down syndrome (DS) is caused by an extra copy of chromosome 21 (Hsa21). Children with DS have an increased frequency of respiratory tract infections, impaired alveolar and vascular development, and pulmonary hypertension. How trisomy 21 causes lung diseases remains poorly understood. In this study, we use the Dp16 mouse model, which contains a segmental chromosomal duplication of the entire Hsa21 syntenic region on mouse chromosome 16, to explore the gene dosage effects on DS-related lung diseases. The Dp16 mice present impaired alveolar development and inflammatory-like pathological changes. Single-cell transcriptome (scRNA-seq) analysis highlights increased APP-related interactions among male Dp16 lung cells. Specifically, altered antigen processing and presentation with increased MHC-II signaling are found in Dp16 immune cells. Reduced angiogenesis and altered inflammatory responses of Dp16 endothelial cells are also suggested. Moreover, scRNA-seq indicates hyperplasia of Dp16 vascular smooth muscle cells, which is validated by tissue immunofluorescence assessment. Transthoracic echocardiography further shows the existence of pulmonary hypertension in young Dp16 mice. Independent scRNA-seq analysis of the female lung cells recapitulates the majority of key findings identified in male mice, confirming the reproducibility of the results. Collectively, our results provide important clues for the further development of therapeutic approaches for DS-related lung diseases.

Genetic architecture of amyotrophic lateral sclerosis: a comprehensive review
Lamei Yuan, Yuewen Yang, Yi Guo, Hao Deng
, Available online  , doi: 10.1016/j.jgg.2025.05.008
Abstract (13) PDF (0)
Abstract:

Amyotrophic lateral sclerosis (ALS), one of the most prevalent neurodegenerative disorders, is pathologically characterized by the progressive degeneration of both upper and lower motor neurons, leading to muscle weakness, paralysis, and death within 2-4 years post-diagnosis. ALS is categorized into familial ALS (FALS) and sporadic ALS, with FALS accounting for approximately 10% of ALS cases. As a genetically heterogeneous disease, ALS exhibits diverse inheritance patterns, including autosomal dominant, autosomal recessive, and X-linked transmission, and genetic factors play pivotal roles in disease pathogenesis. To date, at least 34 disease-causing loci and 32 genes for ALS have been identified. The investigations of mutant protein products and the establishment of animal models have unraveled potential pathogenic pathways, offering insights into the mechanisms of neurodegeneration in ALS. This review focuses on ALS clinical characteristics, neuropathological features, causative loci/genes, genetic susceptibility factors, animal models, and pathogenic mechanisms, with particular attention to recent advances in genetic findings and pathogenic pathways of ALS. Elucidation of the genetic basis of ALS could provide the scientific foundation for personalized treatments to address this recalcitrant disease.

Xgr is involved in body size control in Drosophila through promoting glucose uptake in the Malpighian tubules
Zhiwei Lin, Zihao He, Jianfeng Guo, Xiaofang Ji, Ze Hu, Yingsen Tang, Chuanxian Wei, Jiyong Liu, Wenqi Wu, Jun Ma, Renjie Jiao
, Available online  , doi: 10.1016/j.jgg.2025.05.007
Abstract (233) PDF (0)
Abstract:

Body size control is fundamental to development and requires proper energy engagement. One of the key energy sensing factors is AMP-activated protein kinase (AMPK), which regulates glucose uptake to ensure ATP production and nutrition supply during development. Here, we identify that the mutation of xgr, a gene encoding an ATPase, results in a reduced body size in Drosophila. Xgr is primarily expressed in the epithelial cells of the Malpighian tubules and the midguts. Loss of xgr leads to the inactivation of the AMPK signaling due to an increased ATP level. Glucose reabsorption in the Malpighian tubules is significantly reduced, as the Glut1 translocation to the plasma membrane is significantly disrupted in the absence of Xgr function. Our results suggest that Xgr function in the Malpighian tubules is essential to systemic glucose supply and energy homeostasis at the organismal level, thereby impacting body size. Our findings provide a mechanistic connection between energy homeostasis and animal size control during development.

Decoding the complexity of coding and non-coding RNAs across maize anther development at the isoform level
Ge Yan, Xuxu Ma, Wei Huang, Chunyu Wang, Yingjia Han, Shufang Wang, Han Liu, Mei Zhang
, Available online  , doi: 10.1016/j.jgg.2025.05.005
Abstract (312) PDF (1)
Abstract:

Anther is a key male reproductive organ that is essential for the plant life cycle, from the sporophyte to the gametophyte generation. To explore the isoform-level transcriptional landscape of developing anthers in maize (Zea mays L.), we analyzed Iso-Seq data from anthers collected at 10 developmental stages, together with strand-specific RNA-seq, CAGE-seq, and PAS-seq data. Of the 152,026 high-confidence full-length isoforms identified, 68.8% have not been described; these include 22,365 isoforms that originate from previously unannotated loci and 82,167 novel isoforms that originate from annotated protein-coding genes. Using our newly developed strategy to detect dynamic expression patterns of isoforms, we identified 13,899 differentially variable regions (DVRs); surprisingly, 1275 genes contain more than two DVRs, revealing highly efficient utilization of limited genic regions. We identified 7876 long non-coding RNAs (lncRNAs) from 4098 loci, most of which were preferentially expressed during cell differentiation and meiosis. We also detected 371 long-range interactions involving intergenic lncRNAs (lincRNAs); interestingly, 243 were lincRNA–gene ones, and the interacting genes were highly expressed in anthers, suggesting that many potential lncRNA regulators of key genes are required for anther development. This study provides valuable resources and fundamental information for studying the essential transcripts of key genes during anther development.

Two-pore-domain potassium channel Sandman regulates intestinal stem cell homeostasis and tumorigenesis in Drosophila melanogaster
Chen Zheng, Jiadong Zheng, Xin Wang, Yue Zhang, Xianjue Ma, Li He
, Available online  , doi: 10.1016/j.jgg.2025.05.003
Abstract (171) PDF (0)
Abstract:

Potassium channels regulate diverse biological processes, ranging from cell proliferation to immune responses. However, the functions of potassium homeostasis and its regulatory mechanisms in adult stem cells and tumors remain poorly characterized. Here, we identify Sandman, a two-pore-domain potassium channel in Drosophila, as an essential regulator for the proliferation of intestinal stem cells and malignant tumors, while dispensable for the normal development processes. Mechanistically, loss of sandman elevates intracellular K+ concentration, leading to growth inhibition. This phenotype is rescued by pharmacological reduction of intracellular K+ levels using the K+ ionophore. Conversely, overexpression of sandman triggers stem cell death in most regions of the midgut, inhibits tumor growth, and induces a Notch loss-of-function phenotype in the posterior midgut. These effects are mediated predominantly via the induction of endoplasmic reticulum (ER) stress, as demonstrated by the complete rescue of phenotypes through the co-expression of Ire1 or Xbp1s. Additionally, human homologs of Sandman demonstrated similar ER stress-inducing capabilities, suggesting an evolutionarily conserved relationship between this channel and ER stress. Together, our findings identify Sandman as a shared regulatory node that governs Drosophila adult stem cell dynamics and tumorigenesis through bioelectric homeostasis, and reveal a link between the two-pore potassium channel and ER stress signaling.

DNA methylation landscapes of in vitro matured oocytes retrieved during endoscopic gynaecological procedures
Cui-Ling Lu, Xue-Ling Song, Xiao-Ying Zheng, Tian-Shu Song, Xiao-Na Wang, Jie Yan, Rui Yang, Rong Li, Jie Qiao
, Available online  , doi: 10.1016/j.jgg.2025.05.002
Abstract (29) PDF (0)
Abstract:

In vitro maturation (IVM) of human oocytes offers cost efficiency and minimal invasiveness, serving as a valuable supplementary tool in assisted reproduction for fertility preservation, ovarian hyperstimulation syndrome prevention, and other reproductive strategies. Despite its availability for three decades, the clinical use of IVM remains limited due to efficacy and safety concerns. This study examines the DNA methylation profile of IVM oocytes collected during laparoscopic/hysteroscopic surgeries compared to in vivo matured oocytes via reduced representation bisulfite sequencing. Results indicate IVM oocytes exhibit a higher global methylation level. Differentially methylated regions (DMR) analysis reveals that the in vitro group displays more hypermethylated and fewer hypomethylated DMRs compared to the in vivo group. Additionally, the in vitro group exhibits a higher level of non-CpG methylation than the in vivo group. However, no significant correlation between methylation levels and transcriptional activity in these oocytes is found, especially for those specific imprinted genes or genes related to embryonic development. These findings shed light on the epigenetic landscape of IVM oocytes, contributing to the ongoing assessment of their clinical feasibility and safety in assisted reproduction.

Unveiling cell-type-specific mode of evolution in comparative single-cell expression data
Tian Qin, Hongjiu Zhang, Zhengting Zou
, Available online  , doi: 10.1016/j.jgg.2025.04.022
Abstract (207) PDF (0)
Abstract:

While methodology for determining the mode of evolution in coding sequences has been well established, evaluation of adaptation events in emerging types of phenotype data needs further development. Here we propose an analysis framework (expression variance decomposition, EVaDe) for comparative single-cell expression data based on phenotypic evolution theory. After decomposing the gene expression variance into separate components, we use two strategies to identify genes exhibiting large between-taxon expression divergence and small within-cell-type expression noise in certain cell types, attributing this pattern to putative adaptive evolution. In a dataset of primate prefrontal cortex, we find that such human-specific key genes enrich with neurodevelopment-related functions, while most other genes exhibit neutral evolution patterns. Specific neuron types are found to harbor more of these key genes than other cell types, thus likely to have experienced more extensive adaptation. Reassuringly, at molecular sequence level, the key genes are significantly associated with the rapidly evolving conserved non-coding elements. An additional case analysis comparing the naked mole-rat (NMR) with the mouse suggests that innate-immunity-related genes and cell types have undergone putative expression adaptation in NMR. Overall, the EVaDe framework may effectively probe adaptive evolution mode in single-cell expression data.

Evolutionary divergence on the Qinghai-Tibet Plateau: How life-history traits shape the diversity of plateau zokor and pika populations
Yunyang Wei, Tao Zhang, Zifeng Li, Qinyang Hua, Liduo Yin, Menglong Lei, Shilei Zhao, Shanshan Gu, Xin Zhang, Hao He, Xuemei Lu
, Available online  , doi: 10.1016/j.jgg.2025.04.019
Abstract (358) PDF (0)
Abstract:

Understanding how species diverge and adapt is fundamental to unraveling biodiversity. While environmental impacts on species evolution are well-documented, the roles of intrinsic life-history traits remain underexplored. The Qinghai-Tibet Plateau, with its harsh conditions and unique biodiversity, offers a natural laboratory for such investigations. Here, we examined two sympatric small mammals—the solitary, low-dispersal plateau zokor (Eospalax baileyi) and the social, high-dispersal plateau pika (Ochotona curzoniae)—to elucidate how life-history traits shape population structures and adaptive strategies. Through whole-genome sequencing and cardiac-blood phenotype analyses, we reveal striking differences in their evolutionary trajectories. Despite enduring similar environmental pressures, plateau zokor populations exhibit pronounced genetic subdivisions, high inbreeding, and distinct local adaptations. In contrast, plateau pika populations display genetic panmixia, widespread diversity, and adaptive uniformity. Demographic inference highlights plateau zokors experienced severe population bottlenecks and restricted gene flow during glacial periods, underscoring the impact of dispersal capacity on evolutionary outcomes. Our findings demonstrate that intrinsic biological traits, particularly dispersal ability, fundamentally influence genetic architecture, population connectivity, and local adaptation. This study not only provides empirical evidence of how life-history traits shape evolutionary dynamics but also offers a framework for integrating intrinsic and extrinsic factors in understanding biodiversity formation.

High efficiency of thalassemia prevention by next-generation sequencing: a real-world cohort study in two centers of China
Jinman Zhang, Wenqian Zhang, Haoqing Zhang, Aiqi Cai, Caiyun Li, Ling Liu, Jufang Tan, Yang Yang, Wen Yuan, Jing He, Shiping Chen, Yingli Cao, Yan Zhang, Jie Zhang, Rui Zhou, Shuai Hou, Dongqun Huang, Danjing Chen, Zhiyu Peng, Dongzhu Lei, Baosheng Zhu
, Available online  , doi: 10.1016/j.jgg.2025.04.018
Abstract (31) PDF (0)
Abstract:

The occurrence of severe thalassemia, an inherited blood disorder that is either blood-transfusion-dependent or fatal, can be mitigated through carrier screening. Here, we aim to evaluate the effectiveness and outcomes of pre-conceptional and early pregnancy screening initiatives for severe thalassemia prevention in a diverse population of 28,043 women. Using next-generation sequencing (NGS), we identify 4,226 (15.07%) thalassemia carriers across 29 ethnic groups and categorize them into high- (0.75%), low- (25.86%), and unknown-risk (69.19%) groups based on their spouses’ screening results. Post-screening follow-up reveals 59 fetuses with severe thalassemia exclusively in high-risk couples, underscoring the efficacy of risk classification. Among 25,053 live births over 6 months of age, two severe thalassemia infants were born to unknown-risk couples, which was attributed to incomplete screening and late NGS-based testing for a rare variant. Notably, 64 rare variants are identified in 287 individuals, highlighting the genetic heterogeneity of thalassemia. We also observe that migrant flow significantly impacts carrier rates, with 93.90% of migrants to Chenzhou originating from high-prevalence regions in southern China. Our study demonstrates that NGS-based screening during pre-conception and early pregnancy is effective for severe thalassemia prevention, emphasizing the need for continuous screening efforts in areas with high and underestimated prevalence.

Near-gapless telomere-to-telomere reference nuclear genome and variable mitochondrial genome of Amborella trichopoda
Zhonglong Guo, Jing-Fang Guo, Zhi-Yan Wei, Ren-Gang Zhang, Scott McMahan, Shuai Nie, Xue-Mei Yan, Shan-Shan Zhou, Quan-Zheng Yun, Jia-Yi Wu, Jing Ge, Yong Yang, Jia-Yu Xue, Jian-Feng Mao
, Available online  , doi: 10.1016/j.jgg.2025.04.016
Abstract (24) PDF (0)
Abstract:
Oncogenic Ras, Yki and Notch signals converge to confer clone competitiveness through Upd2
Ying Wang, Rui Huang, Minfeng Deng, Jingjing He, Mingxi Deng, Toyotaka Ishibashi, Cong Yu, Zongzhao Zhai, Yan Yan
, Available online  , doi: 10.1016/j.jgg.2025.04.017
Abstract (18) PDF (0)
Abstract:

Cell competition is an evolutionarily ancient mechanism that functions to remove unfit or dangerous clonal cells in a multicellular community. A classical model is the removal of polarity-deficient clones, such as the precancerous scribble (scrib) mutant clones, in Drosophila imaginal discs. The activation of Ras, Yki, or Notch signaling robustly reverses the scrib mutant clonal fate from elimination to tumorous growth. Whether these signals converge to adopt a common mechanism to overcome the elimination pressure posed by cell competition remains unclear. Using single-cell transcriptomics, we find that a critical converging point downstream of Ras, Yki, and Notch signals is the upregulation of Upd2, an IL-6 family cytokine. Overexpression of Upd2 is sufficient to rescue the scrib mutant clones from elimination. Depletion of Upd2 blocks the growth of the scrib mutant clones with active Ras, Yki, and Notch signals. Moreover, Upd2 overexpression promotes robust intestinal stem cell (ISC) proliferation, while Upd2 is intrinsically required in ISCs for the growth of the adult intestine. Together, these results identify Upd2 as a crucial cell fitness factor that sustains tissue growth but can potentiate tumorigenesis when deregulated.

Wheat PKL genes regulate pre-harvest sprouting and yield-related traits
Wanqing Bai, Ziyi Yang, Xuchang Yu, Shuxian Huang, Yufan Wang, Yexing Jing, Yunwei Zhang, Jiaqiang Sun
, Available online  , doi: 10.1016/j.jgg.2025.04.011
Abstract (23) PDF (0)
Abstract:
RiboParser/RiboShiny: an integrated platform for comprehensive analysis and visualization of Ribo-seq data
Shuchao Ren, Yinan Li, Zhipeng Zhou
, Available online  , doi: 10.1016/j.jgg.2025.04.010
Abstract (38) PDF (0)
Abstract:

Translation is a crucial step in gene expression. Over the past decade, the development and application of Ribosome profiling (Ribo-seq) have significantly advanced our understanding of translational regulation in vivo. However, the analysis and visualization of Ribo-seq data remain challenging. Despite the availability of various analytical pipelines, improvements in comprehensiveness, accuracy, and user-friendliness are still necessary. In this study, we develop RiboParser/RiboShiny, a robust framework for analyzing and visualizing Ribo-seq data. Building on published methods, we optimize ribosome structure-based and start/stop-based models to improve the accuracy and stability of P-site detection, even in species with a high proportion of leaderless transcripts. Leveraging these improvements, RiboParser offers comprehensive analyses, including quality control, gene-level analysis, codon-level analysis, and the analysis of Ribo-seq variants. Meanwhile, RiboShiny provides a user-friendly and adaptable platform for data visualization, facilitating deeper insights into the translational landscape. Furthermore, the integration of standardized genome annotation renders our platform universally applicable to various organisms with sequenced genomes. This framework has the potential to significantly improve the precision and efficiency of Ribo-seq data interpretation, thereby deepening our understanding of translational regulation.

Rhpn2 regulates the development and function of vestibular sensory hair cells through the RhoA signaling in zebrafish
Yubei Dai, Qianqian Li, Jiaju Deng, Sihang Wu, Guiyi Zhang, Yuebo Hu, Yuqian Shen, Dong Liu, Han Wu, Jie Gong
, Available online  , doi: 10.1016/j.jgg.2025.04.006
Abstract (32) PDF (0)
Abstract:

Hearing and balance disorders are significant health issues primarily caused by developmental defects or the irreversible loss of sensory hair cells (HCs). Identifying the underlying genes involved in the morphogenesis and development of HCs is crucial. Our current study highlights rhpn2, a member of rho-binding proteins, as essential for vestibular HC development. The rhpn2 gene is highly expressed in the crista and macula HCs. Loss of rhpn2 function in zebrafish reduces the otic vesicle area and vestibular HC number, accompanied by vestibular dysfunction. Shorter stereocilia and compromised mechanotransduction channel function are found in the crista HCs of rhpn2 mutants. Transcriptome RNA sequencing analysis predicts the potential interaction of rhpn2 with rhoab. Furthermore, co-immunoprecipitation confirms that Rhpn2 directly binds to RhoA, validating the interaction of the two proteins. rhpn2 knockout leads to a decreased expression of rock2b, a canonical RhoA signaling pathway gene. Treatment with the RhoA activator or exogenous rock2b mRNA injection mitigates crista HC stereocilia defects in rhpn2 mutants. This study uncovers the role of rhpn2 in vestibular HC development and stereocilia formation via mediating the RhoA signaling pathway, providing a target for the treatment of balance disorders.

Regulation of maize kernel development via divergent activation of α-Zein genes by transcription factors O11, O2, and PBF1
Runmiao Tian, Zeyuan Yang, Ruihua Yang, Sihao Wang, Qingwen Shen, Guifeng Wang, Hongqiu Wang, Qingqian Zhou, Jihua Tang, Zhiyuan Fu
, Available online  , doi: 10.1016/j.jgg.2025.04.008
Abstract (81) PDF (0)
Abstract:

α-Zeins, the major maize endosperm storage proteins, are transcriptionally regulated by Opaque 2 (O2) and PROLAMIN-BOX BINDING FACTOR1 (PBF1), with Opaque 11 (O11) functioning upstream of them. However, whether O11 directly binds to α-zein genes and its regulatory interactions with O2 and PBF1 remain unclear. Using the small-kernel mutant sw1, which exhibits decreased 19-kDa and increased 22-kDa α-zein, we positionally cloned O11 and found it directly binds to G-box/E-box motifs. O11 activates 19-kDa α-zein transcription, stronger than PBF1 but weaker than O2. Notably, PBF1 competitively binds to overlapping E-box/P-box motif, and represses O11-mediated transactivation. Although O11 does not physically interact with O2, it participates in the O2-centered hierarchical network to enhance α-zein expression. sw1 o2 and sw1 pbf1 double mutants exhibit smaller, more opaque kernels with further reduced 19-kDa and 22-kDa α-zeins compared to the single mutants, suggesting distinct regulatory effects of these transcription factors on 19-kDa and 22-kDa α-zein genes. Promoter motif analysis suggests that O11, PBF1, and O2 directly regulate 19-kDa α-zein genes, while O11 indirectly controls 22-kDa α-zein genes via O2 and PBF1 modulation. These findings identify the unique and coordinated roles of O11, O2, and PBF1 in regulating α-zein genes and kernel development.

Biallelic MED16 variants disrupt neural development and lead to an intellectual disability syndrome
Yan Huang, Zhenglong Xiang, Yaqin Xiang, Hu Pan, Mei He, Zhenming Guo, Oguz Kanca, Chen Liu, Zhao Zhang, Huaizhe Zhan, Yuan Wang, Qing-Ran Bai, Hugo J. Bellen, Hua Wang, Shan Bian, Xiao Mao
, Available online  , doi: 10.1016/j.jgg.2025.04.004
Abstract (20) PDF (0)
Abstract:

Mediator Complex Subunit 16 (MED16, MIM: 604062) is a member of the Mediator complex, which controls many aspects of transcriptional activity in all eukaryotes. Here, we report two individuals from a non-consanguineous family with biallelic variants in MED16 identified by exome sequencing. The affected individuals present with global developmental delay, intellectual disability, and dysmorphisms. To assess the pathogenicity of the variants, functional studies are performed in Drosophila and patient-derived cells. The fly ortholog med16 is expressed in neurons and some glia of the developing central nervous system (CNS). Loss of med16 leads to a reduction in eclosion and lifespan, as well as impaired synaptic transmission. In neurons differentiated from the patient-derived induced pluripotent stem cells (iPSCs), the neurite outgrowth is impaired and rescued by expression of exogenous MED16. The patient-associated variants behave as loss-of-function (LoF) alleles in flies and iPSCs. Additionally, the transcription of genes related to neuronal maturation and function is preferentially altered in patient cells relative to differentiated H9 controls. In summary, our findings support that MED16 is important for appropriate development and function, and that biallelic MED16 variants cause a neurodevelopmental disease.

GCH1 contributes to high-altitude adaptation in Tibetans by regulating blood nitric oxide
Yongbo Guo, Wangshan Zheng, Tian Yue, Baimakangzhuo, Xuebin Qi, Kai Liu, Liya Li, Yaoxi He, Bing Su
, Available online  , doi: 10.1016/j.jgg.2025.04.005
Abstract (294) PDF (0)
Abstract:

Nitric oxide (NO) is a key vasodilator that regulates vascular pressure and blood flow. Tibetans have developed a “blunted” mechanism for regulating NO levels at high altitude, with GTP cyclohydrolase 1 (GCH1) identified as a key candidate gene. Here, we present comprehensive genetic and functional analyses of GCH1, which exhibits strong Darwinian positive selection in Tibetans. We show that Tibetan-enriched GCH1 variants down-regulate its expression in the blood of Tibetans. Based on this observation, we generate the heterozygous Gch1 knockout (Gch1+/–) mouse model to simulate its downregulation in Tibetans. We find that under prolonged hypoxia, the Gch1+/– mice have relatively higher blood NO and blood oxygen saturation levels compared with the wild-type (WT) controls, providing better oxygen supplies to the cardiovascular and pulmonary systems. Markedly, hypoxia-induced cardiac hypertrophy and pulmonary remodeling are significantly attenuated in the Gch1+/– mice compared with the WT controls, likely due to the adaptive changes in molecular regulations related to metabolism, inflammation, circadian rhythm, extracellular matrix, and oxidative stress. This study sheds light on the role of GCH1 in regulating blood NO, contributing to the physiological adaptation of the cardiovascular and pulmonary systems in Tibetans at high altitude.

Deep sequencing reveals SLC35A2 somatic variants in MOGHE: molecular and clinical insights
Huaxia Luo, Xiaoqin Ruan, Xianyu Liu, Qingzhu Liu, Yu Sun, Yao Wang, Jixin Zhang, Lixin Cai, Yuwu Jiang, Ye Wu
, Available online  , doi: 10.1016/j.jgg.2025.04.002
Abstract (25) PDF (0)
Abstract:
Coordinated regulation of cortical astrocyte maturation by OLIG1 and OLIG2 through BMP7 signaling modulation
Ziwu Wang, Yu Tian, Tongye Fu, Feihong Yang, Jialin Li, Lin Yang, Wen Zhang, Wenhui Zheng, Xin Jiang, Zhejun Xu, Yan You, Xiaosu Li, Guoping Liu, Yunli Xie, Zhengang Yang, Dashi Qi, Zhuangzhi Zhang
, Available online  , doi: 10.1016/j.jgg.2025.03.008
Abstract (241) PDF (0)
Abstract:

Astrocyte maturation is crucial for brain function, yet the mechanisms regulating this process remain poorly understood. In this study, we identify the bHLH transcription factors Olig1 and Olig2 as essential coordinators of cortical astrocyte maturation. We demonstrate that Olig1 and Olig2 work synergistically to regulate cortical astrocyte maturation by modulating Bmp7 expression. Genetic ablation of both Olig1 and Olig2 results in defective astrocyte morphology, including reduced process complexity and an immature gene expression profile. Single-cell RNA sequencing reveals a shift towards a less mature astrocyte state, marked by elevated levels of HOPX and GFAP, resembling human astrocytes. Mechanistically, Olig1 and Olig2 bind directly to the Bmp7 enhancer, repressing its expression to promote astrocyte maturation. Overexpression of Bmp7 in vivo replicates the astrocyte defects seen in Olig1/2 double mutants, confirming the critical role of BMP7 signaling in this process. These findings provide insights into the transcriptional and signaling pathways regulating astrocyte development and highlight Olig1 and Olig2 as key regulators of cortical astrocyte maturation, with potential implications for understanding glial dysfunction in neurological diseases.

The phospholipid scramblase PLSCR5 is regulated by POU4F3 and required for hair cell stereocilia homeostasis and auditory functions
Sihao Gong, Qing Liu, Haibo Du, Linqing Zhang, Chengwen Zhu, Zhigang Xu, Xia Gao, Guang-Jie Zhu, Guoqiang Wan
, Available online  , doi: 10.1016/j.jgg.2025.03.003
Abstract (253) PDF (0)
Abstract:

Hearing relies on the structural and functional integrity of cochlear hair cells, particularly their apical F-actin-filled stereocilia. Phospholipid scramblases are important for maintaining membrane asymmetry, but their roles in the stereocilia and auditory functions are not fully understood. Here, we identify Plscr5 as a downstream target of the transcription factor POU4F3 essential for hair cell function, whose mutation causes human DFNA15 deafness. Plscr5 knockout mice exhibit progressive hearing loss due to stereocilia degeneration and hair cell loss. Functional analyses reveal that PLSCR5 contributes to phosphatidylserine externalization in hair cell apical membranes, particularly in inner hair cells, and is important for outer hair cell and stereocilia maintenance. Our findings highlight PLSCR5 as an important downstream effector of POU4F3 and regulator of PS externalization and membrane dynamics required for auditory functions.

Resolving the spatial and cellular architecture of intra-tumor heterogeneity by multi-region dissection of lung adenocarcinoma
Song Mei, Xiaolei Wang, Mengmeng Zhao, Qing Huang, Yixuan Huang, Mingming Su, Xinlei Zhang, Xu Wang, Xueyu Hao, Tianning Wang, Yanhua Wu, Yuanhui Ma, Jingnan Wang, Peng Zhang, Yan Zheng
, Available online  , doi: 10.1016/j.jgg.2025.02.006
Abstract (288) PDF (0)
Abstract:

Although the spatial characteristics within the tumor microenvironment (TME) of lung adenocarcinoma (LUAD) have been identified, the mechanisms by which these factors promote LUAD progression and immune evasion remain unclear. Using spatial transcriptomics (ST) and single-cell RNA-sequencing (scRNA-seq) data from multi-regional LUAD biopsies consisting of tumor core, tumor edge, and normal area, we sought to delineate the spatial heterogeneity and driving factors of cell colocalization. Two cancer cell sub-clusters (Cancer_c1 and Cancer_c2), associated with LUAD initiation and metastasis, respectively, exhibit distinct spatial distributions and immune cell colocalizations. In particular, Cancer_c1, enriched within the tumor core, could directly interact with B cells or indirectly recruit B cells through macrophages. Conversely, Cancer_c2 enriched within the tumor edge exhibits colocalization with CD8+ T cells. Collectively, our work elucidates the spatial distribution of cancer cell subtypes and their interaction with immune cells in the core and edge of LUAD, providing insights for developing therapeutic strategies for cancer intervention.

Diverse functions of sex determination gene doublesex on sexually dimorphic neuronal development and behaviors
Jiangtao Chen, Wen Tu, Ziqi Li, Mingze Ma, Simei Jiang, Wenyue Guan, Rong Wang, Yufeng Pan, Qionglin Peng
, Available online  , doi: 10.1016/j.jgg.2025.02.005
Abstract (177) PDF (0)
Abstract:

Sex-specific neurons play pivotal roles in regulating sexually dimorphic behaviors. In insects, the sex determination gene doublesex (dsx) establishes major sexual dimorphism of the nervous system, in which male-specific dsxM promotes neuronal development, while female-specific dsxF inhibits neuronal development by promoting neuronal apoptosis. In this study, we find that dsx regulates the number of dsx-expressing central neurons in Drosophila in cell-specific manners. Although dsxM overall promotes an increase in the number of dsx neurons, it inhibits the emergence of specific pC1 neurons. dsxF reduces the number of different pC1/pC2 subtypes, but promotes the formation of pC1d. We also find that dsxM and dsxF barely affect the number of some pC2 neurons. Changes in the number of pC1/pC2 neuron numbers alter their roles in regulating different behaviors, including courtship, aggression, and locomotion. Our results illustrate the multifaceted functions of dsx in sexually dimorphic neuronal development and behaviors.

Varying Bifidobacterium species in the maternal-infant gut microbiota correlate with distinct early neurodevelopmental outcomes
Cong Liu, Qun Lu, Qi Xi, Shuxin Xiao, Jiangbo Du, Rui Qin, Jinghan Wang, Bo Xu, Xiumei Han, Kun Zhou, Shiyao Tao, Hong Lv, Yangqian Jiang, Tao Jiang, Kan Ye, Guangfu Jin, Hongxia Ma, Yankai Xia, Hongbing Shen, Xingyin Liu, Yuan Lin, Zhibin Hu
, Available online  , doi: 10.1016/j.jgg.2025.01.015
Abstract (185) PDF (0)
Abstract:

The impact of mother-infant microbiota on neurodevelopment is an area of interest, but longitudinal studies are scarce. Using a cohort of 520 families from the Jiangsu birth cohort in China, we reveal that the maternal gut microbiota during early pregnancy play a substantial role, accounting for 3.34% of the variance in offspring neurodevelopmental scores. This contribution is notably higher than the 1.24% attributed to the infants’ own microbiota at 1 year of age, underscoring the significant influence of maternal gut health on early child development. Remarkably, an elevation in maternal Bifidobacterium pseudocatenulatum is linked to decreased cognitive scores, whereas an enrichment of Bifidobacterium longum at 1 year of age is associated with higher cognitive scores. Furthermore, we find that maternal B. pseudocatenulatum is linked to the heterolactic fermentation metabolic pathway, while infant B. longum is associated with the Bifidobacterium shunt pathway. In summary, our analysis implies that maternal and infant gut microbiota play a distinct role in neurodevelopment, suggesting potential strategies for improving neurodevelopmental outcomes during early pregnancy or infant development by targeting gut microbiota composition.

Genomic predictions of invasiveness and adaptability of the cotton bollworm in response to climate change
Qi Xu, Minghui Jin, Hua Xiao, Yan Peng, Fan Zhang, Hongran Li, Kongming Wu, Yutao Xiao
, Available online  , doi: 10.1016/j.jgg.2025.01.016
Abstract (15) PDF (0)
Abstract:

Agricultural pests cause enormous losses in annual agricultural production. Understanding the evolutionary responses and adaptive capacity of agricultural pests under climate change is crucial for establishing sustainable and environmentally friendly agricultural pest management. In this study, we integrate climate modeling and landscape genomics to investigate the distributional dynamics of the cotton bollworm (Helicoverpa armigera) in the adaptation to local environments and resilience to future climate change. Notably, the predicted inhabitable areas with higher suitability for the cotton bollworm could be eight times larger in the coming decades. Climate change is one of the factors driving the dynamics of distribution and population differentiation of the cotton bollworm. Approximately 19,000 years ago, the cotton bollworm expanded from its ancestral African population, followed by gradual occupations of the European, Asian, Oceanian, and American continents. Furthermore, we identify seven subpopulations with high dispersal and adaptability which may have an increased risk of invasion potential. Additionally, a large number of candidate genes and SNPs linked to climatic adaptation were mapped. These findings could inform sustainable pest management strategies in the face of climate change, aiding future pest forecasting and management planning.

uniLIVER: a human liver cell atlas for data-driven cellular state mapping
Yanhong Wu, Yuhan Fan, Yuxin Miao, Yuman Li, Guifang Du, Zeyu Chen, Jinmei Diao, Yu-Ann Chen, Mingli Ye, Renke You, Amin Chen, Yixin Chen, Wenrui Li, Wenbo Guo, Jiahong Dong, Xuegong Zhang, Yunfang Wang, Jin Gu
, Available online  , doi: 10.1016/j.jgg.2025.01.017
Abstract (8) PDF (0)
Abstract:

The liver performs several vital functions such as metabolism, toxin removal, and glucose storage through the coordination of various cell types. With the recent breakthrough of the single-cell/single-nucleus RNA-seq (sc/snRNA-seq) techniques, there is a great opportunity to establish a reference cell map of the liver at single-cell resolution with transcriptome-wise features. In this study, we build a unified liver cell atlas uniLIVER (http://lifeome.net/database/uniliver) by integrative analysis of a large-scale sc/snRNA-seq data collection of normal human liver with 331,125 cells and 79 samples from 6 datasets. Moreover, we introduce LiverCT, a novel machine learning based method for mapping any query dataset to the liver reference map by introducing the definition of “variant” cellular states analogy to the sequence variants in genomic analysis. Applying LiverCT on liver cancer datasets, we find that the “deviated” states of T cells are highly correlated with the stress pathway activities in hepatocellular carcinoma, and the enrichments of tumor cells with the hepatocyte-cholangiocyte “intermediate” states significantly indicate poor prognosis. Besides, we find that the tumor cells of different patients have different zonation tendencies and this zonation tendency is also significantly associated with the prognosis. This reference atlas mapping framework can also be extended to any other tissues.

Multifaceted interplays between the essential players and lipid peroxidation in ferroptosis
Conghe Liu, Zhihao Liu, Zheng Dong, Sijin Liu, Haidong Kan, Shuping Zhang
, Available online  , doi: 10.1016/j.jgg.2025.01.009
Abstract (10) PDF (0)
Abstract:

Ferroptosis, a type of programmed cell death, represents a distinct paradigm in cell biology. It is characterized by the iron-dependent accumulation of reactive oxygen species, which induce lipid peroxidation (LPO), and is orchestrated by the interplay between iron, lipid peroxides, and glutathione. In this review, we emphasize the frequently overlooked role of iron in LPO beyond the classical iron-driven Fenton reaction in several crucial processes that regulate cellular iron homeostasis, including iron intake and export as well as ferritinophagy, and the emerging roles of endoplasmic reticulum-resident flavoprotein oxidoreductases, especially P450 oxidoreductases, in modulating LPO. We summarize how various types of fatty acids (FAs), including saturated, monounsaturated, and polyunsaturated FAs, differentially influence ferroptosis when incorporated into phospholipids. Furthermore, we highlight the therapeutic potential of targeting LPO to mitigate ferroptosis and discuss the regulatory mechanisms of endogenous lipophilic radical-trapping antioxidants that confer resistance to ferroptosis, shedding light on therapeutic avenues for ferroptosis-associated diseases.

Enhancing crop yields to ensure food security by optimizing photosynthesis
Chunrong Li, Xuejia Du, Cuimin Liu
, Available online  , doi: 10.1016/j.jgg.2025.01.002
Abstract (11) PDF (0)
Abstract:

The crop yields achieved through traditional plant breeding techniques appear to be nearing a plateau. Therefore, it is essential to accelerate advancements in photosynthesis, the fundamental process by which plants convert light energy into chemical energy, to further enhance crop yields. Research focused on improving photosynthesis holds significant promise for increasing sustainable agricultural productivity and addressing challenges related to global food security. This review examines the latest advancements and strategies aimed at boosting crop yields by enhancing photosynthetic efficiency. There has been a linear increase in yield over the years in historically released germplasm selected through traditional breeding methods, and this increase is accompanied by improved photosynthesis. We explore various aspects of the light reactions designed to enhance crop yield, including light harvest efficiency through smart canopy systems, expanding the absorbed light spectrum to include far-red light, optimizing non-photochemical quenching, and accelerating electron transport flux. At the same time, we investigate carbon reactions that can enhance crop yield, such as manipulating Rubisco activity, improving the Calvin-Benson-Bassham (CBB) cycle, introducing CO2 concentrating mechanisms (CCMs) in C3 plants, and optimizing carbon allocation. These strategies could significantly impact crop yield enhancement and help bridge the yield gap.

Original Research
Development and optimization of a cell cycle molecular clock based on the ABE-CRISPR system
Xinyuan Dai, Jiachuan Jin, Zhaojun Wang, Shihua Zhang
, Available online  , doi: 10.1016/j.jgg.2026.10.004
Abstract (90) PDF (2)
Abstract:
Clustered regularly interspaced short palindromic repeats (CRISPR)-guided multiplex mutagenesis has emerged as a tool for lineage tracing; however, existing systems often exhibit disordered mutation patterns and cannot reliably record cell division history. To overcome these limitations, we develop the adenine base editor template number (ATN) system, which implements ordered, stepwise editing of double-stranded DNA through a programmable cascade of modifications. The ATN system incrementally marks each cell cycle, providing a quantitative representation of proliferation events. Once a predetermined number of divisions is reached, the system triggers activation of a downstream gene through frame restoration, effectively serving as a cellular “molecular clock” that links cell age to inducible genetic control. This approach therefore offers a systematic framework for studying cell proliferation dynamics and enables temporally regulated manipulation of cell fate.
A calcium-independent CDPK positively regulates drought resistance in maize by modulating the ABA signaling pathway
Tingting Fang, Yinan Han, Yanyu Chen, Xiaoying Hu, Ziting Zhong, Tingting Wang, Yu Wang, Junsheng Qi, Shuhua Yang, Zhizhong Gong
, Available online  , doi: 10.1016/j.jgg.2026.10.002
Abstract (140) PDF (1)
Abstract:
Calcium-dependent protein kinases (CDPKs/CPKs) play crucial roles in plant responses to drought stress. In this study, we reveal that ZmCPK1, an atypical CDPK whose kinase activity is calcium-independent, acts as a positive regulator of drought resistance in maize. ZmCPK1 is a constitutively active kinase interacting with and phosphorylates multiple components of the abscisic acid (ABA) signaling pathway, including ZmSLAC1, ZmSnRK2.10, and ZmPP2C11. In the absence of ABA, ZmCPK1 may phosphorylate ZmSnRK2.10 and the N-terminus of ZmSLAC1. ZmPP2C11 reverses this phosphorylation by directly dephosphorylating the N-terminus of ZmSLAC1 rather than suppressing ZmCPK1 kinase activity. ABA treatment and dehydration enhance ZmCPK1-mediated phosphorylation of the N-terminus of ZmSLAC1 at the conserved serine 54 (S54) without affecting ZmCPK1 autophosphorylation, consistent with reduced dephosphorylation resulting from the inhibition of ZmPP2C11 by ABA-bound pyrabactin resistance 1-like (PYL) proteins. Notably, ZmCPK1 phosphorylates ZmPP2C11 at S277, a modification that potentially inhibits its phosphatase activity, and phosphorylates multiple sites on ZmSnRK2.10 to activate it. Thus, ZmCPK1 acts as a constitutive kinase; under optimal growth conditions, it interacts with ZmPP2C11, which dephosphorylates the targets phosphorylated by ZmCPK1. Under drought stress, ZmPP2C11 is inhibited, and phosphorylated downstream targets are activated to promote stomatal closure, thereby positively regulating drought resistance in maize.
Plant-mediated RNAi via Panonychus citri-derived miRNAs disrupts adult emergence and confers mite resistance
Chuanzhen Li, Yuchuang Li, Jingyao Zhang, Boyi Li, Menghao Xia, Jinjun Wang, Wei Dou
, Available online  , doi: 10.1016/j.jgg.2026.09.012
Abstract (132) PDF (0)
Abstract:
MicroRNAs (miRNAs), small non-coding RNAs that trigger RNA interference (RNAi), are promising candidates for pest control, yet their efficacy against non-insect pests remains poorly explored. This study evaluates endogenous mite-derived miRNAs as transgenes for crop resistance against Panonychus citri, a major pest with rapid acaricide resistance that severely damages citrus, soybean, and other cash crops. Through developmental miRNA libraries of P. citri, we identify miR-750 and miR-200a-5p as key regulators during the deutonymph-to-adult transition, whose overexpression significantly inhibits adult eclosion. Both in vivo and in vitro assays confirm that miR-750 targets both NFYB and ADRM1, whereas miR-200a-5p specifically targets ADRM1; dsRNA-mediated silencing of these genes recapitulates eclosion defects induced by miRNA overexpression. We generate single-copy transgenic soybean lines with high expression of miR-750 (line 7#6) or miR-200a-5p (line 2#14). Detached-leaf and whole-plant bioassays using T1 generations reveal that both lines exhibit strong resistance against P. citri. Notably, neither transgenic line adversely affects the non-target predatory mite Neoseiulus barkeri, either via direct miRNA exposure or prey-mediated transfer. The resistance against P. citri and biosafety toward N. barkeri are retained in T2 generations. These findings highlight the potential of endogenous miRNAs as effective targets for developing eco-friendly and sustainable pest management strategies.
OsToc159 is essential for amyloplast development and starch granule initiation in rice endosperm
Zhuangzhuang Shan, Yongxiang Wang, Yushuang Fu, Mingqing Ma, Jiaming Duan, Jiahuan Xu, Haigang Yan, Changyuan Zhu, Shuaijie Niu, Qun Ji, Hongming Wu, Hancong Xu, Xiaoqing Hu, Wenxiang Zhang, Feilong Du, Chaofan Wang, Xinlong Fan, Yuan Li, Yunlu Tian, Ling Jiang, Wenwei Zhang, Jianmin Wan
, Available online  , doi: 10.1016/j.jgg.2026.09.014
Abstract (63) PDF (0)
Abstract:
In cereal endosperm, starch is biosynthesized within specialized plastids termed amyloplasts through the coordinated activity of a suite of enzymes. The development of amyloplasts requires the import of nuclear-encoded preproteins from the cytosol via the translocon. However, little is known about how the translocon influences amyloplast development and starch synthesis in cereal endosperm. Here, we functionally characterize the rice floury endosperm mutant b303, which is defective in amyloplast development and starch granule initiation. Map-based cloning of B303 identifies a mutation in OsTOC159, which encodes a GTPase receptor subunit of the translocon at the outer envelope membrane of chloroplasts (TOC) complex. OsToc159 is highly expressed in developing endosperm and localizes to the amyloplast membrane. Combined with biochemical evidence, protoplast-based import assays suggest that OsToc159 potentially modulates the plastid import of a subset of starch biosynthetic proteins. Moreover, OsToc159 likely interacts with another GTPase receptor, OsToc34, to synergistically facilitate the posttranslational translocation of isoamylase 1 (ISA1). Collectively, our data suggest that OsToc159 may function as a receptor involved in the import of a subset of plastid proteins required for amyloplast development and starch biosynthesis.
A transcription factor-focused CRISPR screen identifies SKI as a BCL11A-independent repressor of ζ-globin
Qing Liu, Jianxiang Zhang, Mengli Zhang, Ye Xin, Yuhua Ye, Min Jiang, Xiangmin Xu, Peng Xu
, Available online  , doi: 10.1016/j.jgg.2026.09.003
Abstract (100) PDF (1)
Abstract:
The regulation of α-like globin genes, particularly the embryonic ζ-globin gene (HBZ), remains incompletely understood. To identify transcriptional regulators of HBZ, we establish a GFP reporter system based on the HBZ-P2A-GFP allele in erythroid cell lines and conduct a CRISPR/Cas9 screen targeting 1639 transcription factors. This screen identifies SKI as a potent HBZ repressor. Functional validation shows that SKI loss increases HBZ expression without impairing erythropoiesis, whereas SKI overexpression suppresses HBZ. Tet-on-inducible SKI overexpression and auxin-inducible SKI degradation indicate that SKI rapidly represses HBZ transcription. Transcriptome profiling further reveals that SKI deletion activates HBZ while minimally affecting other erythroid genes. Mechanistically, genome-wide occupancy analyses show that SKI binds the distal enhancers HS-10 and HS-40, with partial co-occupancy by BCL11A. Despite this overlap, dual knockout of SKI and BCL11A synergistically increases HBZ expression, as does base editing of the SKI-binding site within HS-10. We also identify a naturally occurring variant (chr16:193207G>A) within this enhancer in α-thalassemia patients with elevated ζ-globin levels. Together, these findings establish SKI as a direct, BCL11A-independent transcriptional repressor of ζ-globin. This work advances our understanding of globin gene regulation and suggests targeted ζ-globin reactivation as a potential therapeutic strategy for α-thalassemia.
Regulatory divergence of duplicated VIH genes links phosphate signaling to nodule development in Medicago truncatula
Boyu Zhao, Yihan Zhang, Junjie Ye, Meixi Li, Hsihua Wang, Yang Zhang
, Available online  , doi: 10.1016/j.jgg.2026.08.008
Abstract (161) PDF (0)
Abstract:
Primary metabolites and their derivatives often serve as intracellular signals. Inositol pyrophosphates are central regulators of phosphate signaling, but their roles in legume nodulation remain poorly understood. Here, we identify two conserved Vip1 Homolog/Diphosphoinositol Pentakisphosphate Kinase genes, MtVIH1 and MtVIH2, in Medicago truncatula. Biochemical analyses show that the kinase domains of both MtVIH1 and MtVIH2 retain PP-InsP kinase activity. However, transcriptomic and expression analyses reveal regulatory divergence between the duplicated genes, with MtVIH2 being preferentially induced during nodulation and co-expressed with the phosphate starvation marker Mt4. Silencing MtVIH2 reduces nodule formation, nitrogenase activity, and mature nodule marker expression, and alters extractable inorganic Pi accumulation in nodules. Consistently, CRISPR/Cas9-mediated editing of MtVIH2 reproduces these defects, whereas Mtvih1-cas9 shows much weaker effects and Mtvih1/2-cas9 edited roots display phenotypes comparable to Mtvih2-cas9. These results demonstrate that regulatory divergence between duplicated VIH genes is associated with a predominant role of MtVIH2 in nodule development. Promoter analysis and luciferase assays reveal that a P1BS element contributes to MtPHR1-mediated activation of the MtVIH2 promoter. Together, these findings reveal regulatory divergence of duplicated VIH genes and suggest that conserved phosphate-responsive mechanisms contribute to nodule development and nitrogen fixation in M. truncatula.
Prdm15 deficiency perturbs hematopoietic stem and progenitor cell homeostasis
Qiwen Dong, Weiwei Xiao, Junsong Huang, Zheng Sun, Min Liang, Mengjiao Wang, Qingqing Wang, Xinlan Zong, Anzi Li, Yan Wang, Zhongsheng Sun
, Available online  , doi: 10.1016/j.jgg.2026.07.009
Abstract (181) PDF (0)
Abstract:
The maintenance of homeostasis in hematopoietic stem and progenitor cells (HSPCs) is essential for the proper development of the entire hematopoietic system. However, the mechanisms underlying this regulatory equilibrium remain elusive. Here, we report that Prdm15 deficiency in HSPCs induces the accumulation of immature hematopoietic stem cells in mice. A series of transplantation assays shows that these cells display impaired reconstitution capacity and competitive fitness, which are associated with abnormal differentiation trajectories and transcriptional alterations identified by single-cell RNA sequencing. Mechanistically, integrated multi-omics analyses including ATAC-seq and CUT&Tag sequencing of HSPCs indicate that Prdm15 deficiency induces significant transcriptional and epigenetic alterations, particularly affecting the methyltransferase KMT2C and altering H3K4me1 and H3K27ac modifications at the promoters of hematopoietic developmental genes. Collectively, our findings establish PRDM15 as a critical epigenetic regulator of HSPCs, offering valuable insights into the molecular mechanisms underlying hematopoietic homeostasis.
LRRC71 stabilizes SEPT4 protein to ensure sperm annulus integrity and male fertility
Jinmei Cheng, Yu Wang, Yueming Wang, Zifeng Chen, Hehe Ren, Jiahui Li, Yifan Liang, Mingzhe Yang, Yashi Fan, Ziyi Luo, Yitian Xie, Ziqian Qi, Guishuan Wang, Yinchuan Li, Fei Sun
, Available online  , doi: 10.1016/j.jgg.2026.07.008
Abstract (196) PDF (0)
Abstract:
The structural integrity of the sperm flagellum is essential for male fertility, and its impairment is associated with reduced sperm motility. The sperm annulus is a septin-based fibrous ring that demarcates the midpiece and the principal piece. Notably, defects in sperm annulus formation frequently co-occur with abnormalities in flagellar structure; however, the underlying molecular mechanisms remain poorly understood. Herein, we identify an evolutionarily conserved leucine-rich repeat-containing protein, LRRC71, and show that its deficiency results in spermatozoa with a shortened mitochondrial sheath and a defective annulus, ultimately leading to male infertility. Further analysis reveals that LRRC71 deficiency leads to reduced sperm motility and decreased ATP levels following capacitation, effects that are potentially driven by a metabolic shift from oxidative phosphorylation to glycolysis. In addition, the protein levels of SEPT4, SEPT5, and SEPT7 are significantly reduced in Lrrc71-null spermatids. Mechanistically, LRRC71 directly binds SEPT4 via its N-terminal domain, thereby stabilizing the sperm annulus. Furthermore, the exogenous expression of SEPT4 rescues both motility and annulus defects in Lrrc71-null spermatozoa, confirming this functional hierarchy. Collectively, our findings demonstrate that LRRC71 serves as a central hub stabilizing sperm annulus integrity, providing insights into the pathogenic mechanisms underlying infertility associated with downregulation of LRRC71 in humans.
Shaker potassium channel OsKAT1 regulates root K+ uptake and lodging resistance in rice under low K+ stress
Wenxuan Yu, Jun Fu, Yanfeng Li, Shufeng Gao, Shengping Wang, Lei Jin, Ziyi Wang, Wenjing Li, Chen Wang, Niwen Tang, Junping Gao, Tianze Yan, Qunfeng Zhou, Kai Wang, Sheng Luan, Liangbi Chen, Zhonghua Liu, Yuanzhu Yang, Dandan Mao
, Available online  , doi: 10.1016/j.jgg.2026.07.007
Abstract (208) PDF (0)
Abstract:
Widespread potassium (K) deficiency in paddy soils, coupled with the low potassium use efficiency (KUE) of rice, has driven research on genetically improving KUE for sustainable rice production. Breeding high-KUE rice cultivars requires thorough understanding of root K+ uptake molecular mechanisms mediated by specific K+ channels and transporters. Here, we characterize the Shaker-type K+ channel OsKAT1 in rice. Disruption of OsKAT1 impairs root K+ uptake, resulting in reduced K+ accumulation and severe growth retardation under low-K+ stress. Conversely, overexpression of OsKAT1 enhances root K+ acquisition and promotes rice growth. Notably, OsKAT1-overexpressing lines exhibit increased culm diameter and improved bending resistance, thereby enhancing lodging tolerance. OsKAT1 overexpression also significantly increases grain size and weight under both K+-sufficient and low-K+ conditions. Natural variation at the OsKAT1 locus correlates with differential gene expression among haplotypes, with Hap 2 and Hap 3 conferring superior tolerance to low-K+ stress. Additionally, our results suggest that the expression pattern and physiological function of OsKAT1 may be cultivar-dependent. Collectively, these findings establish OsKAT1 as a key integrator of low-K+ adaptation, lodging resistance, and yield enhancement in rice, offering a promising genetic target for breeding rice varieties with improved KUE and yield potential.
LcMADS28 and LcMADS27 coordinately regulate early seed development in Litchi chinensis through direct activation of LcINVERTASE INHIBITOR1
Lijun Wu, Jun Fang, Jiahui Chen, Qiang Chen, Xinyuan Cheng, Zongli Liu, Zhike Zhang, Yonghua Qin, Guibing Hu, Jietang Zhao
, Available online  , doi: 10.1016/j.jgg.2026.05.013
Abstract (222) PDF (0)
Abstract:
Seed abortion is a critical agronomic trait affecting both fruit quality and yield. Cell wall invertase (CWIN) hydrolyzes sucrose into glucose and fructose, which serve as both nutrients and signaling molecules to regulate seed development, with its activity suppressed by invertase inhibitor (INH). However, the transcriptional regulators controlling INH expression remain unknown. Here, we identify the MADS-box transcription factor LcMADS28 as a direct activator of LcINH1. LcMADS28 is highly expressed in the funicle and embryo of the abortive-seeded cultivar ‘Nuomici’ at 15 days after anthesis (DAA), whereas its expression is much lower in the big-seeded cultivar ‘Heiye’. Further analysis reveals that LcMADS28 interacts with LcMADS27 to form a heterodimer that directly binds to the LcINH1 promoter and activates its transcription. Silencing LcMADS28 in ‘Nuomici’ reduces LcINH1 expression, which increases CWIN activity, promotes embryo and seed development, and lowers fruit abscission. RNA-seq analysis indicates that genes associated with auxin and sucrose metabolism are activated after LcMADS28 silencing. In addition, heterologous overexpression of LcMADS28 in tomato and Arabidopsis increases seed abortion. Together, our results indicate that LcMADS28 and LcMADS27 coordinately activate LcINH1 to regulate early seed development in lychee and provide a mechanistic framework linking MADS-box transcription factors with the invertase/INH module.
Non-redundant functions of three CRM proteins in mitochondrial group II intron splicing and maize seed development
Yingbo Li, Mingyue Zhang, Yuhong Wei, Tingting Xu, Yuejiao Dong, Yongyan Wang, Dongsheng Shi, Taoyu Shi, Pengshuai Yan, Hongyu Chen, Jihua Tang, Qingwen Shen, Guifeng Wang
, Available online  , doi: 10.1016/j.jgg.2026.05.003
Abstract (126) PDF (0)
Abstract:
Group II introns are abundant in plant mitochondrial genomes and require numerous nuclear-encoded RNA-binding proteins for their splicing. Chloroplast RNA splicing and ribosome maturation (CRM) proteins constitute an evolutionarily conserved RNA-binding protein family, yet their intron splicing specificity and underlying molecular mechanisms remain poorly understood. Here, we show that three mitochondria-localized CRM proteins play non-redundant roles in mitochondrial group II intron splicing and maize seed development. Genome-wide analysis identifies 16 maize CRM genes, which are classified into four subfamilies and harbor variable numbers of CRM domains. ZmCFM5 and ZmmCSF2 are targeted to mitochondria, whereas ZmCFM3 exhibits dual localization to mitochondria and chloroplasts. Loss-of-function mutants of these three CRM genes display impaired intron splicing and arrested seed development. ZmCFM5 mediates the splicing of nad1 intron4 and nad4 intron1, while ZmmCSF2 and ZmCFM3 act on nad5 intron4 and nad1 intron2, respectively. The cfm5 mutant exhibits abnormal mitochondrial morphology and reduced complex I activity. ZmCFM5 specifically binds to a conserved motif within domain V of target introns and physically interacts with the splicing factor ZmSPR2. Our findings support a model wherein CRM proteins cooperate with other RNA-binding proteins to assemble dynamic ribonucleoprotein complexes, which confer splicing specificity and efficiency to group II introns.
Review
Genome-wide association studies of osteoarthritis: from genetic discovery to precision medicine
Yuezhou Wu, Yang Hong, Xukun Lyu, Peng Wang, Houfeng Zheng, Jiacan Su
, Available online  , doi: 10.1016/j.jgg.2026.10.005
Abstract (59) PDF (0)
Abstract:
Abstract:
Microglial modulation in general anesthesia: molecular
Xiaoman Yuan, Taotao Liu, Yanxue Xue, Xiangyang Guo, Zhaofa Wu, Zhengqian Li
, Available online  , doi: 10.1016/j.jgg.2026.09.001
Abstract (143) PDF (0)
Abstract:
General anesthetics profoundly alter brain function and consciousness, yet the mechanisms underlying these effects remain incompletely understood. Although traditional studies have primarily focused on neuronal targets, accumulating evidence suggests that microglia dynamically respond to anesthetic exposure and may participate in anesthesia-associated neurophysiological changes. Beyond their established immune functions, microglia are increasingly implicated in synaptic remodeling, metabolic regulation, neuronal activity surveillance, and neuron–glia communication. Recent studies indicate that different classes of anesthetic agents modulate microglial activity through diverse and context-dependent mechanisms involving inflammatory signaling, purinergic pathways, calcium dynamics, mitochondrial metabolism, and neural circuit interactions. These responses are associated with postoperative neurocognitive disorders, altered synaptic plasticity, and anesthesia-related changes in brain states. In this review, we summarize current evidence regarding the effects of volatile anesthetics, intravenous anesthetics, and analgesics on microglial function and discuss the molecular, functional, and circuit-level mechanisms underlying anesthesia-associated neuron–microglia interactions. We further highlight the dynamic and heterogeneous nature of microglial responses during anesthesia and discuss current limitations in the field, including the lack of temporally resolved and cell-specific approaches. Understanding these processes may provide insights into anesthesia-associated neurocognitive dysfunction and support the development of neuroimmune-targeted strategies in anesthesiology.
Heat stress in cereal crops: reproductive development and grain filling
Shijun Fan, Nese Sreenivasulu, Qiaoquan Liu, Shigui Li, Peng Qin
, Available online  , doi: 10.1016/j.jgg.2026.09.002
Abstract (232) PDF (0)
Abstract:
Increasingly frequent extreme heat events threaten cereal production and food security under a changing climate. The reproductive-to-grain formation continuum of cereals is particularly vulnerable to elevated temperatures, as heat stress disrupts developmental processes from inflorescence formation and fertilization to grain filling and quality establishment. These disruptions reduce reproductive success, impair yield formation, and compromise grain quality. A comprehensive understanding of the developmental, physiological, molecular, and genetic basis of cereal heat tolerance is therefore essential for developing climate-adapted crops. This review summarizes recent advances in understanding heat stress during cereal reproduction and grain filling across major cereal crops. We first discuss how heat stress affects sequential developmental processes, including inflorescence development, gametophyte development, flowering and pollination, fertilization, and grain filling. We then integrate emerging evidence on cross-cutting mechanisms that connect stage-specific heat responses, focusing on hormonal and redox homeostasis, carbohydrate metabolism and source–sink coordination, proteostasis and endomembrane organization, and genome stability and multilayered gene regulation. Finally, we summarize the genetic basis of cereal heat tolerance by highlighting genetic determinants, favorable alleles, and their potential applications in breeding. We further discuss current bottlenecks and future opportunities for breeding heat-tolerant cereals.
Regulation of root-to-shoot ratio in plant resource allocation by external and internal cues
Qing Li, Yi Tong, Hongning Tong
, Available online  , doi: 10.1016/j.jgg.2026.07.010
Abstract (355) PDF (12)
Abstract:
Plants dynamically adjust their root-to-shoot ratio (R:S) in fluctuating environments, a response often interpreted as adaptive resource allocation, although other explanations have also been proposed. Despite ongoing debates surrounding optimal partitioning theory, the active-regulation framework remains valuable for understanding R:S regulation, which is significant for improving plant resource use efficiency and thus fitness and productivity. This review synthesizes multi-scale advances, from physiology to molecular mechanisms, to outline the regulatory networks associated with coordinated root–shoot growth. We examine how major environmental factors such as light, temperature, osmotic stress, nitrogen, and phosphorus regulate R:S through distinct pathways, highlighting the photoreceptor-mediated carbon-nitrogen allocation, organ-specific thermomorphogenic responses, and the multi-layered osmotic stress networks. For nutrient regulation, we emphasize the OsWRI1a–RNR10–DNR1 module for tissue-specific R:S nitrogen response and the PHR1-centered phosphate starvation response network. We also summarize how endogenous hormones (auxin, cytokinin, abscisic acid, and strigolactone) and signaling hubs like TOR kinase regulate R:S. Finally, we discuss current challenges and future directions for a system-level understanding of R:S regulation and its targeted manipulation.
Research Communications
The zc3h6 deficiency causes gut microbiota dysbiosis and inflammatory bowel disease-like symptoms in zebrafish
Xiangyong Wei, Guozhen Wu, Da Zhang, Si Chen, Xinmiao Tan, Xue Ma, Xiaoqian Fu, Qingliang Zou, Lingfei Luo, Jianlong Ma
, Available online  , doi: 10.1016/j.jgg.2026.10.001
Abstract (73) PDF (0)
Abstract:
SheepVar: an integrated resource for large-scale sheep genomic variation with multi-omics annotation
Maqiang Zhao, Jin Wang, Jiarun Zheng, Shengyu Guan, Weiwei Fu, Lingzhao Fang, Zhangyuan Pan, Quanzhong Liu, Yu Jiang, Ran Li
, Available online  , doi: 10.1016/j.jgg.2026.10.003
Abstract (213) PDF (0)
Abstract:
BioGAIP: a scalable and robust LLM-powered multi-agent system for automated bioinformatics tasks
Jiayu Zhang, Pengfei Guo, Guanghui Jiang, Mengyu Zhou, Gang Wei, Ting Ni
, Available online  , doi: 10.1016/j.jgg.2026.09.013
Abstract (167) PDF (0)
Abstract:
Genomic insights into heading date variation of the different maturity types of japonica rice varieties in Jiangsu Province, China
Hongjia Zhang, Jiaqi Tang, Ying Chen, Zuopeng Xu, Qianfeng Li, Shuzhu Tang, Shimin Zuo, Yuebin Wang, Changquan Zhang, Qiaoquan Liu
, Available online  , doi: 10.1016/j.jgg.2026.08.010
Abstract (264) PDF (1)
Abstract: