Current Articles

2026, Volume 53,  Issue 11

Display Method:
Review
Incomplete penetrance in neurogenetic disorders: current insights and emerging perspectives
Jiao-Jiao Xu, Dian-Fu Chen, Zhi-Ying Wu
2026, 53(11): 2059-2070. doi: 10.1016/j.jgg.2026.07.005
Abstract (0) PDF (0)
Abstract:
Neurogenetic disorders have been recognized clinically for decades, and advances in clinical and genetic studies have identified more than 1700 monogenic causes of neurological diseases. Various types of mutations, including missense mutations, truncating mutations, and repeat expansions, have been reported in patients with neurogenetic disorders. It is now recognized that incomplete penetrance is common, with some individuals carrying disease-causing mutations remaining clinically unaffected. However, there is currently no comprehensive conceptual framework to categorize or explain these observations. Here, we review and integrate decades of evidence on incomplete penetrance in neurogenetic disorders to clarify its biological and mechanistic bases. Accordingly, four major themes are identified, encompassing genetic modifiers, epigenetic modifications, mosaicism, and environmental factors. These factors may act independently or interactively to influence pathogenic burden and functional network balance, ultimately determining whether a pathogenic mutation manifests clinically. Based on these insights, we highlight emerging perspectives and propose future research to fill gaps in our understanding. A deeper understanding of incomplete penetrance will be essential for generating genetic insights to support more effective genetic counseling, therapeutic interventions, and disease prevention in neurogenetic disorders.
Beyond radiogenomics: advancing imaging integration and multi-omics horizons in cancer precision medicine
Yuming Chen, Huiqin Li, Silu Chen, Hongsheng Ji, Junyi Xin, Mulong Du
2026, 53(11): 2071-2083. doi: 10.1016/j.jgg.2026.07.003
Abstract:
Cancer remains the leading cause of death worldwide, presenting substantial challenges to precision medicine due to its complex heterogeneity. Radiogenomics, as a method combining quantitative radiologic data with genomic information, provides a robust analysis framework to assess tumor heterogeneity and cancer progression. Here, we summarize the application of radiogenomics in two key fusion methods: feature-level and decision-level fusion. Feature-level fusion combines multimodal data into a rich feature set to improve the predictive power of models, while decision-level fusion integrates decision results from multiple independent models to improve robustness and reliability. Furthermore, we explore the integration of radiomics with various omics technologies, including transcriptomics, metabolomics, and proteomics. This integration enables a deeper understanding of the dynamic tumor microenvironment, metabolic dysregulation, and cancer progression mechanisms. Finally, we provide a detailed overview of publicly available datasets relevant to radiogenomics research, such as The Cancer Imaging Archive, cBioPortal, UK Biobank, and Human Connectome Project; and further describe multiple types of omics data and sample characteristics for each resource for the benefit to readers. In summary, this review charts a path beyond radiogenomics by advancing radiomics and multi-omics horizons to transform precision medicine in cancer.
The development and application of deep learning for stem cell research
Zhenfu Li, Wenjing Zhang, Yuxin Wu, Yaxuan Ye, Yuxin Chen, Jian Wu, Di Chen
2026, 53(11): 2084-2105. doi: 10.1016/j.jgg.2026.05.008
Abstract:
Stem cells reside in specific microenvironments where they divide to maintain themselves and differentiate into functional cells that replace old, dead, or damaged cells to maintain tissue homeostasis. Some stem cells could also be cultured in vitro for self-renewal and directed differentiation towards specific lineages for both mechanistic interrogation and clinical investigation. A deeper understanding of the regulatory mechanisms underlying stem cell properties is essential for advancing their translational applications. Deep learning (DL), a branch of artificial intelligence, has been widely and deeply incorporated into different fields including biology. The application of DL in stem cells has revolutionized our research strategies and provided significant technical advantages. Here, we review the latest advances of the application of DL in analyzing bioimaging data and exploring large-scale genomics data in stem cells. We also summarize the limitations of current DL models and challenges of the application of these models in stem cell research.
Regulation of root-to-shoot ratio in plant resource allocation by external and internal cues
Qing Li, Yi Tong, Hongning Tong
2026, 53(11): 2106-2123. doi: 10.1016/j.jgg.2026.07.010
Abstract (0) PDF (0)
Abstract:
Plants dynamically adjust their root-to-shoot ratio (R:S) in fluctuating environments, a response often interpreted as adaptive resource allocation, although other explanations have also been proposed. Despite ongoing debates surrounding optimal partitioning theory, the active-regulation framework remains valuable for understanding R:S regulation, which is significant for improving plant resource use efficiency and thus fitness and productivity. This review synthesizes multi-scale advances, from physiology to molecular mechanisms, to outline the regulatory networks associated with coordinated root–shoot growth. We examine how major environmental factors such as light, temperature, osmotic stress, nitrogen, and phosphorus regulate R:S through distinct pathways, highlighting the photoreceptor-mediated carbon-nitrogen allocation, organ-specific thermomorphogenic responses, and the multi-layered osmotic stress networks. For nutrient regulation, we emphasize the OsWRI1a–RNR10–DNR1 module for tissue-specific R:S nitrogen response and the PHR1-centered phosphate starvation response network. We also summarize how endogenous hormones (auxin, cytokinin, abscisic acid, and strigolactone) and signaling hubs like target of rapamycin (TOR) kinase regulate R:S. Finally, we discuss current challenges and future directions for a system-level understanding of R:S regulation and its targeted manipulation.
Original Research
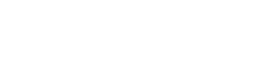
The TNFR2M196R germline variant confers dual impact on CAR-T cell immunotherapy by enhancing T-cell survival and tumor proliferation
Nan Liu, Lu Han, Yuting Lu, Yelei Guo, Jiajia Li, Qi Zhu, Yao Wang, Zhiqiang Wu, Xin Lin, Jianshu Wei, Weidong Han
2026, 53(11): 2124-2135. doi: 10.1016/j.jgg.2026.05.014
Abstract (19) PDF (2)
Abstract:
Despite its remarkable efficacy in hematologic malignancies, chimeric antigen receptor T-cell (CAR-T) therapy is often limited by atypical clinical outcomes, necessitating predictive biomarkers. Here, prompted by a patient with low-tumor-burden non-Hodgkin lymphoma who developed severe and persistent cytokine release syndrome (CRS), we identify the germline TNFR2M196R single-nucleotide polymorphism, rs1061622, as a candidate genetic determinant of atypical responses. The TNFR2M196R variant is found to enhance the antitumor efficacy of CAR-T cells by reducing their apoptosis. Paradoxically, however, patients carrying the TNFR2M196R variant exhibit higher rates of early tumor progression. Further investigation reveals that while the TNFR2M196R mutation increases tumor susceptibility to CAR-T-mediated killing, it also accelerates tumor proliferation kinetics. Together, these results establish the TNFR2M196R SNP as a dual-function genetic determinant that modulates both CAR-T-cell efficacy and intrinsic tumor behavior, highlighting the critical impact of germline genetics on cancer immunotherapy.
Neuroprotective role of Faecalibacterium prausnitzii-derived butyrate in diabetic gastrointestinal autonomic neuropathy
Lin Guo, Jie Li, Jiaming An, Jiahe Miao, Yang Yi, Keying Zhu, Qing Cai, Sijing Wang, Zhan Su, Xiaolong Ye, Yayun Wang, Minglin Pan, Qingmiao Lu, Bota Cui, Faming Zhang, Jianhua Mao, Xingyin Liu, Yibing Lu, Dafa Ding
2026, 53(11): 2136-2149. doi: 10.1016/j.jgg.2026.06.005
Abstract:
Diabetic gastrointestinal autonomic neuropathy (DGAN) is a common yet poorly understood complication of diabetes that is characterized by gastrointestinal dysmotility and enteric neurodegeneration. Here, we investigate whether gut microbiota dysbiosis contributes to DGAN pathogenesis and explore the potential involvement of microbiota-derived metabolites in enteric nervous system (ENS) injury. Gut microbiota profiling reveals disease-associated compositional alterations in patients with DGAN, including depletion of Faecalibacterium-associated signals, with Faecalibacterium prausnitzii identified as a putative species-level annotation and negatively associated with gastrointestinal symptom severity. Fecal microbiota transplantation from patients with DGAN into db/db mice aggravates gut dysmotility and increases apoptosis of ChAT+ and nNOS+myenteric neurons. Further experiments indicate that butyrate, the predominant metabolite produced by F. prausnitzii, attenuates neuronal apoptosis under high-glucose conditions. This biological process is accompanied by enhanced BCL2 expression, downregulation of cleaved caspase-3, and activation of the PI3K/Akt signaling cascade. Collectively, our findings support the presence of gut microbiota-ENS axis in DGAN and identify butyrate as an important candidate neuroprotective metabolite associated with F. prausnitzii. These results provide a rationale for microbiota-targeted therapeutic strategies for diabetic enteric neuropathy.
Population genomic analysis reveals genomic variations associated with domestication and key agronomic traits in bitter gourd (Momordica charantia)
Yu Niu, Rui Wang, Liang Wang, Aizheng Huang, Xu Han, Renbo Yu, Shijun Chen, Juntao Wang, Jiaowen Cheng, Changming Chen, Chengjie Chen, Yan Yang, Yi Liao, Ming Hu
2026, 53(11): 2150-2164. doi: 10.1016/j.jgg.2026.04.017
Abstract (0) PDF (0)
Abstract:
Bitter gourd (Momordica charantia) is a widely cultivated vegetable and medicinal crop for its nutritional and pharmaceutical properties. Yet, the genomic basis of its domestication and key agronomic traits remains largely unexplored. Here, we present a high-quality, chromosome-level genome assembly specifically representing a smooth bitter gourd cultivar (Y1745), with a total assembly size of 327.14 Mb, a contig N50 of 11.33 Mb, and a scaffold N50 of 23.47 Mb. Comparative genomics reveals lineage-specific expansions of gene families related to stress responses and secondary metabolism. Population genomic analysis of 192 globally representative accessions identifies 3 distinct genetic groups (China, Southeast Asia, and South Asia Subcontinent), reflecting complex demographic histories, and uncovers strong signatures of selection associated with domestication and regional adaptation. In the genome-wide association study of 35 traits, we identify 893 significant trait-associated loci. Notably, two candidate genes (McWRKY and McFPF1-like 1) are associated with flowering time regulation, and one candidate gene (McEXLB1) with fruit size determination. Our work provides valuable genomic resources for understanding bitter gourd domestication and offers potential genetic targets for breeding.
The transcription factor SlDOF9 directly activates SlELP5 expression to regulate inflorescence length in tomato
Junling Hu, Can Zhu, Xin Li, Lin Yang, Chen Zhang, Shumin He, Yonglian Wang, Yujie Zhang, Hui Zhang, Zejun Huang, Yanmei Guo, Xiaoxuan Wang, Yongchen Du, Lei Liu, Junming Li
2026, 53(11): 2165-2179. doi: 10.1016/j.jgg.2026.05.002
Abstract:
Tomato inflorescence architecture is an important determinant of yield potential and fruit distribution, with inflorescence length and branching influencing resource allocation and harvestability. Although several regulators of inflorescence development have been identified, the genetic basis of natural variation in inflorescence length remains insufficiently characterized. Using a mapping population derived from LA1256 and Moneymaker, two lines with contrasting inflorescence lengths, we identify a quantitative trait locus, qIFL2.1, for inflorescence length on chromosome 2. Fine mapping and functional analyses identify SlELP5 as a strong candidate gene underlying qIFL2.1. Natural variation in the SlELP5 promoter is associated with altered SlELP5 expression, which is negatively correlated with inflorescence length. Consistently, SlELP5 overexpression in the LA1256 background reduces inflorescence length, supporting a negative role of SlELP5 in regulating this trait. Transcriptomic analysis, dual-luciferase assay, and EMSA further show that SlDOF9 is upregulated in OE-SlELP5 lines and directly activates SlELP5 expression by binding to its promoter, supporting a SlDOF9–SlELP5 regulatory module. These findings provide insights into the molecular regulation of tomato inflorescence architecture and identify a potential target for precision breeding of inflorescence traits.
Non-redundant functions of three CRM proteins in mitochondrial group II intron splicing and maize seed development
Yingbo Li, Mingyue Zhang, Yuhong Wei, Tingting Xu, Yuejiao Dong, Yongyan Wang, Dongsheng Shi, Taoyu Shi, Pengshuai Yan, Hongyu Chen, Jihua Tang, Qingwen Shen, Guifeng Wang
2026, 53(11): 2180-2191. doi: 10.1016/j.jgg.2026.05.003
Abstract:
Group II introns are abundant in plant mitochondrial genomes and require numerous nuclear-encoded RNA-binding proteins for their splicing. Chloroplast RNA splicing and ribosome maturation (CRM) proteins constitute an evolutionarily conserved RNA-binding protein family, yet their intron splicing specificity and underlying molecular mechanisms remain poorly understood. Here, we show that three mitochondria-localized CRM proteins play non-redundant roles in mitochondrial group II intron splicing and maize seed development. Genome-wide analysis identifies 16 maize CRM genes, which are classified into four subfamilies and harbor variable numbers of CRM domains. ZmCFM5 and ZmmCSF2 are targeted to mitochondria, whereas ZmCFM3 exhibits dual localization to mitochondria and chloroplasts. Loss-of-function mutants of these three CRM genes display impaired intron splicing and arrested seed development. ZmCFM5 mediates the splicing of nad1 intron4 and nad4 intron1, while ZmmCSF2 and ZmCFM3 act on nad5 intron4 and nad1 intron2, respectively. The cfm5 mutant exhibits abnormal mitochondrial morphology and reduced complex I activity. ZmCFM5 specifically binds to a conserved motif within domain V of target introns and physically interacts with the splicing factor ZmSPR2. Our findings support a model wherein CRM proteins cooperate with other RNA-binding proteins to assemble dynamic ribonucleoprotein complexes, which confer splicing specificity and efficiency to group II introns.
Genetic dissection of cadmium accumulation in tea plants under nonphytotoxic field conditions
Shijie Luo, Xinwan Zhang, Weiyi Zhang, Yujia Ren, Dawei Gao, Xiali Ye, Alisdair R. Fernie, Weiwei Wen
2026, 53(11): 2192-2202. doi: 10.1016/j.jgg.2026.05.006
Abstract:
Cadmium (Cd) accumulation in tea plants (Camellia sinensis) under chronic, nonphytotoxic field conditions poses a persistent food safety challenge; however, the genetic basis of this process remains uncharacterized. To dissect the genetic architecture of this trait, we quantify leaf Cd content and perform genome-wide association studies (GWAS) across 207 diverse tea accessions which have been cultivated for decades. Quantification analysis of Cd content reveals substantial natural variation in Cd content between young leaves (YL) and mature leaves (ML) under nonphytotoxic environment, with 38.2% of the 173 accessions with paired data intrinsically accumulating more Cd in YL than in ML. GWAS identify 112 quantitative trait loci associated with Cd accumulation. By further integrating the results of GWAS with Cd-responsive transcriptome profiling, we prioritize 14 high-confidence candidate genes. One of these candidate genes, CsHIPP6, which encodes a heavy metal-associated isoprenylated plant protein, is selected for further functional investigation. Heterologous expression in yeast and Arabidopsis demonstrates that CsHIPP6 enhances Cd tolerance and significantly reduces Cd accumulation. Our findings elucidate the genetic architecture underlying long-term Cd accumulation in perennial crops and highlight CsHIPP6 as a valuable target for breeding low-Cd tea cultivars to ensure beverage safety amidst rising environmental contamination.
LcMADS28 and LcMADS27 coordinately regulate early seed development in Litchi chinensis through direct activation of LcINVERTASE INHIBITOR1
Lijun Wu, Jun Fang, Jiahui Chen, Qiang Chen, Xinyuan Cheng, Zongli Liu, Zhike Zhang, Yonghua Qin, Guibing Hu, Jietang Zhao
2026, 53(11): 2203-2214. doi: 10.1016/j.jgg.2026.05.013
Abstract:
Seed abortion is a critical agronomic trait affecting both fruit quality and yield. Cell wall invertase (CWIN) hydrolyzes sucrose into glucose and fructose, which serve as both nutrients and signaling molecules to regulate seed development, with its activity suppressed by invertase inhibitor (INH). However, the transcriptional regulators controlling INH expression remain unknown. Here, we identify the MADS-box transcription factor LcMADS28 as a direct activator of LcINH1. LcMADS28 is highly expressed in the funicle and embryo of the abortive-seeded cultivar ‘Nuomici’ at 15 days after anthesis, whereas its expression is much lower in the big-seeded cultivar ‘Heiye’. Further analysis reveals that LcMADS28 interacts with LcMADS27 to form a heterodimer that directly binds to the LcINH1 promoter and activates its transcription. Silencing LcMADS28 in ‘Nuomici’ reduces LcINH1 expression, which increases CWIN activity, promotes embryo and seed development, and lowers fruit abscission. RNA-seq analysis indicates that genes associated with auxin and sucrose metabolism are activated after LcMADS28 silencing. In addition, heterologous overexpression of LcMADS28 in tomato and Arabidopsis increases seed abortion. Together, our results indicate that LcMADS28 and LcMADS27 coordinately activate LcINH1 to regulate early seed development in lychee and provide a mechanistic framework linking MADS-box transcription factors with the invertase/INH module.
CHST5 gene mutations contribute to high myopia by disrupting collagen fiber organization
Meining He, Senwei Tan, Furong Huang, Zhe Cheng, Lan Yao, Haibin Zhao, Kai Wu, Xiaolong Yu, Hongbing Jiang, Qiuping Liu, Xiangtian Zhou, Qi Tian, Zhengmao Hu, Kun Xia
2026, 53(11): 2215-2224. doi: 10.1016/j.jgg.2026.06.007
Abstract:
High myopia (HM) is a leading cause of irreversible vision loss in working-age adults. Its pathogenesis is characterized by alterations in the microstructure and composition of collagen fibers, and genetic factors make a substantial contribution. In this study, we identify carbohydrate sulfotransferase 5 (CHST5) as a candidate gene for HM in humans and mice, with its mutations disrupting collagen fiber organization. The c.444C>A (p.S148R) variant in CHST5, a gene critical for sulfating corneal keratan sulfate (KS), completely co-segregates with HM in a Chinese family. Screening of CHST5 variants in 320 HM patients identifies two additional ones. We further find that Chst5 is expressed primarily in the cornea and sclera of mouse ocular tissues and that the mutant protein CHST5S148R loses its Golgi localization. Homozygous mutant Chst5S126R mice exhibit HM phenotypes, including myopic refractive error, significantly thinner sclera and cornea, notable microstructural changes in scleral and corneal collagen fibers, and shorter corneal KS chains. Our findings suggest that CHST5 NM_024533.5 c.444C>A (p.S148R) causes the loss of proper protein localization, likely impairing its sulfotransferase function. This defect disrupts the organization of corneal and scleral collagen fibers and ultimately contributes to the development and progression of HM.
Characterizing selection signatures in coding and noncoding regions of 14,886 cancer genomes
Mengyue Zheng, Xiwei Sun, Junren Hou, Minmin Guo, Xinran Liu, Wen Yang, Jian Yang
2026, 53(11): 2225-2239. doi: 10.1016/j.jgg.2026.06.008
Abstract (0) PDF (0)
Abstract:
Clonal selection drives cancer development, but quantifying selection on noncoding somatic mutations remains largely unexplored. Here, we introduce dNdS-Fun, an extension of the dN/dS framework to quantify selection of both coding and noncoding somatic mutations, thereby enhancing the discovery of driver genes. Applying dNdS-Fun to whole-genome sequencing data from 14,886 cancer patients across 31 cancer types, we identify 175 genes under positive selection across multiple cancer types or datasets, as well as 20 previously known driver genes detected through noncoding mutations. Of these, 69 are previously unrecognized as drivers, and 30 are identified solely through noncoding mutations. Furthermore, we observe evidence of negative selection throughout the genome, with significant enrichment in essential and cancer-dependent genes. Sixteen genes exhibit an overall signature of negative selection but show positive selection in noncoding elements, indicating both their conserved functions and adaptive regulatory roles in tumorigenesis. Our study reveals evidence consistent with negative selection of noncoding mutations, providing important insights for future research on their roles in cancer progression.
Tapt1 deficiency in mice impairs pulmonary lipid homeostasis and normal postnatal respiration by targeting ABCA3 for autophagy-lysosomal degradation
Dandan Tan, Lu Cai, Li Yuan
2026, 53(11): 2240-2254. doi: 10.1016/j.jgg.2026.06.012
Abstract (17) PDF (2)
Abstract:
TAPT1, which encodes a highly conserved multi-pass transmembrane protein termed transmembrane anterior posterior transformation 1 (TAPT1), has been reported as a disease-causing gene, but its physiological role in mice remains to be elucidated. Using Tapt1 knockout and knock-in mice, we indicate that TAPT1 localizes to the endoplasmic reticulum and that Tapt1 deletion causes neonatal lethality due to atelectasis-induced respiratory distress. We further reveal that TAPT1 interacts with the ATP-binding cassette transporter A3 (ABCA3), thereby regulating autophagy-lysosomal degradation of ABCA3. ABCA3 insufficiency reduces surfactant lipids, leading to defective lamellar body formation and surfactant production. This study presents a hitherto unrecognized pathway for ABCA3 protein degradation. Our findings may advance the understanding of genetic determinants of neonatal respiratory distress syndrome, providing valuable insights into the treatment of lung surfactant disorders.
Mov10 mitigates hematopoietic stem cell exhaustion under stress by modulating the Camp-mediated inflammatory pathway
Changlin Zhen, Junbing Pan, Wei Wang, Jiankun Fan, Yonghui Zhang, Dan Liu, Shiheng Zhang, Xiaohuan Huang, Yalan Xiao, Yu Hou, Xueya Zhao, Zhigang Li
2026, 53(11): 2255-2269. doi: 10.1016/j.jgg.2026.05.012
Abstract:
The integrity of hematopoietic stem cell (HSC) function is crucial for robust hematopoietic regeneration following stress. Inflammatory responses are pivotal drivers of HSC stress response, yet the precise modulation of inflammatory pathways remains incompletely defined. In this study, we identify the RNA helicase MOV10 as a negative regulator of stress-induced inflammatory pathways in HSCs. Our study indicates that Mov10, which is critically required for HSC maintenance, is highly expressed in HSCs, and its loss adversely affects HSC fitness and survival during hematopoietic stress induced by bone marrow transplantation and irradiation. Mechanistically, Mov10 mitigates excessive inflammatory activation to sustain HSC functional integrity during hematopoietic stress, primarily by enhancing the translation of CAMP, which inhibits the interaction between TNF-α and its receptor TNFR1 and suppresses NF-κB activation. Overall, our results imply that Mov10 plays a critical role in averting functional failure of hematopoiesis under stress, presenting viable paths for therapeutic intervention in relevant diseases.
Multi-omics identification and verification of Dnajb14 as a modulator of retinal ganglion cell survival in glaucoma through ferroptosis
Yu Chen, Jie Wen, Qian Yang, Zixi Chen, Haibo Li, Zhenkai Wu, Quan Cheng, Dan Ji
2026, 53(11): 2270-2285. doi: 10.1016/j.jgg.2026.05.011
Abstract:
Glaucoma is a leading cause of irreversible blindness and necessitates the identification of unreported therapeutic targets. Here, we report eight stable plasma protein targets for glaucoma, identified through a large-scale proteome-wide association study integrated with Mendelian randomization and colocalization analyses. Among these candidates, we show that the DnaJ heat shock protein family member B14 (DNAJB14) acts as a critical neuroprotective factor. Using single-cell and bulk transcriptomics, we demonstrate that Dnajb14 is characteristically expressed in retinal ganglion cells (RGCs) and that its abundance is inversely correlated with ferroptosis under ischemic stress. Furthermore, our results reveal that Dnajb14 preserves RGC survival and structural and functional integrity in multiple acute and chronic murine models. Mechanistically, Dnajb14 overexpression robustly alleviates ischemia-reperfusion-induced retinal injury, whereas targeted knockdown exacerbates lipid peroxidation, mitochondrial dysfunction, and ferroptosis. Taken together, our findings highlight that Dnajb14 is a key regulator of RGC survival through inhibition of ferroptosis, thereby making it a promising therapeutic candidate for glaucoma treatment.
Research Communications
Evolution of resistance to KRASG12C inhibitor in a non-small cell lung cancer responder
Jiahui Xu, Shijia Wang, Ziming Wang, Jumin Huang, Chun Xie, Tingting Zhao, Yuqing Niu, Yabing Cao, Chen Ming, Elaine Lai-Han Leung
2026, 53(11): 2286-2289. doi: 10.1016/j.jgg.2026.07.002
Abstract:
A high-quality genome assembly delineates the genetic basis of freezing tolerance in the elite wheat cultivar Jimai 22
Guang Qi, Guangwei Li, Rongzhi Zhang, Wei Li, Weilong Guo, Jun Zheng, Haosheng Li, Jianjun Liu, Takao Komatsuda, Cheng Liu
2026, 53(11): 2290-2293. doi: 10.1016/j.jgg.2026.06.001
Abstract:
Molecular basis of blade–pseudo-petiole–sheath axis patterning in Spodiopogon sagittifolius: roles of auxin signaling and lineage-specific regulators
Chao Xu, Yang Wang, Liping Qin, Renyi Ma, Wei Bi, Feng Liu, Zhenwen Liu
2026, 53(11): 2294-2297. doi: 10.1016/j.jgg.2026.05.007
Abstract:
Knockout of TpNRAMP5 reduces grain cadmium concentration in Polish wheat
Yiran Cheng, Xing Chen, Hongzhu Zhou, Jian Ma, Rui Wang, Yongjun Dong, Xi Li, Dan Long, Jian Zeng, Xing Fan, Houyang Kang, Dandan Wu, Yonghong Zhou, Ke Wang, Yi Wang
2026, 53(11): 2298-2301. doi: 10.1016/j.jgg.2026.04.021
Abstract:
Metabolic reprogramming of sucrose flux boosts natural rubber in Taraxacum kok-saghyz
Xiuli Fan, Qingwen Chen, Xiaoguang Song, Liquan Kou, Guodong Wang, Jinfang Chu, Xia Cui, Hong Yu, Xia Xu, Jiayang Li
2026, 53(11): 2302-2305. doi: 10.1016/j.jgg.2026.05.009
Abstract: